Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -2611.643428 |
| Energy at 298.15K | -2611.647301 |
| HF Energy | -2611.367876 |
| Nuclear repulsion energy | 80.077056 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3189 |
3060 |
6.47 |
|
|
|
| 2 |
A' |
1395 |
1338 |
21.61 |
|
|
|
| 3 |
A' |
712 |
683 |
16.65 |
|
|
|
| 4 |
A' |
322 |
309 |
58.16 |
|
|
|
| 5 |
A" |
3340 |
3204 |
0.02 |
|
|
|
| 6 |
A" |
933 |
895 |
0.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4944.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 4744.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.007 |
1.496 |
0.000 |
| Br2 |
-0.007 |
-0.371 |
0.000 |
| H3 |
0.142 |
1.998 |
0.959 |
| H4 |
0.142 |
1.998 |
-0.959 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8671 | 1.0926 | 1.0926 |
Br2 | 1.8671 | | 2.5597 | 2.5597 | H3 | 1.0926 | 2.5597 | | 1.9189 | H4 | 1.0926 | 2.5597 | 1.9189 | |
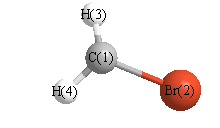 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.310 |
|
Br2 |
C1 |
H4 |
117.310 |
| H3 |
C1 |
H4 |
122.826 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -2611.643366 |
| Energy at 298.15K | |
| HF Energy | -2611.367668 |
| Nuclear repulsion energy | 80.132570 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3198 |
3068 |
4.98 |
|
|
|
| 2 |
A1 |
1394 |
1337 |
22.78 |
|
|
|
| 3 |
A1 |
714 |
685 |
15.38 |
|
|
|
| 4 |
B1 |
232i |
222i |
67.64 |
|
|
|
| 5 |
B2 |
3352 |
3216 |
0.22 |
|
|
|
| 6 |
B2 |
925 |
888 |
0.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4675.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 4485.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.494 |
| Br2 |
0.000 |
0.000 |
0.371 |
| H3 |
0.000 |
0.964 |
-2.007 |
| H4 |
0.000 |
-0.964 |
-2.007 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8645 | 1.0916 | 1.0916 |
Br2 | 1.8645 | | 2.5653 | 2.5653 | H3 | 1.0916 | 2.5653 | | 1.9272 | H4 | 1.0916 | 2.5653 | 1.9272 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
118.027 |
|
Br2 |
C1 |
H4 |
118.027 |
| H3 |
C1 |
H4 |
123.947 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
