return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3OCH2 (methoxymethyl radical)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-153.938315
Energy at 298.15K-153.943354
HF Energy-153.448558
Nuclear repulsion energy77.003351
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3274 3141 19.40      
2 A 3174 3045 23.14      
3 A 3123 2997 23.79      
4 A 3107 2981 53.45      
5 A 3034 2911 49.24      
6 A 1505 1444 8.96      
7 A 1504 1443 5.95      
8 A 1488 1428 5.23      
9 A 1462 1402 9.81      
10 A 1298 1245 143.02      
11 A 1266 1214 29.42      
12 A 1179 1131 2.89      
13 A 1140 1094 4.29      
14 A 994 954 27.84      
15 A 709 680 29.98      
16 A 442 424 3.22      
17 A 305 292 8.13      
18 A 183 176 1.46      

Unscaled Zero Point Vibrational Energy (zpe) 14593.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 14000.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
1.51912 0.35887 0.30956

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.089 -0.557 -0.045
C2 -1.129 0.173 0.015
C3 1.195 0.234 0.082
H4 -1.947 -0.553 -0.118
H5 -1.241 0.680 0.992
H6 -1.177 0.931 -0.792
H7 2.132 -0.319 -0.037
H8 1.123 1.275 -0.270

Atom - Atom Distances (Å)
  O1 C2 C3 H4 H5 H6 H7 H8
O11.42111.36582.03712.09182.09192.05712.1153
C21.42112.32601.10111.10691.10853.29852.5231
C31.36582.32603.24522.63922.62271.09451.1009
H42.03711.10113.24521.80281.80204.08653.5755
H52.09181.10692.63921.80281.80293.66602.7450
H62.09191.10852.62271.80201.80293.61742.3836
H72.05713.29851.09454.08653.66603.61741.9009
H82.11532.52311.10093.57552.74502.38361.9009

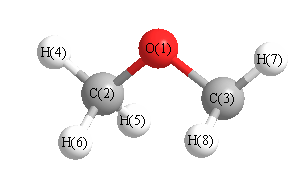
picture of methoxymethyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 H4 107.061 O1 C2 H5 111.069
O1 C2 H6 110.980 O1 C3 H7 112.998
O1 C3 H8 117.685 C2 O1 C3 113.143
H4 C2 H5 109.468 H4 C2 H6 109.282
H5 C2 H6 108.944 H7 C3 H8 119.957
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability