return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH2 (methylallyl radical)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 2A"
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-156.097955
Energy at 298.15K-156.104223
HF Energy-155.519260
Nuclear repulsion energy108.119681
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3277 3144 11.52      
2 A' 3174 3045 22.04      
3 A' 3173 3044 18.59      
4 A' 3163 3035 4.83      
5 A' 3138 3011 13.63      
6 A' 3038 2915 33.78      
7 A' 1530 1468 2.98      
8 A' 1517 1455 3.85      
9 A' 1477 1417 9.43      
10 A' 1418 1360 1.35      
11 A' 1339 1284 0.74      
12 A' 1285 1233 2.14      
13 A' 1193 1144 0.35      
14 A' 1149 1103 0.22      
15 A' 988 948 2.53      
16 A' 889 853 3.61      
17 A' 502 482 0.54      
18 A' 285 273 0.15      
19 A" 3100 2974 23.37      
20 A" 1474 1415 4.74      
21 A" 1026 985 0.62      
22 A" 985 945 23.09      
23 A" 754 724 38.60      
24 A" 722 692 0.00      
25 A" 538 516 3.99      
26 A" 216 207 1.43      
27 A" 140 134 0.66      

Unscaled Zero Point Vibrational Energy (zpe) 20744.2 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19902.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
1.23956 0.13135 0.12149

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.373 -1.425 0.000
C2 0.146 -0.754 0.000
C3 0.000 0.638 0.000
C4 -1.327 1.350 0.000
H5 2.319 -0.869 0.000
H6 1.424 -2.518 0.000
H7 -0.772 -1.361 0.000
H8 0.909 1.257 0.000
H9 -2.167 0.632 0.000
H10 -1.433 2.002 0.889
H11 -1.433 2.002 -0.889

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
C11.39922.47833.87171.09691.09472.14622.72164.09424.51754.5175
C21.39921.39942.56762.17672.17881.10032.15082.69573.29783.2978
C32.47831.39941.50582.76613.46262.14301.10002.16672.16882.1688
C43.87172.56761.50584.26844.74632.76702.23841.10451.10771.1077
H51.09692.17672.76614.26841.87653.12992.55124.73044.80764.8076
H61.09472.17883.46264.74631.87652.48183.81014.77645.42055.4205
H72.14621.10032.14302.76703.12992.48183.11142.43263.54073.5407
H82.72162.15081.10002.23842.55123.81013.11143.13892.61382.6138
H94.09422.69572.16671.10454.73044.77642.43263.13891.79041.7904
H104.51753.29782.16881.10774.80765.42053.54072.61381.79041.7780
H114.51753.29782.16881.10774.80765.42053.54072.61381.79041.7780

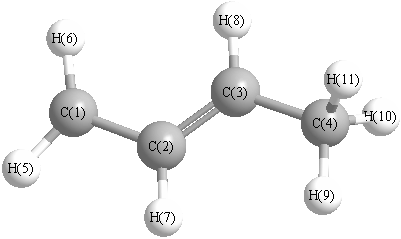
picture of methylallyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.637 C1 C2 H7 117.831
C2 C1 H5 120.912 C2 C1 H6 121.289
C2 C3 C4 124.173 C2 C3 H8 118.258
C3 C2 H7 117.532 C3 C4 H9 111.276
C3 C4 H10 111.251 C3 C4 H11 111.251
C4 C3 H8 117.569 H5 C1 H6 117.798
H9 C4 H10 108.058 H9 C4 H11 108.058
H10 C4 H11 106.755
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability