return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H5OH (Cyclopropanol)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-192.570244
Energy at 298.15K-192.577256
HF Energy-191.924473
Nuclear repulsion energy123.659831
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3839 3683 24.34      
2 A 3270 3137 12.58      
3 A 3254 3122 2.91      
4 A 3172 3043 4.66      
5 A 3159 3031 13.08      
6 A 3140 3013 36.61      
7 A 1530 1468 18.18      
8 A 1450 1391 1.49      
9 A 1430 1372 4.09      
10 A 1334 1280 102.67      
11 A 1251 1201 27.58      
12 A 1202 1153 4.94      
13 A 1197 1148 0.27      
14 A 1135 1089 0.47      
15 A 1077 1033 1.91      
16 A 1064 1021 15.06      
17 A 1004 963 8.59      
18 A 951 912 24.01      
19 A 842 808 9.82      
20 A 823 790 5.18      
21 A 762 731 2.99      
22 A 410 393 27.00      
23 A 405 388 5.26      
24 A 338 324 96.58      

Unscaled Zero Point Vibrational Energy (zpe) 19018.6 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 18246.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.55059 0.22902 0.19731

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.243 -0.015 0.485
C2 0.913 -0.747 -0.136
C3 0.887 0.779 -0.138
O4 -1.463 -0.113 -0.200
H5 -0.313 -0.018 1.583
H6 1.625 -1.260 0.519
H7 0.704 -1.246 -1.088
H8 1.580 1.317 0.518
H9 0.671 1.266 -1.095
H10 -1.902 0.742 -0.097

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.50311.51501.40291.10022.24502.21072.25752.23021.9138
C21.50311.52662.46032.23451.09491.09482.26572.24343.1848
C31.51501.52662.51442.24482.26582.24461.09551.09542.7889
O41.40292.46032.51442.12383.37162.60133.43802.69400.9666
H51.10022.23452.24482.12382.53603.11082.54963.12922.4335
H62.24501.09492.26583.37162.53601.85242.57713.14604.1016
H72.21071.09482.24462.60133.11081.85243.14922.51293.4239
H82.25752.26571.09553.43802.54962.57713.14921.85263.5817
H92.23022.24341.09542.69403.12923.14602.51291.85262.8087
H101.91383.18482.78890.96662.43354.10163.42393.58172.8087

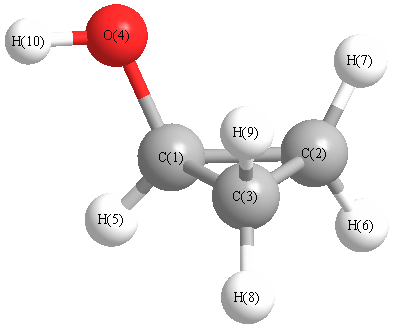
picture of Cyclopropanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.998 C1 C2 H6 118.722
C1 C2 H7 115.736 C1 C3 C2 59.232
C1 C3 H8 118.840 C1 C3 H9 116.458
C1 O4 H10 106.281 C2 C1 C3 60.770
C2 C1 O4 115.650 C2 C1 H5 117.418
C2 C3 H8 118.627 C2 C3 H9 116.689
C3 C1 O4 118.969 C3 C1 H5 117.388
C3 C2 H6 118.688 C3 C2 H7 116.838
O4 C1 H5 115.559 H6 C2 H7 115.554
H8 C3 H9 115.460
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability