Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -595.495171 |
| Energy at 298.15K | -595.485309 |
| Nuclear repulsion energy | 315.079173 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3056 |
3038 |
39.41 |
|
|
|
| 2 |
A |
3052 |
3035 |
6.15 |
|
|
|
| 3 |
A |
3049 |
3031 |
52.29 |
|
|
|
| 4 |
A |
3046 |
3029 |
38.44 |
|
|
|
| 5 |
A |
3045 |
3027 |
0.37 |
|
|
|
| 6 |
A |
3043 |
3025 |
7.87 |
|
|
|
| 7 |
A |
3032 |
3014 |
5.56 |
|
|
|
| 8 |
A |
2975 |
2958 |
22.26 |
|
|
|
| 9 |
A |
2968 |
2951 |
16.66 |
|
|
|
| 10 |
A |
2964 |
2947 |
30.83 |
|
|
|
| 11 |
A |
2960 |
2943 |
22.61 |
|
|
|
| 12 |
A |
2615 |
2600 |
7.22 |
|
|
|
| 13 |
A |
1440 |
1432 |
10.02 |
|
|
|
| 14 |
A |
1429 |
1421 |
9.54 |
|
|
|
| 15 |
A |
1426 |
1418 |
4.54 |
|
|
|
| 16 |
A |
1404 |
1396 |
0.16 |
|
|
|
| 17 |
A |
1403 |
1395 |
0.03 |
|
|
|
| 18 |
A |
1396 |
1388 |
0.27 |
|
|
|
| 19 |
A |
1396 |
1387 |
2.44 |
|
|
|
| 20 |
A |
1358 |
1350 |
7.42 |
|
|
|
| 21 |
A |
1334 |
1326 |
8.50 |
|
|
|
| 22 |
A |
1329 |
1321 |
6.76 |
|
|
|
| 23 |
A |
1250 |
1243 |
1.09 |
|
|
|
| 24 |
A |
1243 |
1236 |
8.02 |
|
|
|
| 25 |
A |
1225 |
1218 |
11.49 |
|
|
|
| 26 |
A |
1168 |
1161 |
19.16 |
|
|
|
| 27 |
A |
1095 |
1089 |
2.87 |
|
|
|
| 28 |
A |
1005 |
1000 |
0.05 |
|
|
|
| 29 |
A |
999 |
993 |
0.69 |
|
|
|
| 30 |
A |
927 |
922 |
1.79 |
|
|
|
| 31 |
A |
918 |
912 |
0.11 |
|
|
|
| 32 |
A |
913 |
908 |
0.42 |
|
|
|
| 33 |
A |
911 |
905 |
0.76 |
|
|
|
| 34 |
A |
848 |
843 |
3.61 |
|
|
|
| 35 |
A |
845 |
840 |
4.09 |
|
|
|
| 36 |
A |
752 |
748 |
4.30 |
|
|
|
| 37 |
A |
683 |
679 |
0.23 |
|
|
|
| 38 |
A |
439 |
437 |
1.44 |
|
|
|
| 39 |
A |
401 |
399 |
0.00 |
|
|
|
| 40 |
A |
380 |
378 |
0.14 |
|
|
|
| 41 |
A |
325 |
323 |
0.06 |
|
|
|
| 42 |
A |
306 |
305 |
0.30 |
|
|
|
| 43 |
A |
264 |
263 |
0.04 |
|
|
|
| 44 |
A |
255 |
254 |
0.09 |
|
|
|
| 45 |
A |
208 |
207 |
0.07 |
|
|
|
| 46 |
A |
192 |
191 |
1.44 |
|
|
|
| 47 |
A |
147 |
147 |
6.92 |
|
|
|
| 48 |
A |
51 |
51 |
9.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33735.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33540.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.800 |
-0.005 |
0.001 |
| C2 |
2.024 |
0.933 |
-0.050 |
| H3 |
2.965 |
0.346 |
-0.022 |
| S4 |
-2.039 |
-0.077 |
-0.001 |
| H5 |
-2.860 |
1.015 |
0.012 |
| C6 |
-0.457 |
0.896 |
-0.034 |
| C7 |
0.844 |
-0.828 |
1.302 |
| C8 |
0.829 |
-0.946 |
-1.220 |
| H9 |
-0.449 |
1.577 |
0.842 |
| H10 |
-0.446 |
1.513 |
-0.957 |
| H11 |
1.765 |
-1.446 |
1.340 |
| H12 |
1.757 |
-1.554 |
-1.220 |
| H13 |
-0.027 |
-1.508 |
1.386 |
| H14 |
-0.033 |
-1.642 |
-1.222 |
| H15 |
0.841 |
-0.168 |
2.195 |
| H16 |
0.799 |
-0.372 |
-2.169 |
| H17 |
2.031 |
1.539 |
-0.979 |
| H18 |
2.035 |
1.630 |
0.813 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
S4 |
H5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5432 | 2.1939 | 2.8392 | 3.7991 | 1.5470 | 1.5408 | 1.5411 | 2.1839 | 2.1846 | 2.1915 | 2.1920 | 2.2048 | 2.2064 | 2.2002 | 2.2006 | 2.2053 | 2.2045 |
C2 | 1.5432 | | 1.1094 | 4.1865 | 4.8849 | 2.4816 | 2.5143 | 2.5147 | 2.7066 | 2.6947 | 2.7676 | 2.7608 | 3.4967 | 3.4979 | 2.7656 | 2.7736 | 1.1098 | 1.1098 | H3 | 2.1939 | 1.1094 | | 5.0217 | 5.8634 | 3.4664 | 2.7624 | 2.7684 | 3.7306 | 3.7247 | 2.5512 | 2.5499 | 3.7914 | 3.7923 | 3.1126 | 3.1333 | 1.7925 | 1.7922 | S4 | 2.8392 | 4.1865 | 5.0217 | | 1.3664 | 1.8570 | 3.2519 | 3.2353 | 2.4438 | 2.4442 | 4.2592 | 4.2517 | 2.8314 | 2.8221 | 3.6224 | 3.5833 | 4.4871 | 4.4911 | H5 | 3.7991 | 4.8849 | 5.8634 | 1.3664 | | 2.4059 | 4.3343 | 4.3557 | 2.6110 | 2.6475 | 5.4048 | 5.4253 | 4.0346 | 4.0712 | 4.4568 | 4.4794 | 5.0180 | 4.9979 | C6 | 1.5470 | 2.4816 | 3.4664 | 1.8570 | 2.4059 | | 2.5405 | 2.5404 | 1.1094 | 1.1096 | 3.5090 | 3.5089 | 2.8250 | 2.8345 | 2.7902 | 2.7827 | 2.7387 | 2.7327 | C7 | 1.5408 | 2.5143 | 2.7624 | 3.2519 | 4.3343 | 2.5405 | | 2.5250 | 2.7694 | 3.4998 | 1.1094 | 2.7786 | 1.1085 | 2.7936 | 1.1098 | 3.5018 | 3.4959 | 2.7748 | C8 | 1.5411 | 2.5147 | 2.7684 | 3.2353 | 4.3557 | 2.5404 | 2.5250 | | 3.4997 | 2.7821 | 2.7710 | 1.1094 | 2.8002 | 1.1087 | 3.5018 | 1.1099 | 2.7710 | 3.4958 | H9 | 2.1839 | 2.7066 | 3.7306 | 2.4438 | 2.6110 | 1.1094 | 2.7694 | 3.4997 | | 1.7996 | 3.7801 | 4.3496 | 3.1607 | 3.8464 | 2.5572 | 3.7973 | 3.0772 | 2.4845 | H10 | 2.1846 | 2.6947 | 3.7247 | 2.4442 | 2.6475 | 1.1096 | 3.4998 | 2.7821 | 1.7996 | | 4.3497 | 3.7853 | 3.8453 | 3.1928 | 3.7963 | 2.5634 | 2.4781 | 3.0502 | H11 | 2.1915 | 2.7676 | 2.5512 | 4.2592 | 5.4048 | 3.5090 | 1.1094 | 2.7710 | 3.7801 | 4.3497 | | 2.5619 | 1.7939 | 3.1359 | 1.7937 | 3.7951 | 3.7902 | 3.1327 | H12 | 2.1920 | 2.7608 | 2.5499 | 4.2517 | 5.4253 | 3.5089 | 2.7786 | 1.1094 | 4.3496 | 3.7853 | 2.5619 | | 3.1586 | 1.7926 | 3.7968 | 1.7940 | 3.1148 | 3.7877 | H13 | 2.2048 | 3.4967 | 3.7914 | 2.8314 | 4.0346 | 2.8250 | 1.1085 | 2.8002 | 3.1607 | 3.8453 | 1.7939 | 3.1586 | | 2.6113 | 1.7898 | 3.8228 | 4.3726 | 3.7982 | H14 | 2.2064 | 3.4979 | 3.7923 | 2.8221 | 4.0712 | 2.8345 | 2.7936 | 1.1087 | 3.8464 | 3.1928 | 3.1359 | 1.7926 | 2.6113 | | 3.8222 | 1.7901 | 3.8006 | 4.3733 | H15 | 2.2002 | 2.7656 | 3.1126 | 3.6224 | 4.4568 | 2.7902 | 1.1098 | 3.5018 | 2.5572 | 3.7963 | 1.7937 | 3.7968 | 1.7898 | 3.8222 | | 4.3686 | 3.7954 | 2.5622 | H16 | 2.2006 | 2.7736 | 3.1333 | 3.5833 | 4.4794 | 2.7827 | 3.5018 | 1.1099 | 3.7973 | 2.5634 | 3.7951 | 1.7940 | 3.8228 | 1.7901 | 4.3686 | | 2.5665 | 3.7986 | H17 | 2.2053 | 1.1098 | 1.7925 | 4.4871 | 5.0180 | 2.7387 | 3.4959 | 2.7710 | 3.0772 | 2.4781 | 3.7902 | 3.1148 | 4.3726 | 3.8006 | 3.7954 | 2.5665 | | 1.7949 | H18 | 2.2045 | 1.1098 | 1.7922 | 4.4911 | 4.9979 | 2.7327 | 2.7748 | 3.4958 | 2.4845 | 3.0502 | 3.1327 | 3.7877 | 3.7982 | 4.3733 | 2.5622 | 3.7986 | 1.7949 | |
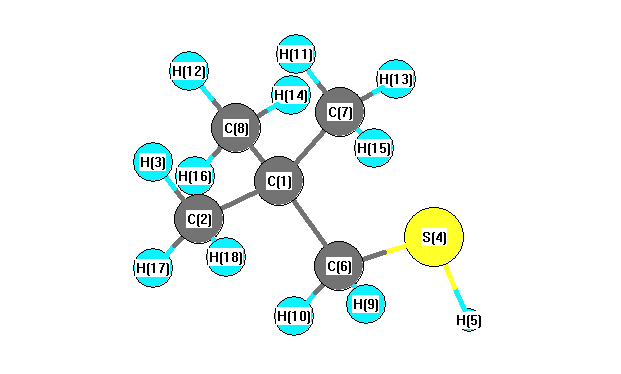 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H3 |
110.533 |
|
C1 |
C2 |
H17 |
111.412 |
| C1 |
C2 |
H18 |
111.347 |
|
C1 |
C6 |
S4 |
112.723 |
| C1 |
C6 |
H9 |
109.489 |
|
C1 |
C6 |
H10 |
109.539 |
| C1 |
C7 |
H11 |
110.511 |
|
C1 |
C7 |
H13 |
111.621 |
| C1 |
C7 |
H15 |
111.174 |
|
C1 |
C8 |
H12 |
110.521 |
| C1 |
C8 |
H14 |
111.702 |
|
C1 |
C8 |
H16 |
111.174 |
| C2 |
C1 |
C6 |
106.846 |
|
C2 |
C1 |
C7 |
109.234 |
| C2 |
C1 |
C8 |
109.240 |
|
H3 |
C2 |
H17 |
107.740 |
| H3 |
C2 |
H18 |
107.717 |
|
S4 |
C6 |
H9 |
108.289 |
| S4 |
C6 |
H10 |
108.314 |
|
H5 |
S4 |
C6 |
95.352 |
| C6 |
C1 |
C7 |
110.720 |
|
C6 |
C1 |
C8 |
110.698 |
| C7 |
C1 |
C8 |
110.031 |
|
H9 |
C6 |
H10 |
108.385 |
| H11 |
C7 |
H13 |
107.961 |
|
H11 |
C7 |
H15 |
107.847 |
| H12 |
C8 |
H14 |
107.830 |
|
H12 |
C8 |
H16 |
107.869 |
| H13 |
C7 |
H15 |
107.570 |
|
H14 |
C8 |
H16 |
107.582 |
| H17 |
C2 |
H18 |
107.931 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.342 |
|
|
|
| 2 |
C |
0.028 |
|
|
|
| 3 |
H |
0.026 |
|
|
|
| 4 |
S |
-0.117 |
|
|
|
| 5 |
H |
0.086 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
C |
0.035 |
|
|
|
| 8 |
C |
0.033 |
|
|
|
| 9 |
H |
0.051 |
|
|
|
| 10 |
H |
0.051 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
H |
0.024 |
|
|
|
| 13 |
H |
0.036 |
|
|
|
| 14 |
H |
0.036 |
|
|
|
| 15 |
H |
0.022 |
|
|
|
| 16 |
H |
0.023 |
|
|
|
| 17 |
H |
0.022 |
|
|
|
| 18 |
H |
0.022 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.871 |
1.182 |
-0.013 |
1.468 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.484 |
-2.920 |
-0.054 |
| y |
-2.920 |
-46.242 |
0.003 |
| z |
-0.054 |
0.003 |
-48.907 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.090 |
-2.920 |
-0.054 |
| y |
-2.920 |
1.954 |
0.003 |
| z |
-0.054 |
0.003 |
-2.044 |
|
| Polar |
|---|
| 3z2-r2 | -4.088 |
|---|
| x2-y2 | -1.242 |
|---|
| xy | -2.920 |
|---|
| xz | -0.054 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.167 |
-0.033 |
-0.025 |
| y |
-0.033 |
10.657 |
-0.006 |
| z |
-0.025 |
-0.006 |
9.797 |
<r2> (average value of r
2) Å
2
| <r2> |
246.260 |
| (<r2>)1/2 |
15.693 |
