Jump to
S2C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -2612.257609 |
| Energy at 298.15K | -2612.260322 |
| HF Energy | -2612.257609 |
| Nuclear repulsion energy | 69.596072 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.027 |
1.567 |
0.000 |
| Br2 |
0.027 |
-0.319 |
0.000 |
| H3 |
-1.098 |
1.749 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
| C1 | | 1.8855 | 1.1390 |
Br2 | 1.8855 | | 2.3531 | H3 | 1.1390 | 2.3531 | |
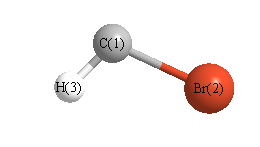 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
99.174 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.174 |
|
|
|
| 2 |
Br |
0.074 |
|
|
|
| 3 |
H |
0.100 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.319 |
-0.889 |
0.000 |
1.590 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.155 |
-2.797 |
0.000 |
| y |
-2.797 |
-25.630 |
0.000 |
| z |
0.000 |
0.000 |
-23.512 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.583 |
-2.797 |
0.000 |
| y |
-2.797 |
-1.296 |
0.000 |
| z |
0.000 |
0.000 |
1.880 |
|
| Polar |
|---|
| 3z2-r2 | 3.760 |
|---|
| x2-y2 | 0.475 |
|---|
| xy | -2.797 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.012 |
-0.349 |
0.000 |
| y |
-0.349 |
6.196 |
0.000 |
| z |
0.000 |
0.000 |
2.313 |
<r2> (average value of r
2) Å
2
| <r2> |
38.044 |
| (<r2>)1/2 |
6.168 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -2612.253522 |
| Energy at 298.15K | -2612.256219 |
| HF Energy | -2612.253522 |
| Nuclear repulsion energy | 70.726521 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.021 |
1.507 |
0.000 |
| Br2 |
0.021 |
-0.320 |
0.000 |
| H3 |
-0.871 |
2.158 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
| C1 | | 1.8271 | 1.1047 |
Br2 | 1.8271 | | 2.6338 | H3 | 1.1047 | 2.6338 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.181 |
|
|
|
| 2 |
Br |
0.078 |
|
|
|
| 3 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.749 |
-0.086 |
0.000 |
0.754 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.028 |
-1.716 |
0.000 |
| y |
-1.716 |
-22.201 |
0.000 |
| z |
0.000 |
0.000 |
-25.218 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.318 |
-1.716 |
0.000 |
| y |
-1.716 |
2.422 |
0.000 |
| z |
0.000 |
0.000 |
-2.104 |
|
| Polar |
|---|
| 3z2-r2 | -4.207 |
|---|
| x2-y2 | -1.827 |
|---|
| xy | -1.716 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.500 |
-0.234 |
0.000 |
| y |
-0.234 |
5.832 |
0.000 |
| z |
0.000 |
0.000 |
2.087 |
<r2> (average value of r
2) Å
2
| <r2> |
37.522 |
| (<r2>)1/2 |
6.126 |
