Jump to
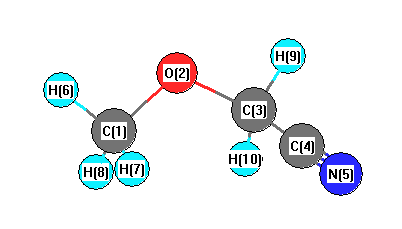
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -247.185591 |
| Energy at 298.15K | -247.191028 |
| HF Energy | -247.185591 |
| Nuclear repulsion energy | 155.179179 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3052 |
3057 |
17.38 |
|
|
|
| 2 |
A' |
2885 |
2889 |
61.91 |
|
|
|
| 3 |
A' |
2862 |
2867 |
31.05 |
|
|
|
| 4 |
A' |
2268 |
2272 |
0.28 |
|
|
|
| 5 |
A' |
1445 |
1448 |
4.19 |
|
|
|
| 6 |
A' |
1421 |
1424 |
6.26 |
|
|
|
| 7 |
A' |
1409 |
1411 |
2.55 |
|
|
|
| 8 |
A' |
1346 |
1348 |
43.45 |
|
|
|
| 9 |
A' |
1162 |
1164 |
20.07 |
|
|
|
| 10 |
A' |
1086 |
1087 |
157.24 |
|
|
|
| 11 |
A' |
936 |
938 |
26.30 |
|
|
|
| 12 |
A' |
919 |
921 |
9.48 |
|
|
|
| 13 |
A' |
524 |
525 |
1.09 |
|
|
|
| 14 |
A' |
365 |
366 |
1.30 |
|
|
|
| 15 |
A' |
173 |
173 |
1.67 |
|
|
|
| 16 |
A" |
2929 |
2934 |
60.52 |
|
|
|
| 17 |
A" |
2881 |
2886 |
44.92 |
|
|
|
| 18 |
A" |
1410 |
1412 |
6.08 |
|
|
|
| 19 |
A" |
1195 |
1197 |
3.07 |
|
|
|
| 20 |
A" |
1123 |
1125 |
3.24 |
|
|
|
| 21 |
A" |
985 |
986 |
1.65 |
|
|
|
| 22 |
A" |
361 |
361 |
0.67 |
|
|
|
| 23 |
A" |
229 |
229 |
3.45 |
|
|
|
| 24 |
A" |
85 |
85 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16525.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 16551.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.150 |
-0.381 |
0.000 |
| O2 |
-0.738 |
-0.624 |
0.000 |
| C3 |
0.000 |
0.596 |
0.000 |
| C4 |
1.444 |
0.289 |
0.000 |
| N5 |
2.598 |
0.075 |
0.000 |
| H6 |
-2.638 |
-1.373 |
0.000 |
| H7 |
-2.474 |
0.186 |
0.905 |
| H8 |
-2.474 |
0.186 |
-0.905 |
| H9 |
-0.231 |
1.221 |
-0.899 |
| H10 |
-0.231 |
1.221 |
0.899 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4324 | 2.3615 | 3.6563 | 4.7693 | 1.1055 | 1.1165 | 1.1165 | 2.6570 | 2.6570 |
O2 | 1.4324 | | 1.4258 | 2.3660 | 3.4082 | 2.0423 | 2.1188 | 2.1188 | 2.1144 | 2.1144 | C3 | 2.3615 | 1.4258 | | 1.4766 | 2.6494 | 3.2918 | 2.6660 | 2.6660 | 1.1191 | 1.1191 | C4 | 3.6563 | 2.3660 | 1.4766 | | 1.1730 | 4.4080 | 4.0230 | 4.0230 | 2.1172 | 2.1172 | N5 | 4.7693 | 3.4082 | 2.6494 | 1.1730 | | 5.4322 | 5.1530 | 5.1530 | 3.1814 | 3.1814 | H6 | 1.1055 | 2.0423 | 3.2918 | 4.4080 | 5.4322 | | 1.8106 | 1.8106 | 3.6517 | 3.6517 | H7 | 1.1165 | 2.1188 | 2.6660 | 4.0230 | 5.1530 | 1.8106 | | 1.8102 | 3.0592 | 2.4707 | H8 | 1.1165 | 2.1188 | 2.6660 | 4.0230 | 5.1530 | 1.8106 | 1.8102 | | 2.4707 | 3.0592 | H9 | 2.6570 | 2.1144 | 1.1191 | 2.1172 | 3.1814 | 3.6517 | 3.0592 | 2.4707 | | 1.7978 | H10 | 2.6570 | 2.1144 | 1.1191 | 2.1172 | 3.1814 | 3.6517 | 2.4707 | 3.0592 | 1.7978 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.423 |
|
O2 |
C1 |
H6 |
106.453 |
| O2 |
C1 |
H7 |
111.859 |
|
O2 |
C1 |
H8 |
111.859 |
| O2 |
C3 |
C4 |
109.194 |
|
O2 |
C3 |
H9 |
111.808 |
| O2 |
C3 |
H10 |
111.808 |
|
C3 |
C4 |
N5 |
178.553 |
| C4 |
C3 |
H9 |
108.517 |
|
C4 |
C3 |
H10 |
108.517 |
| H6 |
C1 |
H7 |
109.144 |
|
H6 |
C1 |
H8 |
109.144 |
| H7 |
C1 |
H8 |
108.321 |
|
H9 |
C3 |
H10 |
106.883 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.140 |
|
|
|
| 2 |
O |
-0.269 |
|
|
|
| 3 |
C |
0.209 |
|
|
|
| 4 |
C |
-0.129 |
|
|
|
| 5 |
N |
-0.069 |
|
|
|
| 6 |
H |
0.030 |
|
|
|
| 7 |
H |
0.007 |
|
|
|
| 8 |
H |
0.007 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.010 |
1.499 |
0.000 |
4.281 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.539 |
0.650 |
0.000 |
| y |
0.650 |
-29.142 |
0.000 |
| z |
0.000 |
0.000 |
-29.365 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.285 |
0.650 |
0.000 |
| y |
0.650 |
3.310 |
0.000 |
| z |
0.000 |
0.000 |
2.975 |
|
| Polar |
|---|
| 3z2-r2 | 5.951 |
|---|
| x2-y2 | -6.397 |
|---|
| xy | 0.650 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.717 |
0.112 |
0.000 |
| y |
0.112 |
5.023 |
0.000 |
| z |
0.000 |
0.000 |
4.681 |
<r2> (average value of r
2) Å
2
| <r2> |
145.581 |
| (<r2>)1/2 |
12.066 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -247.188189 |
| Energy at 298.15K | -247.193694 |
| HF Energy | -247.188189 |
| Nuclear repulsion energy | 157.741532 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3055 |
3060 |
17.61 |
|
|
|
| 2 |
A |
3007 |
3012 |
6.39 |
|
|
|
| 3 |
A |
2959 |
2964 |
40.27 |
|
|
|
| 4 |
A |
2898 |
2903 |
53.23 |
|
|
|
| 5 |
A |
2884 |
2888 |
35.27 |
|
|
|
| 6 |
A |
2248 |
2252 |
0.10 |
|
|
|
| 7 |
A |
1437 |
1439 |
9.95 |
|
|
|
| 8 |
A |
1415 |
1417 |
0.69 |
|
|
|
| 9 |
A |
1411 |
1413 |
6.03 |
|
|
|
| 10 |
A |
1393 |
1395 |
3.05 |
|
|
|
| 11 |
A |
1328 |
1330 |
17.43 |
|
|
|
| 12 |
A |
1252 |
1254 |
7.03 |
|
|
|
| 13 |
A |
1158 |
1160 |
12.67 |
|
|
|
| 14 |
A |
1123 |
1125 |
3.36 |
|
|
|
| 15 |
A |
1077 |
1079 |
129.90 |
|
|
|
| 16 |
A |
984 |
985 |
7.49 |
|
|
|
| 17 |
A |
889 |
891 |
30.33 |
|
|
|
| 18 |
A |
863 |
864 |
15.52 |
|
|
|
| 19 |
A |
572 |
573 |
1.77 |
|
|
|
| 20 |
A |
375 |
376 |
2.55 |
|
|
|
| 21 |
A |
347 |
347 |
0.69 |
|
|
|
| 22 |
A |
243 |
244 |
7.34 |
|
|
|
| 23 |
A |
169 |
169 |
2.90 |
|
|
|
| 24 |
A |
113 |
113 |
6.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16599.9 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 16626.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.609 |
-0.764 |
0.140 |
| O2 |
1.083 |
0.436 |
-0.453 |
| C3 |
-0.073 |
0.929 |
0.214 |
| C4 |
-1.269 |
0.057 |
0.044 |
| N5 |
-2.193 |
-0.653 |
-0.101 |
| H6 |
2.507 |
-1.030 |
-0.447 |
| H7 |
0.879 |
-1.604 |
0.094 |
| H8 |
1.899 |
-0.599 |
1.205 |
| H9 |
-0.293 |
1.921 |
-0.227 |
| H10 |
0.100 |
1.068 |
1.310 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4383 | 2.3877 | 2.9945 | 3.8107 | 1.1056 | 1.1134 | 1.1158 | 3.3109 | 2.6462 |
O2 | 1.4383 | | 1.4228 | 2.4339 | 3.4699 | 2.0440 | 2.1219 | 2.1179 | 2.0368 | 2.1146 | C3 | 2.3877 | 1.4228 | | 1.4897 | 2.6635 | 3.3064 | 2.7086 | 2.6846 | 1.1082 | 1.1176 | C4 | 2.9945 | 2.4339 | 1.4897 | | 1.1740 | 3.9605 | 2.7162 | 3.4379 | 2.1219 | 2.1210 | N5 | 3.8107 | 3.4699 | 2.6635 | 1.1740 | | 4.7277 | 3.2214 | 4.2958 | 3.2023 | 3.1952 | H6 | 1.1056 | 2.0440 | 3.3064 | 3.9605 | 4.7277 | | 1.8092 | 1.8115 | 4.0738 | 3.6444 | H7 | 1.1134 | 2.1219 | 2.7086 | 2.7162 | 3.2214 | 1.8092 | | 1.8124 | 3.7287 | 3.0374 | H8 | 1.1158 | 2.1179 | 2.6846 | 3.4379 | 4.2958 | 1.8115 | 1.8124 | | 3.6343 | 2.4556 | H9 | 3.3109 | 2.0368 | 1.1082 | 2.1219 | 3.2023 | 4.0738 | 3.7287 | 3.6343 | | 1.8009 | H10 | 2.6462 | 2.1146 | 1.1176 | 2.1210 | 3.1952 | 3.6444 | 3.0374 | 2.4556 | 1.8009 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.127 |
|
O2 |
C1 |
H6 |
106.190 |
| O2 |
C1 |
H7 |
111.888 |
|
O2 |
C1 |
H8 |
111.408 |
| O2 |
C3 |
C4 |
113.348 |
|
O2 |
C3 |
H9 |
106.505 |
| O2 |
C3 |
H10 |
112.128 |
|
C3 |
C4 |
N5 |
178.422 |
| C4 |
C3 |
H9 |
108.639 |
|
C4 |
C3 |
H10 |
108.028 |
| H6 |
C1 |
H7 |
109.237 |
|
H6 |
C1 |
H8 |
109.268 |
| H7 |
C1 |
H8 |
108.784 |
|
H9 |
C3 |
H10 |
108.017 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.119 |
|
|
|
| 2 |
O |
-0.261 |
|
|
|
| 3 |
C |
0.208 |
|
|
|
| 4 |
C |
-0.137 |
|
|
|
| 5 |
N |
-0.078 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
H |
0.007 |
|
|
|
| 9 |
H |
0.051 |
|
|
|
| 10 |
H |
0.034 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.444 |
1.176 |
1.238 |
2.981 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.593 |
-5.092 |
0.443 |
| y |
-5.092 |
-28.114 |
0.480 |
| z |
0.443 |
0.480 |
-29.733 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.669 |
-5.092 |
0.443 |
| y |
-5.092 |
4.548 |
0.480 |
| z |
0.443 |
0.480 |
2.121 |
|
| Polar |
|---|
| 3z2-r2 | 4.242 |
|---|
| x2-y2 | -7.478 |
|---|
| xy | -5.092 |
|---|
| xz | 0.443 |
|---|
| yz | 0.480 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.357 |
0.324 |
0.197 |
| y |
0.324 |
5.905 |
0.219 |
| z |
0.197 |
0.219 |
4.693 |
<r2> (average value of r
2) Å
2
| <r2> |
126.057 |
| (<r2>)1/2 |
11.228 |
