Jump to
S1C2
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -246.629663 |
| Energy at 298.15K | -246.635182 |
| HF Energy | -245.805891 |
| Nuclear repulsion energy | 156.007560 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3168 |
3038 |
|
|
|
|
| 2 |
A' |
3004 |
2881 |
|
|
|
|
| 3 |
A' |
2997 |
2874 |
|
|
|
|
| 4 |
A' |
2286 |
2192 |
|
|
|
|
| 5 |
A' |
1512 |
1450 |
|
|
|
|
| 6 |
A' |
1492 |
1430 |
|
|
|
|
| 7 |
A' |
1470 |
1410 |
|
|
|
|
| 8 |
A' |
1407 |
1350 |
|
|
|
|
| 9 |
A' |
1229 |
1179 |
|
|
|
|
| 10 |
A' |
1171 |
1123 |
|
|
|
|
| 11 |
A' |
1011 |
970 |
|
|
|
|
| 12 |
A' |
949 |
910 |
|
|
|
|
| 13 |
A' |
533 |
511 |
|
|
|
|
| 14 |
A' |
378 |
363 |
|
|
|
|
| 15 |
A' |
173 |
166 |
|
|
|
|
| 16 |
A" |
3065 |
2940 |
|
|
|
|
| 17 |
A" |
3040 |
2915 |
|
|
|
|
| 18 |
A" |
1480 |
1419 |
|
|
|
|
| 19 |
A" |
1255 |
1203 |
|
|
|
|
| 20 |
A" |
1178 |
1129 |
|
|
|
|
| 21 |
A" |
1031 |
989 |
|
|
|
|
| 22 |
A" |
352 |
338 |
|
|
|
|
| 23 |
A" |
229 |
220 |
|
|
|
|
| 24 |
A" |
84 |
81 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17246.6 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 16539.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.140 |
-0.312 |
0.000 |
| O2 |
-0.750 |
-0.599 |
0.000 |
| C3 |
0.000 |
0.594 |
0.000 |
| C4 |
1.442 |
0.244 |
0.000 |
| N5 |
2.592 |
-0.006 |
0.000 |
| H6 |
-2.667 |
-1.279 |
0.000 |
| H7 |
-2.437 |
0.265 |
0.902 |
| H8 |
-2.437 |
0.265 |
-0.902 |
| H9 |
-0.209 |
1.216 |
-0.897 |
| H10 |
-0.209 |
1.216 |
0.897 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4198 | 2.3239 | 3.6253 | 4.7420 | 1.1011 | 1.1108 | 1.1108 | 2.6204 | 2.6204 |
O2 | 1.4198 | | 1.4092 | 2.3486 | 3.3939 | 2.0338 | 2.0991 | 2.0991 | 2.0956 | 2.0956 | C3 | 2.3239 | 1.4092 | | 1.4842 | 2.6604 | 3.2586 | 2.6191 | 2.6191 | 1.1116 | 1.1116 | C4 | 3.6253 | 2.3486 | 1.4842 | | 1.1764 | 4.3821 | 3.9826 | 3.9826 | 2.1160 | 2.1160 | N5 | 4.7420 | 3.3939 | 2.6604 | 1.1764 | | 5.4105 | 5.1161 | 5.1161 | 3.1850 | 3.1850 | H6 | 1.1011 | 2.0338 | 3.2586 | 4.3821 | 5.4105 | | 1.8026 | 1.8026 | 3.6148 | 3.6148 | H7 | 1.1108 | 2.0991 | 2.6191 | 3.9826 | 5.1161 | 1.8026 | | 1.8034 | 3.0170 | 2.4220 | H8 | 1.1108 | 2.0991 | 2.6191 | 3.9826 | 5.1161 | 1.8026 | 1.8034 | | 2.4220 | 3.0170 | H9 | 2.6204 | 2.0956 | 1.1116 | 2.1160 | 3.1850 | 3.6148 | 3.0170 | 2.4220 | | 1.7946 | H10 | 2.6204 | 2.0956 | 1.1116 | 2.1160 | 3.1850 | 3.6148 | 2.4220 | 3.0170 | 1.7946 | |
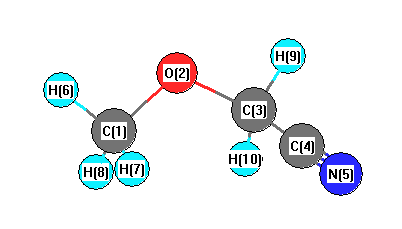 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.461 |
|
O2 |
C1 |
H6 |
106.884 |
| O2 |
C1 |
H7 |
111.512 |
|
O2 |
C1 |
H8 |
111.512 |
| O2 |
C3 |
C4 |
108.500 |
|
O2 |
C3 |
H9 |
111.930 |
| O2 |
C3 |
H10 |
111.930 |
|
C3 |
C4 |
N5 |
178.616 |
| C4 |
C3 |
H9 |
108.356 |
|
C4 |
C3 |
H10 |
108.356 |
| H6 |
C1 |
H7 |
109.169 |
|
H6 |
C1 |
H8 |
109.169 |
| H7 |
C1 |
H8 |
108.544 |
|
H9 |
C3 |
H10 |
107.654 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -246.632437 |
| Energy at 298.15K | -246.638072 |
| HF Energy | -245.807724 |
| Nuclear repulsion energy | 158.843737 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3040 |
|
|
|
|
| 2 |
A |
3139 |
3010 |
|
|
|
|
| 3 |
A |
3086 |
2959 |
|
|
|
|
| 4 |
A |
3024 |
2900 |
|
|
|
|
| 5 |
A |
3012 |
2888 |
|
|
|
|
| 6 |
A |
2268 |
2175 |
|
|
|
|
| 7 |
A |
1503 |
1442 |
|
|
|
|
| 8 |
A |
1483 |
1422 |
|
|
|
|
| 9 |
A |
1477 |
1417 |
|
|
|
|
| 10 |
A |
1464 |
1404 |
|
|
|
|
| 11 |
A |
1387 |
1330 |
|
|
|
|
| 12 |
A |
1311 |
1257 |
|
|
|
|
| 13 |
A |
1221 |
1171 |
|
|
|
|
| 14 |
A |
1180 |
1132 |
|
|
|
|
| 15 |
A |
1166 |
1118 |
|
|
|
|
| 16 |
A |
1029 |
987 |
|
|
|
|
| 17 |
A |
961 |
922 |
|
|
|
|
| 18 |
A |
895 |
858 |
|
|
|
|
| 19 |
A |
586 |
562 |
|
|
|
|
| 20 |
A |
396 |
379 |
|
|
|
|
| 21 |
A |
345 |
331 |
|
|
|
|
| 22 |
A |
255 |
244 |
|
|
|
|
| 23 |
A |
174 |
166 |
|
|
|
|
| 24 |
A |
117 |
112 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17323.4 cm
-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 16613.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.556 |
-0.782 |
0.139 |
| O2 |
1.092 |
0.430 |
-0.448 |
| C3 |
-0.041 |
0.935 |
0.214 |
| C4 |
-1.247 |
0.066 |
0.038 |
| N5 |
-2.184 |
-0.634 |
-0.099 |
| H6 |
2.464 |
-1.072 |
-0.412 |
| H7 |
0.803 |
-1.591 |
0.054 |
| H8 |
1.807 |
-0.637 |
1.211 |
| H9 |
-0.259 |
1.925 |
-0.219 |
| H10 |
0.131 |
1.057 |
1.304 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4243 | 2.3453 | 2.9301 | 3.7501 | 1.1010 | 1.1086 | 1.1104 | 3.2784 | 2.6014 |
O2 | 1.4243 | | 1.4055 | 2.4168 | 3.4621 | 2.0347 | 2.1026 | 2.0981 | 2.0279 | 2.0942 | C3 | 2.3453 | 1.4055 | | 1.4971 | 2.6743 | 3.2697 | 2.6675 | 2.6227 | 1.1022 | 1.1107 | C4 | 2.9301 | 2.4168 | 1.4971 | | 1.1773 | 3.9078 | 2.6359 | 3.3463 | 2.1204 | 2.1177 | N5 | 3.7501 | 3.4621 | 2.6743 | 1.1773 | | 4.6791 | 3.1399 | 4.2002 | 3.2038 | 3.1917 | H6 | 1.1010 | 2.0347 | 3.2697 | 3.9078 | 4.6791 | | 1.8018 | 1.8034 | 4.0538 | 3.5939 | H7 | 1.1086 | 2.1026 | 2.6675 | 2.6359 | 3.1399 | 1.8018 | | 1.8045 | 3.6827 | 3.0040 | H8 | 1.1104 | 2.0981 | 2.6227 | 3.3463 | 4.2002 | 1.8034 | 1.8045 | | 3.5884 | 2.3845 | H9 | 3.2784 | 2.0279 | 1.1022 | 2.1204 | 3.2038 | 4.0538 | 3.6827 | 3.5884 | | 1.7959 | H10 | 2.6014 | 2.0942 | 1.1107 | 2.1177 | 3.1917 | 3.5939 | 3.0040 | 2.3845 | 1.7959 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.952 |
|
O2 |
C1 |
H6 |
106.664 |
| O2 |
C1 |
H7 |
111.616 |
|
O2 |
C1 |
H8 |
111.140 |
| O2 |
C3 |
C4 |
112.705 |
|
O2 |
C3 |
H9 |
107.312 |
| O2 |
C3 |
H10 |
112.141 |
|
C3 |
C4 |
N5 |
178.991 |
| C4 |
C3 |
H9 |
108.372 |
|
C4 |
C3 |
H10 |
107.683 |
| H6 |
C1 |
H7 |
109.266 |
|
H6 |
C1 |
H8 |
109.277 |
| H7 |
C1 |
H8 |
108.826 |
|
H9 |
C3 |
H10 |
108.500 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
