Jump to
S1C2
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -246.588313 |
| Energy at 298.15K | -246.593909 |
| HF Energy | -245.809229 |
| Nuclear repulsion energy | 156.789120 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3210 |
3018 |
17.72 |
|
|
|
| 2 |
A' |
3055 |
2872 |
37.00 |
|
|
|
| 3 |
A' |
3046 |
2865 |
30.50 |
|
|
|
| 4 |
A' |
2441 |
2296 |
0.68 |
|
|
|
| 5 |
A' |
1529 |
1437 |
3.12 |
|
|
|
| 6 |
A' |
1508 |
1418 |
7.30 |
|
|
|
| 7 |
A' |
1491 |
1402 |
0.18 |
|
|
|
| 8 |
A' |
1434 |
1349 |
56.54 |
|
|
|
| 9 |
A' |
1259 |
1184 |
96.01 |
|
|
|
| 10 |
A' |
1205 |
1133 |
78.37 |
|
|
|
| 11 |
A' |
1044 |
982 |
30.76 |
|
|
|
| 12 |
A' |
974 |
915 |
8.70 |
|
|
|
| 13 |
A' |
552 |
519 |
0.92 |
|
|
|
| 14 |
A' |
388 |
364 |
1.76 |
|
|
|
| 15 |
A' |
181 |
170 |
2.12 |
|
|
|
| 16 |
A" |
3114 |
2928 |
43.57 |
|
|
|
| 17 |
A" |
3094 |
2909 |
37.24 |
|
|
|
| 18 |
A" |
1496 |
1407 |
7.25 |
|
|
|
| 19 |
A" |
1279 |
1203 |
3.89 |
|
|
|
| 20 |
A" |
1195 |
1124 |
4.92 |
|
|
|
| 21 |
A" |
1046 |
984 |
2.91 |
|
|
|
| 22 |
A" |
369 |
347 |
1.43 |
|
|
|
| 23 |
A" |
227 |
213 |
4.17 |
|
|
|
| 24 |
A" |
86 |
81 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17609.8 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 16558.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.131 |
-0.322 |
0.000 |
| O2 |
-0.746 |
-0.599 |
0.000 |
| C3 |
0.000 |
0.588 |
0.000 |
| C4 |
1.440 |
0.248 |
0.000 |
| N5 |
2.578 |
0.011 |
0.000 |
| H6 |
-2.653 |
-1.288 |
0.000 |
| H7 |
-2.431 |
0.251 |
0.899 |
| H8 |
-2.431 |
0.251 |
-0.899 |
| H9 |
-0.209 |
1.208 |
-0.894 |
| H10 |
-0.209 |
1.208 |
0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4125 | 2.3175 | 3.6166 | 4.7206 | 1.0980 | 1.1072 | 1.1072 | 2.6144 | 2.6144 |
O2 | 1.4125 | | 1.4022 | 2.3445 | 3.3792 | 2.0278 | 2.0904 | 2.0904 | 2.0862 | 2.0862 | C3 | 2.3175 | 1.4022 | | 1.4802 | 2.6416 | 3.2496 | 2.6140 | 2.6140 | 1.1075 | 1.1075 | C4 | 3.6166 | 2.3445 | 1.4802 | | 1.1617 | 4.3720 | 3.9746 | 3.9746 | 2.1076 | 2.1076 | N5 | 4.7206 | 3.3792 | 2.6416 | 1.1617 | | 5.3896 | 5.0947 | 5.0947 | 3.1620 | 3.1620 | H6 | 1.0980 | 2.0278 | 3.2496 | 4.3720 | 5.3896 | | 1.7959 | 1.7959 | 3.6060 | 3.6060 | H7 | 1.1072 | 2.0904 | 2.6140 | 3.9746 | 5.0947 | 1.7959 | | 1.7972 | 3.0113 | 2.4197 | H8 | 1.1072 | 2.0904 | 2.6140 | 3.9746 | 5.0947 | 1.7959 | 1.7972 | | 2.4197 | 3.0113 | H9 | 2.6144 | 2.0862 | 1.1075 | 2.1076 | 3.1620 | 3.6060 | 3.0113 | 2.4197 | | 1.7877 | H10 | 2.6144 | 2.0862 | 1.1075 | 2.1076 | 3.1620 | 3.6060 | 2.4197 | 3.0113 | 1.7877 | |
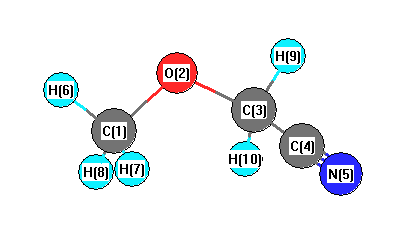 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.842 |
|
O2 |
C1 |
H6 |
107.081 |
| O2 |
C1 |
H7 |
111.550 |
|
O2 |
C1 |
H8 |
111.550 |
| O2 |
C3 |
C4 |
108.826 |
|
O2 |
C3 |
H9 |
111.922 |
| O2 |
C3 |
H10 |
111.922 |
|
C3 |
C4 |
N5 |
178.438 |
| C4 |
C3 |
H9 |
108.210 |
|
C4 |
C3 |
H10 |
108.210 |
| H6 |
C1 |
H7 |
109.050 |
|
H6 |
C1 |
H8 |
109.050 |
| H7 |
C1 |
H8 |
108.503 |
|
H9 |
C3 |
H10 |
107.621 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -246.590866 |
| Energy at 298.15K | -246.596585 |
| HF Energy | -245.811086 |
| Nuclear repulsion energy | 159.607077 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3212 |
3020 |
18.21 |
|
|
|
| 2 |
A |
3183 |
2993 |
6.17 |
|
|
|
| 3 |
A |
3131 |
2944 |
37.08 |
|
|
|
| 4 |
A |
3075 |
2892 |
29.41 |
|
|
|
| 5 |
A |
3057 |
2875 |
34.11 |
|
|
|
| 6 |
A |
2426 |
2281 |
1.09 |
|
|
|
| 7 |
A |
1519 |
1429 |
9.60 |
|
|
|
| 8 |
A |
1501 |
1412 |
1.98 |
|
|
|
| 9 |
A |
1497 |
1407 |
7.19 |
|
|
|
| 10 |
A |
1483 |
1394 |
0.97 |
|
|
|
| 11 |
A |
1412 |
1327 |
25.90 |
|
|
|
| 12 |
A |
1335 |
1255 |
11.65 |
|
|
|
| 13 |
A |
1253 |
1178 |
94.96 |
|
|
|
| 14 |
A |
1202 |
1131 |
41.00 |
|
|
|
| 15 |
A |
1192 |
1121 |
19.14 |
|
|
|
| 16 |
A |
1048 |
986 |
11.12 |
|
|
|
| 17 |
A |
991 |
932 |
25.56 |
|
|
|
| 18 |
A |
922 |
867 |
20.26 |
|
|
|
| 19 |
A |
606 |
570 |
1.71 |
|
|
|
| 20 |
A |
404 |
380 |
3.00 |
|
|
|
| 21 |
A |
362 |
341 |
0.85 |
|
|
|
| 22 |
A |
260 |
244 |
9.94 |
|
|
|
| 23 |
A |
178 |
167 |
3.47 |
|
|
|
| 24 |
A |
120 |
113 |
7.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17683.7 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 16628.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.561 |
-0.771 |
0.136 |
| O2 |
1.085 |
0.431 |
-0.444 |
| C3 |
-0.047 |
0.925 |
0.212 |
| C4 |
-1.246 |
0.056 |
0.035 |
| N5 |
-2.177 |
-0.629 |
-0.097 |
| H6 |
2.466 |
-1.053 |
-0.417 |
| H7 |
0.818 |
-1.585 |
0.056 |
| H8 |
1.818 |
-0.627 |
1.203 |
| H9 |
-0.272 |
1.912 |
-0.217 |
| H10 |
0.121 |
1.048 |
1.299 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4167 | 2.3384 | 2.9284 | 3.7477 | 1.0980 | 1.1054 | 1.1069 | 3.2678 | 2.5955 |
O2 | 1.4167 | | 1.3985 | 2.4095 | 3.4471 | 2.0278 | 2.0942 | 2.0900 | 2.0209 | 2.0853 | C3 | 2.3384 | 1.3985 | | 1.4920 | 2.6543 | 3.2600 | 2.6600 | 2.6209 | 1.0989 | 1.1068 | C4 | 2.9284 | 2.4095 | 1.4920 | | 1.1624 | 3.9011 | 2.6374 | 3.3499 | 2.1114 | 2.1102 | N5 | 3.7477 | 3.4471 | 2.6543 | 1.1624 | | 4.6733 | 3.1476 | 4.2009 | 3.1776 | 3.1685 | H6 | 1.0980 | 2.0278 | 3.2600 | 3.9011 | 4.6733 | | 1.7950 | 1.7967 | 4.0407 | 3.5868 | H7 | 1.1054 | 2.0942 | 2.6600 | 2.6374 | 3.1476 | 1.7950 | | 1.7989 | 3.6729 | 2.9950 | H8 | 1.1069 | 2.0900 | 2.6209 | 3.3499 | 4.2009 | 1.7967 | 1.7989 | | 3.5815 | 2.3867 | H9 | 3.2678 | 2.0209 | 1.0989 | 2.1114 | 3.1776 | 4.0407 | 3.6729 | 3.5815 | | 1.7883 | H10 | 2.5955 | 2.0853 | 1.1068 | 2.1102 | 3.1685 | 3.5868 | 2.9950 | 2.3867 | 1.7883 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.321 |
|
O2 |
C1 |
H6 |
106.804 |
| O2 |
C1 |
H7 |
111.672 |
|
O2 |
C1 |
H8 |
111.232 |
| O2 |
C3 |
C4 |
112.899 |
|
O2 |
C3 |
H9 |
107.425 |
| O2 |
C3 |
H10 |
112.162 |
|
C3 |
C4 |
N5 |
179.490 |
| C4 |
C3 |
H9 |
108.204 |
|
C4 |
C3 |
H10 |
107.660 |
| H6 |
C1 |
H7 |
109.113 |
|
H6 |
C1 |
H8 |
109.150 |
| H7 |
C1 |
H8 |
108.810 |
|
H9 |
C3 |
H10 |
108.340 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
