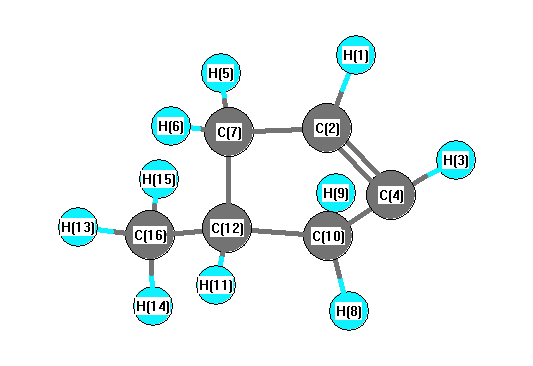
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3138 |
3120 |
28.18 |
|
|
|
| 2 |
A |
3112 |
3094 |
7.37 |
|
|
|
| 3 |
A |
3044 |
3026 |
21.66 |
|
|
|
| 4 |
A |
3038 |
3021 |
38.48 |
|
|
|
| 5 |
A |
2995 |
2978 |
29.47 |
|
|
|
| 6 |
A |
2991 |
2974 |
34.29 |
|
|
|
| 7 |
A |
2960 |
2943 |
36.09 |
|
|
|
| 8 |
A |
2954 |
2937 |
12.45 |
|
|
|
| 9 |
A |
2918 |
2901 |
41.54 |
|
|
|
| 10 |
A |
2917 |
2901 |
26.13 |
|
|
|
| 11 |
A |
1640 |
1631 |
3.86 |
|
|
|
| 12 |
A |
1422 |
1414 |
6.61 |
|
|
|
| 13 |
A |
1418 |
1410 |
2.45 |
|
|
|
| 14 |
A |
1409 |
1401 |
0.63 |
|
|
|
| 15 |
A |
1400 |
1392 |
2.02 |
|
|
|
| 16 |
A |
1344 |
1336 |
2.59 |
|
|
|
| 17 |
A |
1319 |
1312 |
0.09 |
|
|
|
| 18 |
A |
1315 |
1308 |
4.52 |
|
|
|
| 19 |
A |
1276 |
1269 |
2.25 |
|
|
|
| 20 |
A |
1251 |
1243 |
0.07 |
|
|
|
| 21 |
A |
1231 |
1223 |
2.39 |
|
|
|
| 22 |
A |
1183 |
1176 |
0.31 |
|
|
|
| 23 |
A |
1128 |
1122 |
0.06 |
|
|
|
| 24 |
A |
1100 |
1093 |
2.25 |
|
|
|
| 25 |
A |
1091 |
1084 |
0.48 |
|
|
|
| 26 |
A |
1073 |
1067 |
0.27 |
|
|
|
| 27 |
A |
1032 |
1026 |
1.56 |
|
|
|
| 28 |
A |
969 |
963 |
0.21 |
|
|
|
| 29 |
A |
961 |
955 |
11.91 |
|
|
|
| 30 |
A |
923 |
917 |
0.07 |
|
|
|
| 31 |
A |
902 |
897 |
3.84 |
|
|
|
| 32 |
A |
891 |
886 |
4.70 |
|
|
|
| 33 |
A |
870 |
865 |
0.00 |
|
|
|
| 34 |
A |
810 |
805 |
1.50 |
|
|
|
| 35 |
A |
755 |
750 |
0.51 |
|
|
|
| 36 |
A |
676 |
672 |
28.23 |
|
|
|
| 37 |
A |
554 |
551 |
3.54 |
|
|
|
| 38 |
A |
407 |
405 |
1.25 |
|
|
|
| 39 |
A |
383 |
381 |
0.16 |
|
|
|
| 40 |
A |
304 |
303 |
0.07 |
|
|
|
| 41 |
A |
249 |
248 |
0.01 |
|
|
|
| 42 |
A |
103 |
102 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30728.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30550.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.025 |
|
|
|
| 2 |
C |
-0.033 |
|
|
|
| 3 |
H |
-0.025 |
|
|
|
| 4 |
C |
-0.033 |
|
|
|
| 5 |
H |
0.027 |
|
|
|
| 6 |
H |
0.024 |
|
|
|
| 7 |
C |
0.042 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.027 |
|
|
|
| 10 |
C |
0.042 |
|
|
|
| 11 |
H |
0.009 |
|
|
|
| 12 |
C |
-0.160 |
|
|
|
| 13 |
H |
0.027 |
|
|
|
| 14 |
H |
0.027 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
| 16 |
C |
-0.004 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.225 |
0.000 |
0.023 |
0.226 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.435 |
0.000 |
0.346 |
| y |
0.000 |
-37.284 |
0.000 |
| z |
0.346 |
0.000 |
-39.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.151 |
0.000 |
0.346 |
| y |
0.000 |
1.576 |
0.000 |
| z |
0.346 |
0.000 |
-1.425 |
|
| Polar |
|---|
| 3z2-r2 | -2.849 |
|---|
| x2-y2 | -1.152 |
|---|
| xy | 0.000 |
|---|
| xz | 0.346 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.652 |
0.000 |
0.031 |
| y |
0.000 |
10.894 |
0.000 |
| z |
0.031 |
0.000 |
7.351 |
<r2> (average value of r
2) Å
2
| <r2> |
165.895 |
| (<r2>)1/2 |
12.880 |