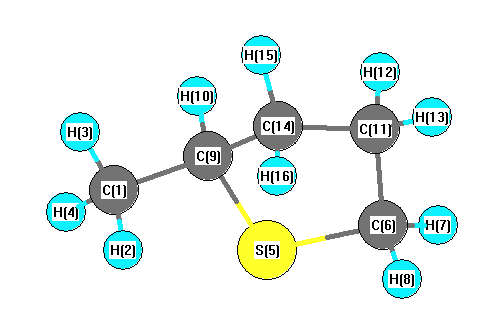
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3061 |
3043 |
20.18 |
|
|
|
| 2 |
A |
3048 |
3030 |
17.86 |
|
|
|
| 3 |
A |
3044 |
3027 |
20.39 |
|
|
|
| 4 |
A |
3027 |
3009 |
29.71 |
|
|
|
| 5 |
A |
3016 |
2999 |
25.97 |
|
|
|
| 6 |
A |
2983 |
2965 |
30.01 |
|
|
|
| 7 |
A |
2973 |
2956 |
23.07 |
|
|
|
| 8 |
A |
2968 |
2951 |
19.19 |
|
|
|
| 9 |
A |
2965 |
2947 |
41.43 |
|
|
|
| 10 |
A |
2959 |
2941 |
10.10 |
|
|
|
| 11 |
A |
1424 |
1416 |
0.80 |
|
|
|
| 12 |
A |
1418 |
1410 |
5.16 |
|
|
|
| 13 |
A |
1408 |
1400 |
5.71 |
|
|
|
| 14 |
A |
1404 |
1396 |
4.25 |
|
|
|
| 15 |
A |
1399 |
1391 |
2.30 |
|
|
|
| 16 |
A |
1341 |
1334 |
3.07 |
|
|
|
| 17 |
A |
1321 |
1314 |
1.67 |
|
|
|
| 18 |
A |
1301 |
1293 |
0.48 |
|
|
|
| 19 |
A |
1262 |
1254 |
0.09 |
|
|
|
| 20 |
A |
1235 |
1228 |
4.91 |
|
|
|
| 21 |
A |
1223 |
1216 |
17.32 |
|
|
|
| 22 |
A |
1183 |
1176 |
4.81 |
|
|
|
| 23 |
A |
1145 |
1138 |
10.77 |
|
|
|
| 24 |
A |
1120 |
1113 |
0.73 |
|
|
|
| 25 |
A |
1073 |
1067 |
0.87 |
|
|
|
| 26 |
A |
1031 |
1025 |
1.29 |
|
|
|
| 27 |
A |
1013 |
1007 |
2.65 |
|
|
|
| 28 |
A |
994 |
989 |
2.93 |
|
|
|
| 29 |
A |
933 |
928 |
2.56 |
|
|
|
| 30 |
A |
926 |
921 |
0.92 |
|
|
|
| 31 |
A |
885 |
880 |
0.88 |
|
|
|
| 32 |
A |
834 |
829 |
0.41 |
|
|
|
| 33 |
A |
826 |
821 |
2.81 |
|
|
|
| 34 |
A |
667 |
663 |
1.94 |
|
|
|
| 35 |
A |
609 |
606 |
3.22 |
|
|
|
| 36 |
A |
512 |
509 |
0.87 |
|
|
|
| 37 |
A |
446 |
444 |
0.21 |
|
|
|
| 38 |
A |
400 |
398 |
0.74 |
|
|
|
| 39 |
A |
275 |
274 |
1.13 |
|
|
|
| 40 |
A |
255 |
254 |
0.19 |
|
|
|
| 41 |
A |
235 |
234 |
0.12 |
|
|
|
| 42 |
A |
93 |
92 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30116.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29941.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.011 |
|
|
|
| 2 |
H |
0.038 |
|
|
|
| 3 |
H |
0.032 |
|
|
|
| 4 |
H |
0.040 |
|
|
|
| 5 |
S |
-0.059 |
|
|
|
| 6 |
C |
-0.108 |
|
|
|
| 7 |
H |
0.055 |
|
|
|
| 8 |
H |
0.061 |
|
|
|
| 9 |
C |
-0.213 |
|
|
|
| 10 |
H |
0.045 |
|
|
|
| 11 |
C |
-0.023 |
|
|
|
| 12 |
H |
0.034 |
|
|
|
| 13 |
H |
0.021 |
|
|
|
| 14 |
C |
0.020 |
|
|
|
| 15 |
H |
0.022 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.322 |
1.880 |
0.000 |
1.907 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.695 |
-0.251 |
0.920 |
| y |
-0.251 |
-47.921 |
-0.311 |
| z |
0.920 |
-0.311 |
-46.070 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.300 |
-0.251 |
0.920 |
| y |
-0.251 |
-3.538 |
-0.311 |
| z |
0.920 |
-0.311 |
-0.762 |
|
| Polar |
|---|
| 3z2-r2 | -1.525 |
|---|
| x2-y2 | 5.226 |
|---|
| xy | -0.251 |
|---|
| xz | 0.920 |
|---|
| yz | -0.311 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.488 |
-0.133 |
0.290 |
| y |
-0.133 |
11.010 |
-0.049 |
| z |
0.290 |
-0.049 |
8.595 |
<r2> (average value of r
2) Å
2
| <r2> |
206.225 |
| (<r2>)1/2 |
14.361 |