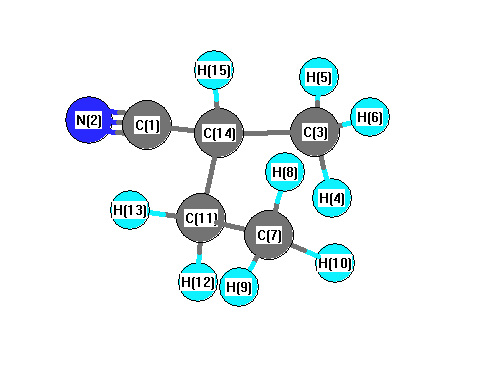
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3044 |
12.99 |
|
|
|
| 2 |
A |
3147 |
3037 |
13.36 |
|
|
|
| 3 |
A |
3136 |
3027 |
26.05 |
|
|
|
| 4 |
A |
3128 |
3018 |
28.35 |
|
|
|
| 5 |
A |
3096 |
2988 |
5.92 |
|
|
|
| 6 |
A |
3058 |
2951 |
13.17 |
|
|
|
| 7 |
A |
3052 |
2945 |
20.60 |
|
|
|
| 8 |
A |
3046 |
2939 |
19.97 |
|
|
|
| 9 |
A |
3034 |
2928 |
3.29 |
|
|
|
| 10 |
A |
2361 |
2278 |
11.54 |
|
|
|
| 11 |
A |
1479 |
1427 |
3.29 |
|
|
|
| 12 |
A |
1475 |
1424 |
7.96 |
|
|
|
| 13 |
A |
1470 |
1418 |
6.51 |
|
|
|
| 14 |
A |
1463 |
1412 |
9.81 |
|
|
|
| 15 |
A |
1457 |
1406 |
0.78 |
|
|
|
| 16 |
A |
1397 |
1348 |
2.05 |
|
|
|
| 17 |
A |
1390 |
1342 |
4.41 |
|
|
|
| 18 |
A |
1364 |
1317 |
0.21 |
|
|
|
| 19 |
A |
1335 |
1289 |
1.49 |
|
|
|
| 20 |
A |
1302 |
1257 |
3.10 |
|
|
|
| 21 |
A |
1269 |
1224 |
1.90 |
|
|
|
| 22 |
A |
1175 |
1134 |
2.79 |
|
|
|
| 23 |
A |
1144 |
1104 |
0.21 |
|
|
|
| 24 |
A |
1108 |
1069 |
4.07 |
|
|
|
| 25 |
A |
1059 |
1022 |
1.95 |
|
|
|
| 26 |
A |
999 |
964 |
2.06 |
|
|
|
| 27 |
A |
968 |
934 |
5.12 |
|
|
|
| 28 |
A |
904 |
873 |
0.40 |
|
|
|
| 29 |
A |
812 |
784 |
0.49 |
|
|
|
| 30 |
A |
769 |
742 |
3.90 |
|
|
|
| 31 |
A |
568 |
548 |
0.07 |
|
|
|
| 32 |
A |
539 |
520 |
1.07 |
|
|
|
| 33 |
A |
396 |
382 |
0.24 |
|
|
|
| 34 |
A |
328 |
317 |
0.51 |
|
|
|
| 35 |
A |
268 |
259 |
0.14 |
|
|
|
| 36 |
A |
224 |
216 |
0.19 |
|
|
|
| 37 |
A |
207 |
200 |
1.79 |
|
|
|
| 38 |
A |
164 |
158 |
4.79 |
|
|
|
| 39 |
A |
85 |
82 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28664.9 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 27661.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.022 |
|
|
|
| 2 |
N |
-0.143 |
|
|
|
| 3 |
C |
-0.029 |
|
|
|
| 4 |
H |
0.060 |
|
|
|
| 5 |
H |
0.062 |
|
|
|
| 6 |
H |
0.051 |
|
|
|
| 7 |
C |
-0.078 |
|
|
|
| 8 |
H |
0.046 |
|
|
|
| 9 |
H |
0.051 |
|
|
|
| 10 |
H |
0.050 |
|
|
|
| 11 |
C |
-0.061 |
|
|
|
| 12 |
H |
0.050 |
|
|
|
| 13 |
H |
0.054 |
|
|
|
| 14 |
C |
-0.161 |
|
|
|
| 15 |
H |
0.069 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.768 |
1.287 |
0.473 |
4.009 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.701 |
-3.707 |
-0.801 |
| y |
-3.707 |
-37.054 |
-0.094 |
| z |
-0.801 |
-0.094 |
-36.543 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.903 |
-3.707 |
-0.801 |
| y |
-3.707 |
5.069 |
-0.094 |
| z |
-0.801 |
-0.094 |
5.834 |
|
| Polar |
|---|
| 3z2-r2 | 11.668 |
|---|
| x2-y2 | -10.648 |
|---|
| xy | -3.707 |
|---|
| xz | -0.801 |
|---|
| yz | -0.094 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
199.387 |
| (<r2>)1/2 |
14.120 |