Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -114.288596 |
| Energy at 298.15K | -114.289999 |
| HF Energy | -114.288596 |
| Nuclear repulsion energy | 30.315799 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3521 |
3500 |
49.19 |
|
|
|
| 2 |
A' |
2695 |
2679 |
190.75 |
|
|
|
| 3 |
A' |
1467 |
1459 |
18.18 |
|
|
|
| 4 |
A' |
1291 |
1284 |
46.23 |
|
|
|
| 5 |
A' |
1163 |
1156 |
115.01 |
|
|
|
| 6 |
A" |
1080 |
1074 |
104.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5608.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 5575.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.751 |
0.000 |
| O2 |
0.011 |
-0.576 |
0.000 |
| H3 |
-1.113 |
0.963 |
0.000 |
| H4 |
0.955 |
-0.860 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3278 | 1.1438 | 1.8670 |
O2 | 1.3278 | | 1.9061 | 0.9852 | H3 | 1.1438 | 1.9061 | | 2.7563 | H4 | 1.8670 | 0.9852 | 2.7563 | |
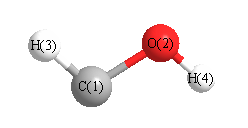 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
106.705 |
|
O2 |
C1 |
H3 |
100.659 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.170 |
|
|
|
| 2 |
O |
-0.052 |
|
|
|
| 3 |
H |
0.041 |
|
|
|
| 4 |
H |
0.181 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.322 |
-1.768 |
0.000 |
1.798 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-10.970 |
-3.335 |
0.000 |
| y |
-3.335 |
-13.556 |
0.000 |
| z |
0.000 |
0.000 |
-11.764 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.690 |
-3.335 |
0.000 |
| y |
-3.335 |
-2.189 |
0.000 |
| z |
0.000 |
0.000 |
0.499 |
|
| Polar |
|---|
| 3z2-r2 | 0.998 |
|---|
| x2-y2 | 2.586 |
|---|
| xy | -3.335 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.680 |
-0.607 |
0.000 |
| y |
-0.607 |
2.538 |
0.000 |
| z |
0.000 |
0.000 |
1.463 |
<r2> (average value of r
2) Å
2
| <r2> |
17.419 |
| (<r2>)1/2 |
4.174 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -114.282107 |
| Energy at 298.15K | -114.283494 |
| HF Energy | -114.282107 |
| Nuclear repulsion energy | 30.312472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3246 |
3227 |
76.34 |
|
|
|
| 2 |
A' |
2610 |
2595 |
271.59 |
|
|
|
| 3 |
A' |
1419 |
1411 |
45.86 |
|
|
|
| 4 |
A' |
1309 |
1302 |
30.77 |
|
|
|
| 5 |
A' |
1129 |
1123 |
11.48 |
|
|
|
| 6 |
A" |
1006 |
1000 |
27.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5359.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 5328.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.125 |
0.746 |
0.000 |
| O2 |
0.125 |
-0.570 |
0.000 |
| H3 |
-0.974 |
1.082 |
0.000 |
| H4 |
-0.774 |
-1.001 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3160 | 1.1487 | 1.9651 |
O2 | 1.3160 | | 1.9840 | 0.9973 | H3 | 1.1487 | 1.9840 | | 2.0930 | H4 | 1.9651 | 0.9973 | 2.0930 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
115.622 |
|
O2 |
C1 |
H3 |
107.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.125 |
|
|
|
| 2 |
O |
-0.044 |
|
|
|
| 3 |
H |
0.021 |
|
|
|
| 4 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.746 |
-1.779 |
0.000 |
3.272 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.262 |
-0.103 |
0.000 |
| y |
-0.103 |
-13.038 |
0.000 |
| z |
0.000 |
0.000 |
-11.739 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.127 |
-0.103 |
0.000 |
| y |
-0.103 |
-1.038 |
0.000 |
| z |
0.000 |
0.000 |
0.911 |
|
| Polar |
|---|
| 3z2-r2 | 1.822 |
|---|
| x2-y2 | 0.776 |
|---|
| xy | -0.103 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.558 |
0.019 |
0.000 |
| y |
0.019 |
3.207 |
0.000 |
| z |
0.000 |
0.000 |
1.473 |
<r2> (average value of r
2) Å
2
| <r2> |
17.589 |
| (<r2>)1/2 |
4.194 |
