Jump to
S1C2
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.581926 |
| Energy at 298.15K | -133.587333 |
| HF Energy | -133.083326 |
| Nuclear repulsion energy | 70.084477 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3371 |
3213 |
|
|
|
|
| 2 |
A' |
3137 |
2990 |
|
|
|
|
| 3 |
A' |
3094 |
2948 |
|
|
|
|
| 4 |
A' |
3037 |
2894 |
|
|
|
|
| 5 |
A' |
1700 |
1620 |
|
|
|
|
| 6 |
A' |
1470 |
1401 |
|
|
|
|
| 7 |
A' |
1426 |
1359 |
|
|
|
|
| 8 |
A' |
1386 |
1321 |
|
|
|
|
| 9 |
A' |
1280 |
1220 |
|
|
|
|
| 10 |
A' |
1065 |
1015 |
|
|
|
|
| 11 |
A' |
912 |
870 |
|
|
|
|
| 12 |
A' |
479 |
457 |
|
|
|
|
| 13 |
A" |
3119 |
2973 |
|
|
|
|
| 14 |
A" |
1463 |
1395 |
|
|
|
|
| 15 |
A" |
1135 |
1082 |
|
|
|
|
| 16 |
A" |
1057 |
1008 |
|
|
|
|
| 17 |
A" |
673 |
641 |
|
|
|
|
| 18 |
A" |
165 |
157 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14984.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 14281.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.032 |
-0.622 |
0.000 |
| C2 |
0.000 |
0.488 |
0.000 |
| N3 |
1.284 |
0.375 |
0.000 |
| H4 |
-0.547 |
-1.617 |
0.000 |
| H5 |
-1.684 |
-0.542 |
0.891 |
| H6 |
-1.684 |
-0.542 |
-0.891 |
| H7 |
-0.404 |
1.519 |
0.000 |
| H8 |
1.518 |
-0.636 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5156 | 2.5212 | 1.1061 | 1.1074 | 1.1074 | 2.2318 | 2.5496 |
C2 | 1.5156 | | 1.2891 | 2.1747 | 2.1654 | 2.1654 | 1.1076 | 1.8887 | N3 | 2.5212 | 1.2891 | | 2.7055 | 3.2314 | 3.2314 | 2.0395 | 1.0372 | H4 | 1.1061 | 2.1747 | 2.7055 | | 1.8006 | 1.8006 | 3.1394 | 2.2864 | H5 | 1.1074 | 2.1654 | 3.2314 | 1.8006 | | 1.7827 | 2.5846 | 3.3248 | H6 | 1.1074 | 2.1654 | 3.2314 | 1.8006 | 1.7827 | | 2.5846 | 3.3248 | H7 | 2.2318 | 1.1076 | 2.0395 | 3.1394 | 2.5846 | 2.5846 | | 2.8876 | H8 | 2.5496 | 1.8887 | 1.0372 | 2.2864 | 3.3248 | 3.3248 | 2.8876 | |
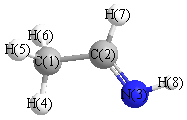 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.851 |
|
C1 |
C2 |
H7 |
115.728 |
| C2 |
C1 |
H4 |
111.136 |
|
C2 |
C1 |
H5 |
110.325 |
| C2 |
C1 |
H6 |
110.325 |
|
C2 |
N3 |
H8 |
108.075 |
| N3 |
C2 |
H7 |
116.421 |
|
H4 |
C1 |
H5 |
108.873 |
| H4 |
C1 |
H6 |
108.873 |
|
H5 |
C1 |
H6 |
107.207 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
