Jump to
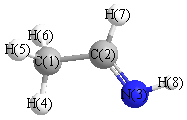
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.891131 |
| Energy at 298.15K | -133.896548 |
| HF Energy | -133.891131 |
| Nuclear repulsion energy | 70.068085 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3265 |
3270 |
10.55 |
|
|
|
| 2 |
A' |
3053 |
3058 |
13.43 |
|
|
|
| 3 |
A' |
2942 |
2947 |
12.73 |
|
|
|
| 4 |
A' |
2871 |
2876 |
89.66 |
|
|
|
| 5 |
A' |
1663 |
1666 |
61.51 |
|
|
|
| 6 |
A' |
1398 |
1400 |
10.88 |
|
|
|
| 7 |
A' |
1381 |
1383 |
22.71 |
|
|
|
| 8 |
A' |
1326 |
1328 |
9.09 |
|
|
|
| 9 |
A' |
1232 |
1234 |
32.48 |
|
|
|
| 10 |
A' |
1018 |
1020 |
27.17 |
|
|
|
| 11 |
A' |
896 |
897 |
7.62 |
|
|
|
| 12 |
A' |
472 |
473 |
13.40 |
|
|
|
| 13 |
A" |
2993 |
2998 |
17.02 |
|
|
|
| 14 |
A" |
1408 |
1410 |
6.62 |
|
|
|
| 15 |
A" |
1080 |
1081 |
7.37 |
|
|
|
| 16 |
A" |
1035 |
1037 |
8.22 |
|
|
|
| 17 |
A" |
659 |
660 |
41.66 |
|
|
|
| 18 |
A" |
204 |
204 |
0.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14448.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 14471.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.040 |
-0.641 |
0.000 |
| C2 |
0.000 |
0.455 |
0.000 |
| N3 |
1.257 |
0.184 |
0.000 |
| H4 |
-0.551 |
-1.633 |
0.000 |
| H5 |
-1.700 |
-0.552 |
0.891 |
| H6 |
-1.700 |
-0.552 |
-0.891 |
| H7 |
-0.413 |
1.496 |
0.000 |
| H8 |
1.806 |
1.067 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5108 | 2.4399 | 1.1066 | 1.1119 | 1.1119 | 2.2261 | 3.3186 |
C2 | 1.5108 | | 1.2856 | 2.1601 | 2.1672 | 2.1672 | 1.1191 | 1.9066 | N3 | 2.4399 | 1.2856 | | 2.5628 | 3.1738 | 3.1738 | 2.1234 | 1.0400 | H4 | 1.1066 | 2.1601 | 2.5628 | | 1.8115 | 1.8115 | 3.1318 | 3.5839 | H5 | 1.1119 | 2.1672 | 3.1738 | 1.8115 | | 1.7813 | 2.5770 | 3.9626 | H6 | 1.1119 | 2.1672 | 3.1738 | 1.8115 | 1.7813 | | 2.5770 | 3.9626 | H7 | 2.2261 | 1.1191 | 2.1234 | 3.1318 | 2.5770 | 2.5770 | | 2.2601 | H8 | 3.3186 | 1.9066 | 1.0400 | 3.5839 | 3.9626 | 3.9626 | 2.2601 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.290 |
|
C1 |
C2 |
H7 |
114.845 |
| C2 |
C1 |
H4 |
110.281 |
|
C2 |
C1 |
H5 |
110.531 |
| C2 |
C1 |
H6 |
110.531 |
|
C2 |
N3 |
H8 |
109.682 |
| N3 |
C2 |
H7 |
123.865 |
|
H4 |
C1 |
H5 |
109.478 |
| H4 |
C1 |
H6 |
109.478 |
|
H5 |
C1 |
H6 |
106.462 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.049 |
|
|
|
| 2 |
C |
-0.046 |
|
|
|
| 3 |
N |
-0.109 |
|
|
|
| 4 |
H |
0.017 |
|
|
|
| 5 |
H |
0.025 |
|
|
|
| 6 |
H |
0.025 |
|
|
|
| 7 |
H |
-0.031 |
|
|
|
| 8 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.212 |
1.202 |
0.000 |
1.707 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.520 |
2.274 |
0.000 |
| y |
2.274 |
-18.110 |
0.000 |
| z |
0.000 |
0.000 |
-19.610 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.659 |
2.274 |
0.000 |
| y |
2.274 |
1.455 |
0.000 |
| z |
0.000 |
0.000 |
-0.795 |
|
| Polar |
|---|
| 3z2-r2 | -1.591 |
|---|
| x2-y2 | -1.409 |
|---|
| xy | 2.274 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.019 |
0.454 |
0.000 |
| y |
0.454 |
4.622 |
0.000 |
| z |
0.000 |
0.000 |
3.009 |
<r2> (average value of r
2) Å
2
| <r2> |
51.151 |
| (<r2>)1/2 |
7.152 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.890629 |
| Energy at 298.15K | -133.896003 |
| HF Energy | -133.890629 |
| Nuclear repulsion energy | 69.965442 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3209 |
3214 |
19.90 |
|
|
|
| 2 |
A' |
3027 |
3032 |
26.86 |
|
|
|
| 3 |
A' |
2955 |
2960 |
34.34 |
|
|
|
| 4 |
A' |
2935 |
2940 |
14.57 |
|
|
|
| 5 |
A' |
1659 |
1661 |
60.24 |
|
|
|
| 6 |
A' |
1407 |
1409 |
24.61 |
|
|
|
| 7 |
A' |
1381 |
1383 |
24.75 |
|
|
|
| 8 |
A' |
1332 |
1334 |
4.39 |
|
|
|
| 9 |
A' |
1244 |
1246 |
53.47 |
|
|
|
| 10 |
A' |
1030 |
1032 |
12.48 |
|
|
|
| 11 |
A' |
875 |
876 |
5.25 |
|
|
|
| 12 |
A' |
470 |
471 |
5.75 |
|
|
|
| 13 |
A" |
2992 |
2997 |
13.70 |
|
|
|
| 14 |
A" |
1401 |
1403 |
7.02 |
|
|
|
| 15 |
A" |
1105 |
1107 |
37.54 |
|
|
|
| 16 |
A" |
1027 |
1029 |
4.67 |
|
|
|
| 17 |
A" |
665 |
666 |
5.41 |
|
|
|
| 18 |
A" |
175 |
176 |
1.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14444.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 14467.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.029 |
-0.632 |
0.000 |
| C2 |
0.000 |
0.481 |
0.000 |
| N3 |
1.282 |
0.392 |
0.000 |
| H4 |
-0.553 |
-1.633 |
0.000 |
| H5 |
-1.691 |
-0.550 |
0.890 |
| H6 |
-1.691 |
-0.550 |
-0.890 |
| H7 |
-0.421 |
1.512 |
0.000 |
| H8 |
1.557 |
-0.616 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5155 | 2.5279 | 1.1090 | 1.1120 | 1.1120 | 2.2282 | 2.5869 |
C2 | 1.5155 | | 1.2852 | 2.1846 | 2.1711 | 2.1711 | 1.1137 | 1.9047 | N3 | 2.5279 | 1.2852 | | 2.7323 | 3.2433 | 3.2433 | 2.0382 | 1.0444 | H4 | 1.1090 | 2.1846 | 2.7323 | | 1.8060 | 1.8060 | 3.1476 | 2.3424 | H5 | 1.1120 | 2.1711 | 3.2433 | 1.8060 | | 1.7797 | 2.5800 | 3.3690 | H6 | 1.1120 | 2.1711 | 3.2433 | 1.8060 | 1.7797 | | 2.5800 | 3.3690 | H7 | 2.2282 | 1.1137 | 2.0382 | 3.1476 | 2.5800 | 2.5800 | | 2.9051 | H8 | 2.5869 | 1.9047 | 1.0444 | 2.3424 | 3.3690 | 3.3690 | 2.9051 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.814 |
|
C1 |
C2 |
H7 |
115.027 |
| C2 |
C1 |
H4 |
111.756 |
|
C2 |
C1 |
H5 |
110.500 |
| C2 |
C1 |
H6 |
110.500 |
|
C2 |
N3 |
H8 |
109.253 |
| N3 |
C2 |
H7 |
116.159 |
|
H4 |
C1 |
H5 |
108.805 |
| H4 |
C1 |
H6 |
108.805 |
|
H5 |
C1 |
H6 |
106.301 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.033 |
|
|
|
| 2 |
C |
-0.051 |
|
|
|
| 3 |
N |
-0.108 |
|
|
|
| 4 |
H |
0.007 |
|
|
|
| 5 |
H |
0.031 |
|
|
|
| 6 |
H |
0.031 |
|
|
|
| 7 |
H |
-0.002 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.749 |
-1.313 |
0.000 |
2.187 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.889 |
-2.804 |
0.000 |
| y |
-2.804 |
-19.172 |
0.000 |
| z |
0.000 |
0.000 |
-19.583 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.512 |
-2.804 |
0.000 |
| y |
-2.804 |
1.064 |
0.000 |
| z |
0.000 |
0.000 |
0.448 |
|
| Polar |
|---|
| 3z2-r2 | 0.895 |
|---|
| x2-y2 | -1.718 |
|---|
| xy | -2.804 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.843 |
-0.055 |
0.000 |
| y |
-0.055 |
4.573 |
0.000 |
| z |
0.000 |
0.000 |
3.001 |
<r2> (average value of r
2) Å
2
| <r2> |
51.284 |
| (<r2>)1/2 |
7.161 |
