Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.909856 |
| Energy at 298.15K | -133.915326 |
| HF Energy | -133.909856 |
| Nuclear repulsion energy | 70.728217 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3434 |
3314 |
1.69 |
|
|
|
| 2 |
A' |
3162 |
3052 |
8.44 |
|
|
|
| 3 |
A' |
3039 |
2933 |
10.58 |
|
|
|
| 4 |
A' |
2996 |
2891 |
72.32 |
|
|
|
| 5 |
A' |
1751 |
1690 |
69.96 |
|
|
|
| 6 |
A' |
1436 |
1386 |
15.17 |
|
|
|
| 7 |
A' |
1426 |
1376 |
23.33 |
|
|
|
| 8 |
A' |
1360 |
1313 |
10.42 |
|
|
|
| 9 |
A' |
1264 |
1220 |
32.04 |
|
|
|
| 10 |
A' |
1053 |
1016 |
28.56 |
|
|
|
| 11 |
A' |
931 |
899 |
6.38 |
|
|
|
| 12 |
A' |
483 |
466 |
15.89 |
|
|
|
| 13 |
A" |
3109 |
3001 |
11.27 |
|
|
|
| 14 |
A" |
1446 |
1395 |
9.19 |
|
|
|
| 15 |
A" |
1122 |
1083 |
7.73 |
|
|
|
| 16 |
A" |
1063 |
1026 |
9.15 |
|
|
|
| 17 |
A" |
679 |
655 |
49.81 |
|
|
|
| 18 |
A" |
204 |
196 |
0.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14978.9 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 14454.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.031 |
-0.634 |
0.000 |
| C2 |
0.000 |
0.449 |
0.000 |
| N3 |
1.245 |
0.183 |
0.000 |
| H4 |
-0.544 |
-1.619 |
0.000 |
| H5 |
-1.684 |
-0.547 |
0.885 |
| H6 |
-1.684 |
-0.547 |
-0.885 |
| H7 |
-0.402 |
1.484 |
0.000 |
| H8 |
1.787 |
1.057 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4959 | 2.4181 | 1.0986 | 1.1032 | 1.1032 | 2.2096 | 3.2868 |
C2 | 1.4959 | | 1.2729 | 2.1391 | 2.1473 | 2.1473 | 1.1097 | 1.8876 | N3 | 2.4181 | 1.2729 | | 2.5394 | 3.1452 | 3.1452 | 2.0986 | 1.0286 | H4 | 1.0986 | 2.1391 | 2.5394 | | 1.7978 | 1.7978 | 3.1063 | 3.5494 | H5 | 1.1032 | 2.1473 | 3.1452 | 1.7978 | | 1.7700 | 2.5591 | 3.9247 | H6 | 1.1032 | 2.1473 | 3.1452 | 1.7978 | 1.7700 | | 2.5591 | 3.9247 | H7 | 2.2096 | 1.1097 | 2.0986 | 3.1063 | 2.5591 | 2.5591 | | 2.2306 | H8 | 3.2868 | 1.8876 | 1.0286 | 3.5494 | 3.9247 | 3.9247 | 2.2306 | |
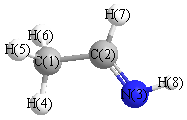 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.486 |
|
C1 |
C2 |
H7 |
115.185 |
| C2 |
C1 |
H4 |
110.129 |
|
C2 |
C1 |
H5 |
110.505 |
| C2 |
C1 |
H6 |
110.505 |
|
C2 |
N3 |
H8 |
109.747 |
| N3 |
C2 |
H7 |
123.330 |
|
H4 |
C1 |
H5 |
109.474 |
| H4 |
C1 |
H6 |
109.474 |
|
H5 |
C1 |
H6 |
106.686 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.059 |
|
|
|
| 2 |
C |
-0.057 |
|
|
|
| 3 |
N |
-0.171 |
|
|
|
| 4 |
H |
0.058 |
|
|
|
| 5 |
H |
0.060 |
|
|
|
| 6 |
H |
0.060 |
|
|
|
| 7 |
H |
0.013 |
|
|
|
| 8 |
H |
0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.331 |
1.299 |
0.000 |
1.860 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.302 |
2.338 |
0.000 |
| y |
2.338 |
-17.632 |
0.000 |
| z |
0.000 |
0.000 |
-19.448 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.762 |
2.338 |
0.000 |
| y |
2.338 |
1.743 |
0.000 |
| z |
0.000 |
0.000 |
-0.981 |
|
| Polar |
|---|
| 3z2-r2 | -1.961 |
|---|
| x2-y2 | -1.670 |
|---|
| xy | 2.338 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.770 |
0.383 |
0.000 |
| y |
0.383 |
4.353 |
0.000 |
| z |
0.000 |
0.000 |
2.941 |
<r2> (average value of r
2) Å
2
| <r2> |
50.251 |
| (<r2>)1/2 |
7.089 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.909217 |
| Energy at 298.15K | -133.914652 |
| HF Energy | -133.909217 |
| Nuclear repulsion energy | 70.670608 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3386 |
3268 |
6.93 |
|
|
|
| 2 |
A' |
3138 |
3028 |
19.65 |
|
|
|
| 3 |
A' |
3065 |
2958 |
31.20 |
|
|
|
| 4 |
A' |
3033 |
2926 |
10.27 |
|
|
|
| 5 |
A' |
1754 |
1692 |
70.26 |
|
|
|
| 6 |
A' |
1444 |
1394 |
26.96 |
|
|
|
| 7 |
A' |
1418 |
1369 |
24.21 |
|
|
|
| 8 |
A' |
1367 |
1319 |
6.28 |
|
|
|
| 9 |
A' |
1273 |
1229 |
53.11 |
|
|
|
| 10 |
A' |
1063 |
1026 |
17.06 |
|
|
|
| 11 |
A' |
914 |
882 |
5.22 |
|
|
|
| 12 |
A' |
482 |
465 |
6.51 |
|
|
|
| 13 |
A" |
3110 |
3001 |
8.27 |
|
|
|
| 14 |
A" |
1439 |
1389 |
9.70 |
|
|
|
| 15 |
A" |
1143 |
1103 |
43.01 |
|
|
|
| 16 |
A" |
1059 |
1022 |
8.00 |
|
|
|
| 17 |
A" |
687 |
663 |
6.00 |
|
|
|
| 18 |
A" |
179 |
172 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14976.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 14452.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.018 |
-0.623 |
0.000 |
| C2 |
0.000 |
0.478 |
0.000 |
| N3 |
1.268 |
0.383 |
0.000 |
| H4 |
-0.544 |
-1.617 |
0.000 |
| H5 |
-1.673 |
-0.542 |
0.884 |
| H6 |
-1.673 |
-0.542 |
-0.884 |
| H7 |
-0.414 |
1.503 |
0.000 |
| H8 |
1.537 |
-0.614 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5002 | 2.4983 | 1.1009 | 1.1033 | 1.1033 | 2.2108 | 2.5552 |
C2 | 1.5002 | | 1.2719 | 2.1648 | 2.1499 | 2.1499 | 1.1054 | 1.8853 | N3 | 2.4983 | 1.2719 | | 2.6991 | 3.2078 | 3.2078 | 2.0212 | 1.0321 | H4 | 1.1009 | 2.1648 | 2.6991 | | 1.7922 | 1.7922 | 3.1229 | 2.3103 | H5 | 1.1033 | 2.1499 | 3.2078 | 1.7922 | | 1.7683 | 2.5593 | 3.3303 | H6 | 1.1033 | 2.1499 | 3.2078 | 1.7922 | 1.7683 | | 2.5593 | 3.3303 | H7 | 2.2108 | 1.1054 | 2.0212 | 3.1229 | 2.5593 | 2.5593 | | 2.8787 | H8 | 2.5552 | 1.8853 | 1.0321 | 2.3103 | 3.3303 | 3.3303 | 2.8787 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.449 |
|
C1 |
C2 |
H7 |
115.257 |
| C2 |
C1 |
H4 |
111.746 |
|
C2 |
C1 |
H5 |
110.407 |
| C2 |
C1 |
H6 |
110.407 |
|
C2 |
N3 |
H8 |
109.377 |
| N3 |
C2 |
H7 |
116.294 |
|
H4 |
C1 |
H5 |
108.798 |
| H4 |
C1 |
H6 |
108.798 |
|
H5 |
C1 |
H6 |
106.526 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.074 |
|
|
|
| 2 |
C |
-0.057 |
|
|
|
| 3 |
N |
-0.171 |
|
|
|
| 4 |
H |
0.044 |
|
|
|
| 5 |
H |
0.067 |
|
|
|
| 6 |
H |
0.067 |
|
|
|
| 7 |
H |
0.039 |
|
|
|
| 8 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.892 |
-1.384 |
0.000 |
2.344 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.732 |
-2.983 |
0.000 |
| y |
-2.983 |
-18.777 |
0.000 |
| z |
0.000 |
0.000 |
-19.413 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.638 |
-2.983 |
0.000 |
| y |
-2.983 |
1.296 |
0.000 |
| z |
0.000 |
0.000 |
0.342 |
|
| Polar |
|---|
| 3z2-r2 | 0.684 |
|---|
| x2-y2 | -1.956 |
|---|
| xy | -2.983 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.606 |
-0.069 |
0.000 |
| y |
-0.069 |
4.332 |
0.000 |
| z |
0.000 |
0.000 |
2.929 |
<r2> (average value of r
2) Å
2
| <r2> |
50.311 |
| (<r2>)1/2 |
7.093 |
