Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.567332 |
| Energy at 298.15K | -133.572803 |
| HF Energy | -133.085124 |
| Nuclear repulsion energy | 70.296603 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3447 |
3307 |
4.15 |
|
|
|
| 2 |
A' |
3176 |
3047 |
11.68 |
|
|
|
| 3 |
A' |
3061 |
2937 |
20.85 |
|
|
|
| 4 |
A' |
3046 |
2922 |
54.95 |
|
|
|
| 5 |
A' |
1739 |
1669 |
48.27 |
|
|
|
| 6 |
A' |
1474 |
1414 |
6.61 |
|
|
|
| 7 |
A' |
1446 |
1387 |
27.03 |
|
|
|
| 8 |
A' |
1391 |
1335 |
6.03 |
|
|
|
| 9 |
A' |
1283 |
1231 |
28.33 |
|
|
|
| 10 |
A' |
1068 |
1025 |
26.47 |
|
|
|
| 11 |
A' |
939 |
901 |
3.61 |
|
|
|
| 12 |
A' |
483 |
464 |
15.40 |
|
|
|
| 13 |
A" |
3132 |
3005 |
18.08 |
|
|
|
| 14 |
A" |
1480 |
1420 |
6.89 |
|
|
|
| 15 |
A" |
1124 |
1078 |
6.62 |
|
|
|
| 16 |
A" |
1080 |
1037 |
11.52 |
|
|
|
| 17 |
A" |
681 |
654 |
46.03 |
|
|
|
| 18 |
A" |
195 |
187 |
1.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15122.4 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 14508.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.041 |
-0.636 |
0.000 |
| C2 |
0.000 |
0.456 |
0.000 |
| N3 |
1.254 |
0.178 |
0.000 |
| H4 |
-0.545 |
-1.620 |
0.000 |
| H5 |
-1.691 |
-0.550 |
0.891 |
| H6 |
-1.691 |
-0.550 |
-0.891 |
| H7 |
-0.393 |
1.495 |
0.000 |
| H8 |
1.791 |
1.058 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5090 | 2.4345 | 1.1022 | 1.1060 | 1.1060 | 2.2276 | 3.2999 |
C2 | 1.5090 | | 1.2842 | 2.1473 | 2.1601 | 2.1601 | 1.1104 | 1.8893 | N3 | 2.4345 | 1.2842 | | 2.5435 | 3.1611 | 3.1611 | 2.1084 | 1.0314 | H4 | 1.1022 | 2.1473 | 2.5435 | | 1.8031 | 1.8031 | 3.1194 | 3.5542 | H5 | 1.1060 | 2.1601 | 3.1611 | 1.8031 | | 1.7818 | 2.5813 | 3.9374 | H6 | 1.1060 | 2.1601 | 3.1611 | 1.8031 | 1.7818 | | 2.5813 | 3.9374 | H7 | 2.2276 | 1.1104 | 2.1084 | 3.1194 | 2.5813 | 2.5813 | | 2.2269 | H8 | 3.2999 | 1.8893 | 1.0314 | 3.5542 | 3.9374 | 3.9374 | 2.2269 | |
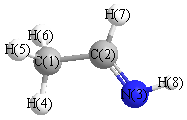 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.081 |
|
C1 |
C2 |
H7 |
115.678 |
| C2 |
C1 |
H4 |
109.659 |
|
C2 |
C1 |
H5 |
110.441 |
| C2 |
C1 |
H6 |
110.441 |
|
C2 |
N3 |
H8 |
108.867 |
| N3 |
C2 |
H7 |
123.241 |
|
H4 |
C1 |
H5 |
109.474 |
| H4 |
C1 |
H6 |
109.474 |
|
H5 |
C1 |
H6 |
107.312 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.566570 |
| Energy at 298.15K | -133.572001 |
| HF Energy | -133.084114 |
| Nuclear repulsion energy | 70.245411 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3407 |
3269 |
8.57 |
|
|
|
| 2 |
A' |
3152 |
3024 |
27.37 |
|
|
|
| 3 |
A' |
3111 |
2985 |
25.67 |
|
|
|
| 4 |
A' |
3053 |
2929 |
12.50 |
|
|
|
| 5 |
A' |
1737 |
1667 |
49.94 |
|
|
|
| 6 |
A' |
1479 |
1419 |
21.34 |
|
|
|
| 7 |
A' |
1438 |
1379 |
22.01 |
|
|
|
| 8 |
A' |
1399 |
1343 |
3.46 |
|
|
|
| 9 |
A' |
1291 |
1238 |
46.50 |
|
|
|
| 10 |
A' |
1075 |
1031 |
16.23 |
|
|
|
| 11 |
A' |
922 |
885 |
2.53 |
|
|
|
| 12 |
A' |
485 |
465 |
7.77 |
|
|
|
| 13 |
A" |
3134 |
3007 |
13.87 |
|
|
|
| 14 |
A" |
1473 |
1413 |
7.17 |
|
|
|
| 15 |
A" |
1150 |
1104 |
43.13 |
|
|
|
| 16 |
A" |
1071 |
1028 |
5.26 |
|
|
|
| 17 |
A" |
685 |
657 |
8.09 |
|
|
|
| 18 |
A" |
167 |
160 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15114.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 14501.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.028 |
-0.624 |
0.000 |
| C2 |
0.000 |
0.487 |
0.000 |
| N3 |
1.279 |
0.377 |
0.000 |
| H4 |
-0.543 |
-1.617 |
0.000 |
| H5 |
-1.679 |
-0.544 |
0.890 |
| H6 |
-1.679 |
-0.544 |
-0.890 |
| H7 |
-0.407 |
1.515 |
0.000 |
| H8 |
1.523 |
-0.628 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5136 | 2.5145 | 1.1046 | 1.1060 | 1.1060 | 2.2277 | 2.5506 |
C2 | 1.5136 | | 1.2836 | 2.1728 | 2.1624 | 2.1624 | 1.1061 | 1.8875 | N3 | 2.5145 | 1.2836 | | 2.7010 | 3.2236 | 3.2236 | 2.0345 | 1.0344 | H4 | 1.1046 | 2.1728 | 2.7010 | | 1.7980 | 1.7980 | 3.1352 | 2.2904 | H5 | 1.1060 | 2.1624 | 3.2236 | 1.7980 | | 1.7805 | 2.5794 | 3.3245 | H6 | 1.1060 | 2.1624 | 3.2236 | 1.7980 | 1.7805 | | 2.5794 | 3.3245 | H7 | 2.2277 | 1.1061 | 2.0345 | 3.1352 | 2.5794 | 2.5794 | | 2.8845 | H8 | 2.5506 | 1.8875 | 1.0344 | 2.2904 | 3.3245 | 3.3245 | 2.8845 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.850 |
|
C1 |
C2 |
H7 |
115.626 |
| C2 |
C1 |
H4 |
111.214 |
|
C2 |
C1 |
H5 |
110.306 |
| C2 |
C1 |
H6 |
110.306 |
|
C2 |
N3 |
H8 |
108.557 |
| N3 |
C2 |
H7 |
116.524 |
|
H4 |
C1 |
H5 |
108.850 |
| H4 |
C1 |
H6 |
108.850 |
|
H5 |
C1 |
H6 |
107.208 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
