Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.779119 |
| Energy at 298.15K | -133.784535 |
| HF Energy | -133.779119 |
| Nuclear repulsion energy | 70.284455 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3292 |
5.67 |
|
|
|
| 2 |
A' |
3084 |
3067 |
8.53 |
|
|
|
| 3 |
A' |
2964 |
2947 |
9.22 |
|
|
|
| 4 |
A' |
2895 |
2879 |
83.10 |
|
|
|
| 5 |
A' |
1686 |
1676 |
65.13 |
|
|
|
| 6 |
A' |
1387 |
1378 |
14.92 |
|
|
|
| 7 |
A' |
1377 |
1369 |
19.64 |
|
|
|
| 8 |
A' |
1311 |
1303 |
11.04 |
|
|
|
| 9 |
A' |
1226 |
1219 |
33.01 |
|
|
|
| 10 |
A' |
1019 |
1013 |
30.21 |
|
|
|
| 11 |
A' |
906 |
901 |
5.45 |
|
|
|
| 12 |
A' |
470 |
467 |
14.41 |
|
|
|
| 13 |
A" |
3032 |
3014 |
10.12 |
|
|
|
| 14 |
A" |
1397 |
1389 |
8.43 |
|
|
|
| 15 |
A" |
1079 |
1073 |
8.01 |
|
|
|
| 16 |
A" |
1028 |
1022 |
7.56 |
|
|
|
| 17 |
A" |
657 |
653 |
45.49 |
|
|
|
| 18 |
A" |
206 |
205 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14516.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 14432.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.037 |
-0.632 |
0.000 |
| C2 |
0.000 |
0.453 |
0.000 |
| N3 |
1.253 |
0.176 |
0.000 |
| H4 |
-0.549 |
-1.625 |
0.000 |
| H5 |
-1.696 |
-0.542 |
0.890 |
| H6 |
-1.696 |
-0.542 |
-0.890 |
| H7 |
-0.408 |
1.496 |
0.000 |
| H8 |
1.800 |
1.058 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5011 | 2.4285 | 1.1065 | 1.1111 | 1.1111 | 2.2188 | 3.3028 |
C2 | 1.5011 | | 1.2835 | 2.1497 | 2.1587 | 2.1587 | 1.1194 | 1.8993 | N3 | 2.4285 | 1.2835 | | 2.5478 | 3.1633 | 3.1633 | 2.1217 | 1.0387 | H4 | 1.1065 | 2.1497 | 2.5478 | | 1.8113 | 1.8113 | 3.1240 | 3.5669 | H5 | 1.1111 | 2.1587 | 3.1633 | 1.8113 | | 1.7797 | 2.5699 | 3.9472 | H6 | 1.1111 | 2.1587 | 3.1633 | 1.8113 | 1.7797 | | 2.5699 | 3.9472 | H7 | 2.2188 | 1.1194 | 2.1217 | 3.1240 | 2.5699 | 2.5699 | | 2.2510 | H8 | 3.3028 | 1.8993 | 1.0387 | 3.5669 | 3.9472 | 3.9472 | 2.2510 | |
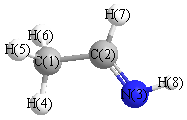 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.208 |
|
C1 |
C2 |
H7 |
114.933 |
| C2 |
C1 |
H4 |
110.139 |
|
C2 |
C1 |
H5 |
110.569 |
| C2 |
C1 |
H6 |
110.569 |
|
C2 |
N3 |
H8 |
109.295 |
| N3 |
C2 |
H7 |
123.859 |
|
H4 |
C1 |
H5 |
109.531 |
| H4 |
C1 |
H6 |
109.531 |
|
H5 |
C1 |
H6 |
106.427 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.030 |
|
|
|
| 2 |
C |
-0.059 |
|
|
|
| 3 |
N |
-0.138 |
|
|
|
| 4 |
H |
0.044 |
|
|
|
| 5 |
H |
0.052 |
|
|
|
| 6 |
H |
0.052 |
|
|
|
| 7 |
H |
-0.005 |
|
|
|
| 8 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.251 |
1.220 |
0.000 |
1.748 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.335 |
2.314 |
0.000 |
| y |
2.314 |
-17.942 |
0.000 |
| z |
0.000 |
0.000 |
-19.562 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.583 |
2.314 |
0.000 |
| y |
2.314 |
1.506 |
0.000 |
| z |
0.000 |
0.000 |
-0.923 |
|
| Polar |
|---|
| 3z2-r2 | -1.847 |
|---|
| x2-y2 | -1.393 |
|---|
| xy | 2.314 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.988 |
0.437 |
0.000 |
| y |
0.437 |
4.594 |
0.000 |
| z |
0.000 |
0.000 |
3.005 |
<r2> (average value of r
2) Å
2
| <r2> |
50.759 |
| (<r2>)1/2 |
7.125 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.778660 |
| Energy at 298.15K | -133.784035 |
| HF Energy | -133.778660 |
| Nuclear repulsion energy | 70.190124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3253 |
3234 |
14.49 |
|
|
|
| 2 |
A' |
3060 |
3043 |
18.86 |
|
|
|
| 3 |
A' |
2976 |
2959 |
32.74 |
|
|
|
| 4 |
A' |
2956 |
2939 |
11.33 |
|
|
|
| 5 |
A' |
1685 |
1676 |
62.97 |
|
|
|
| 6 |
A' |
1395 |
1387 |
27.25 |
|
|
|
| 7 |
A' |
1375 |
1367 |
25.58 |
|
|
|
| 8 |
A' |
1317 |
1309 |
7.39 |
|
|
|
| 9 |
A' |
1236 |
1228 |
49.12 |
|
|
|
| 10 |
A' |
1032 |
1026 |
13.94 |
|
|
|
| 11 |
A' |
886 |
881 |
6.28 |
|
|
|
| 12 |
A' |
468 |
465 |
5.75 |
|
|
|
| 13 |
A" |
3030 |
3012 |
7.64 |
|
|
|
| 14 |
A" |
1390 |
1382 |
8.97 |
|
|
|
| 15 |
A" |
1105 |
1099 |
40.24 |
|
|
|
| 16 |
A" |
1020 |
1014 |
5.48 |
|
|
|
| 17 |
A" |
662 |
658 |
4.46 |
|
|
|
| 18 |
A" |
177 |
176 |
1.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14511.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 14427.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.026 |
-0.624 |
0.000 |
| C2 |
0.000 |
0.478 |
0.000 |
| N3 |
1.279 |
0.385 |
0.000 |
| H4 |
-0.552 |
-1.626 |
0.000 |
| H5 |
-1.688 |
-0.540 |
0.889 |
| H6 |
-1.688 |
-0.540 |
-0.889 |
| H7 |
-0.416 |
1.512 |
0.000 |
| H8 |
1.542 |
-0.625 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5054 | 2.5162 | 1.1088 | 1.1114 | 1.1114 | 2.2210 | 2.5680 |
C2 | 1.5054 | | 1.2829 | 2.1753 | 2.1622 | 2.1622 | 1.1142 | 1.8961 | N3 | 2.5162 | 1.2829 | | 2.7197 | 3.2325 | 3.2325 | 2.0357 | 1.0433 | H4 | 1.1088 | 2.1753 | 2.7197 | | 1.8054 | 1.8054 | 3.1408 | 2.3209 | H5 | 1.1114 | 2.1622 | 3.2325 | 1.8054 | | 1.7779 | 2.5726 | 3.3510 | H6 | 1.1114 | 2.1622 | 3.2325 | 1.8054 | 1.7779 | | 2.5726 | 3.3510 | H7 | 2.2210 | 1.1142 | 2.0357 | 3.1408 | 2.5726 | 2.5726 | | 2.8982 | H8 | 2.5680 | 1.8961 | 1.0433 | 2.3209 | 3.3510 | 3.3510 | 2.8982 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.780 |
|
C1 |
C2 |
H7 |
115.137 |
| C2 |
C1 |
H4 |
111.734 |
|
C2 |
C1 |
H5 |
110.530 |
| C2 |
C1 |
H6 |
110.530 |
|
C2 |
N3 |
H8 |
108.759 |
| N3 |
C2 |
H7 |
116.083 |
|
H4 |
C1 |
H5 |
108.820 |
| H4 |
C1 |
H6 |
108.820 |
|
H5 |
C1 |
H6 |
106.233 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.045 |
|
|
|
| 2 |
C |
-0.062 |
|
|
|
| 3 |
N |
-0.138 |
|
|
|
| 4 |
H |
0.033 |
|
|
|
| 5 |
H |
0.058 |
|
|
|
| 6 |
H |
0.058 |
|
|
|
| 7 |
H |
0.025 |
|
|
|
| 8 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.813 |
-1.336 |
0.000 |
2.252 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.779 |
-2.825 |
0.000 |
| y |
-2.825 |
-18.971 |
0.000 |
| z |
0.000 |
0.000 |
-19.536 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.526 |
-2.825 |
0.000 |
| y |
-2.825 |
1.186 |
0.000 |
| z |
0.000 |
0.000 |
0.339 |
|
| Polar |
|---|
| 3z2-r2 | 0.679 |
|---|
| x2-y2 | -1.808 |
|---|
| xy | -2.825 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.820 |
-0.038 |
0.000 |
| y |
-0.038 |
4.553 |
0.000 |
| z |
0.000 |
0.000 |
2.996 |
<r2> (average value of r
2) Å
2
| <r2> |
50.894 |
| (<r2>)1/2 |
7.134 |
