Jump to
S2C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.115346 |
| Energy at 298.15K | -498.115205 |
| HF Energy | -497.826641 |
| Nuclear repulsion energy | 38.049678 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.046 |
1.215 |
0.000 |
| Cl2 |
0.046 |
-0.513 |
0.000 |
| H3 |
-1.062 |
1.435 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.7286 | 1.1293 |
Cl2 | 1.7286 | | 2.2415 | H3 | 1.1293 | 2.2415 | |
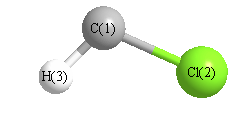 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
101.240 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.108262 |
| Energy at 298.15K | -498.108113 |
| HF Energy | -497.851401 |
| Nuclear repulsion energy | 38.408778 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.037 |
1.171 |
0.000 |
| Cl2 |
0.037 |
-0.520 |
0.000 |
| H3 |
-0.857 |
1.810 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.6911 | 1.0995 |
Cl2 | 1.6911 | | 2.4961 | H3 | 1.0995 | 2.4961 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
125.545 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
