Jump to
S2C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.531597 |
| Energy at 298.15K | -498.531448 |
| HF Energy | -498.531597 |
| Nuclear repulsion energy | 38.044138 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.047 |
1.216 |
0.000 |
| Cl2 |
0.047 |
-0.513 |
0.000 |
| H3 |
-1.075 |
1.417 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.7285 | 1.1395 |
Cl2 | 1.7285 | | 2.2319 | H3 | 1.1395 | 2.2319 | |
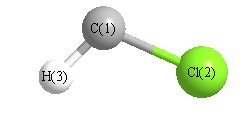 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
100.164 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.149 |
|
|
|
| 2 |
Cl |
0.051 |
|
|
|
| 3 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.383 |
-0.717 |
0.000 |
1.558 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.389 |
-2.370 |
0.000 |
| y |
-2.370 |
-19.859 |
0.000 |
| z |
0.000 |
0.000 |
-17.856 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.532 |
-2.370 |
0.000 |
| y |
-2.370 |
-1.236 |
0.000 |
| z |
0.000 |
0.000 |
1.768 |
|
| Polar |
|---|
| 3z2-r2 | 3.536 |
|---|
| x2-y2 | 0.470 |
|---|
| xy | -2.370 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.515 |
-0.359 |
0.000 |
| y |
-0.359 |
4.783 |
0.000 |
| z |
0.000 |
0.000 |
1.857 |
<r2> (average value of r
2) Å
2
| <r2> |
28.440 |
| (<r2>)1/2 |
5.333 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.527477 |
| Energy at 298.15K | -498.527314 |
| HF Energy | -498.527477 |
| Nuclear repulsion energy | 38.616062 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.037 |
1.163 |
0.000 |
| Cl2 |
0.037 |
-0.517 |
0.000 |
| H3 |
-0.859 |
1.807 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
| C1 | | 1.6800 | 1.1041 |
Cl2 | 1.6800 | | 2.4911 | H3 | 1.1041 | 2.4911 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl2 |
C1 |
H3 |
125.693 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.147 |
|
|
|
| 2 |
Cl |
0.042 |
|
|
|
| 3 |
H |
0.105 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.763 |
0.153 |
0.000 |
0.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.257 |
-1.474 |
0.000 |
| y |
-1.474 |
-17.054 |
0.000 |
| z |
0.000 |
0.000 |
-19.413 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.023 |
-1.474 |
0.000 |
| y |
-1.474 |
1.781 |
0.000 |
| z |
0.000 |
0.000 |
-1.758 |
|
| Polar |
|---|
| 3z2-r2 | -3.516 |
|---|
| x2-y2 | -1.203 |
|---|
| xy | -1.474 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.999 |
-0.239 |
0.000 |
| y |
-0.239 |
4.561 |
0.000 |
| z |
0.000 |
0.000 |
1.581 |
<r2> (average value of r
2) Å
2
| <r2> |
28.089 |
| (<r2>)1/2 |
5.300 |
