Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -193.216134 |
| Energy at 298.15K | |
| HF Energy | -193.216134 |
| Nuclear repulsion energy | 147.591764 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3190 |
3172 |
0.01 |
|
|
|
| 2 |
A1 |
3183 |
3164 |
7.55 |
|
|
|
| 3 |
A1 |
3145 |
3127 |
1.41 |
|
|
|
| 4 |
A1 |
1535 |
1526 |
0.01 |
|
|
|
| 5 |
A1 |
1422 |
1414 |
2.75 |
|
|
|
| 6 |
A1 |
1115 |
1108 |
0.02 |
|
|
|
| 7 |
A1 |
1044 |
1038 |
7.56 |
|
|
|
| 8 |
A1 |
1012 |
1006 |
13.17 |
|
|
|
| 9 |
A1 |
810 |
805 |
0.33 |
|
|
|
| 10 |
A2 |
865 |
860 |
0.00 |
|
|
|
| 11 |
A2 |
796 |
791 |
0.00 |
|
|
|
| 12 |
A2 |
505 |
502 |
0.00 |
|
|
|
| 13 |
B1 |
850 |
845 |
5.21 |
|
|
|
| 14 |
B1 |
694 |
690 |
3.79 |
|
|
|
| 15 |
B1 |
656 |
652 |
69.67 |
|
|
|
| 16 |
B1 |
468 |
466 |
9.25 |
|
|
|
| 17 |
B2 |
3165 |
3146 |
12.45 |
|
|
|
| 18 |
B2 |
3146 |
3128 |
9.01 |
|
|
|
| 19 |
B2 |
1349 |
1341 |
37.78 |
|
|
|
| 20 |
B2 |
1229 |
1222 |
0.00 |
|
|
|
| 21 |
B2 |
1168 |
1161 |
2.03 |
|
|
|
| 22 |
B2 |
948 |
943 |
0.00 |
|
|
|
| 23 |
B2 |
889 |
884 |
3.76 |
|
|
|
| 24 |
B2 |
67i |
66i |
8.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16558.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 16462.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.233 |
| C2 |
0.000 |
1.131 |
0.394 |
| C3 |
0.000 |
0.686 |
-1.011 |
| C4 |
0.000 |
-0.686 |
-1.011 |
| C5 |
0.000 |
-1.131 |
0.394 |
| H6 |
0.000 |
0.000 |
2.328 |
| H7 |
0.000 |
2.180 |
0.722 |
| H8 |
0.000 |
1.350 |
-1.883 |
| H9 |
0.000 |
-1.350 |
-1.883 |
| H10 |
0.000 |
-2.180 |
0.722 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4080 | 2.3468 | 2.3468 | 1.4080 | 1.0950 | 2.2390 | 3.3964 | 3.3964 | 2.2390 |
C2 | 1.4080 | | 1.4743 | 2.2971 | 2.2618 | 2.2402 | 1.0988 | 2.2882 | 3.3680 | 3.3269 | C3 | 2.3468 | 1.4743 | | 1.3718 | 2.2971 | 3.4091 | 2.2879 | 1.0963 | 2.2150 | 3.3489 | C4 | 2.3468 | 2.2971 | 1.3718 | | 1.4743 | 3.4091 | 3.3489 | 2.2150 | 1.0963 | 2.2879 | C5 | 1.4080 | 2.2618 | 2.2971 | 1.4743 | | 2.2402 | 3.3269 | 3.3680 | 2.2882 | 1.0988 | H6 | 1.0950 | 2.2402 | 3.4091 | 3.4091 | 2.2402 | | 2.7078 | 4.4226 | 4.4226 | 2.7078 | H7 | 2.2390 | 1.0988 | 2.2879 | 3.3489 | 3.3269 | 2.7078 | | 2.7339 | 4.3871 | 4.3596 | H8 | 3.3964 | 2.2882 | 1.0963 | 2.2150 | 3.3680 | 4.4226 | 2.7339 | | 2.7004 | 4.3871 | H9 | 3.3964 | 3.3680 | 2.2150 | 1.0963 | 2.2882 | 4.4226 | 4.3871 | 2.7004 | | 2.7339 | H10 | 2.2390 | 3.3269 | 3.3489 | 2.2879 | 1.0988 | 2.7078 | 4.3596 | 4.3871 | 2.7339 | |
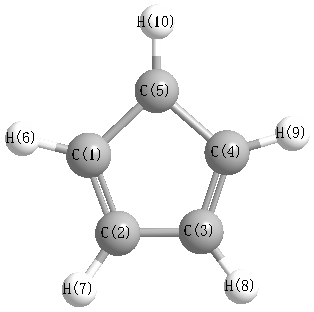 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.996 |
|
C1 |
C2 |
H7 |
126.104 |
| C1 |
C5 |
C4 |
108.996 |
|
C1 |
C5 |
H10 |
126.104 |
| C2 |
C1 |
C5 |
106.873 |
|
C2 |
C1 |
H6 |
126.563 |
| C2 |
C3 |
C4 |
107.568 |
|
C2 |
C3 |
H8 |
125.136 |
| C3 |
C2 |
H7 |
124.901 |
|
C3 |
C4 |
C5 |
107.568 |
| C3 |
C4 |
H9 |
127.296 |
|
C4 |
C3 |
H8 |
127.296 |
| C4 |
C5 |
H10 |
124.901 |
|
C5 |
C1 |
H6 |
126.563 |
| C5 |
C4 |
H9 |
125.136 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.003 |
|
|
|
| 2 |
C |
0.032 |
|
|
|
| 3 |
C |
0.008 |
|
|
|
| 4 |
C |
0.008 |
|
|
|
| 5 |
C |
0.032 |
|
|
|
| 6 |
H |
-0.023 |
|
|
|
| 7 |
H |
-0.009 |
|
|
|
| 8 |
H |
-0.019 |
|
|
|
| 9 |
H |
-0.019 |
|
|
|
| 10 |
H |
-0.009 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.060 |
0.060 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.744 |
0.000 |
0.000 |
| y |
0.000 |
-25.838 |
0.000 |
| z |
0.000 |
0.000 |
-27.646 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.001 |
0.000 |
0.000 |
| y |
0.000 |
4.357 |
0.000 |
| z |
0.000 |
0.000 |
1.645 |
|
| Polar |
|---|
| 3z2-r2 | 3.289 |
|---|
| x2-y2 | -6.905 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.207 |
0.000 |
0.000 |
| y |
0.000 |
9.377 |
0.000 |
| z |
0.000 |
0.000 |
8.641 |
<r2> (average value of r
2) Å
2
| <r2> |
88.911 |
| (<r2>)1/2 |
9.429 |
