return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-270.094638
Energy at 298.15K-270.101500
HF Energy-269.160652
Nuclear repulsion energy258.013263
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3233 3101 13.25      
2 A1 3212 3082 5.10      
3 A1 3196 3067 5.84      
4 A1 3174 3045 5.96      
5 A1 1624 1558 1.11      
6 A1 1515 1454 5.39      
7 A1 1498 1437 0.53      
8 A1 1292 1239 2.39      
9 A1 1180 1132 0.01      
10 A1 1041 998 1.39      
11 A1 983 943 0.13      
12 A1 833 799 0.24      
13 A1 527 505 0.24      
14 A2 971 932 0.00      
15 A2 840 806 0.00      
16 A2 520 499 0.00      
17 A2 387 371 0.00      
18 B1 965 926 0.43      
19 B1 890 853 1.90      
20 B1 742 712 72.14      
21 B1 700 671 8.96      
22 B1 640 614 13.31      
23 B1 469 450 10.65      
24 B1 199 191 1.06      
25 B2 3280 3147 7.69      
26 B2 3219 3089 34.38      
27 B2 3199 3070 0.86      
28 B2 1610 1544 0.58      
29 B2 1484 1423 5.48      
30 B2 1343 1289 0.49      
31 B2 1312 1259 0.42      
32 B2 1156 1109 0.05      
33 B2 1119 1074 2.24      
34 B2 977 937 1.54      
35 B2 619 594 0.01      
36 B2 355 341 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 25151.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 24130.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.18186 0.08905 0.05978

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.000
C2 0.000 1.227 0.254
C3 0.000 1.220 -1.139
C4 0.000 0.000 -1.850
C5 0.000 -1.220 -1.139
C6 0.000 -1.227 0.254
C7 0.000 0.000 2.414
H8 0.000 2.178 0.801
H9 0.000 2.170 -1.687
H10 0.000 0.000 -2.946
H11 0.000 -2.170 -1.687
H12 0.000 -2.178 0.801
H13 0.000 0.939 2.977
H14 0.000 -0.939 2.977

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.43582.46272.85032.46271.43581.41382.18663.45363.94563.45362.18662.18862.1886
C21.43581.39342.43612.81622.45392.48371.09662.15813.42713.91233.44822.73763.4792
C32.46271.39341.41252.44072.81623.75662.16341.09612.18013.43403.91274.12544.6480
C42.85032.43611.41251.41252.43614.26403.43092.17581.09532.17583.43094.91764.9176
C52.46272.81622.44071.41251.39343.75663.91273.43402.18011.09612.16344.64804.1254
C61.43582.45392.81622.43611.39342.48373.44823.91233.42712.15811.09663.47922.7376
C71.41382.48373.75664.26403.75662.48372.70984.63945.35944.63942.70981.09511.0951
H82.18661.09662.16343.43093.91273.44822.70982.48804.33355.00894.35512.50343.8012
H93.45362.15811.09612.17583.43403.91234.63942.48802.50834.33935.00894.82335.6051
H103.94563.42712.18011.09532.18013.42715.35944.33352.50832.50834.33355.99645.9964
H113.45363.91233.43402.17581.09612.15814.63945.00894.33932.50832.48805.60514.8233
H122.18663.44823.91273.43092.16341.09662.70984.35515.00894.33352.48803.80122.5034
H132.18862.73764.12544.91764.64803.47921.09512.50344.82335.99645.60513.80121.8787
H142.18863.47924.64804.91764.12542.73761.09513.80125.60515.99644.82332.50341.8787

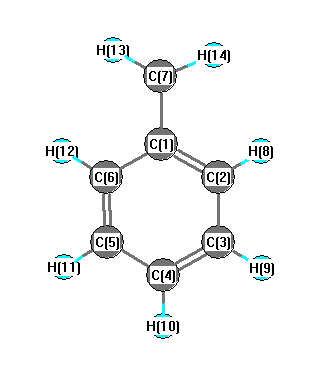
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.016 C1 C2 H8 118.807
C1 C6 C5 121.016 C1 C6 H12 118.807
C1 C7 H13 120.934 C1 C7 H14 120.934
C2 C1 C6 117.422 C2 C1 C7 121.289
C2 C3 C4 120.507 C2 C3 H9 119.721
C3 C2 H8 120.177 C3 C4 C5 119.532
C3 C4 H10 120.234 C4 C3 H9 119.773
C4 C5 C6 120.507 C4 C5 H11 119.773
C5 C1 C7 150.295 C5 C4 H10 120.234
C5 C6 H12 120.177 C6 C5 H11 119.721
H13 C7 H14 118.132
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability