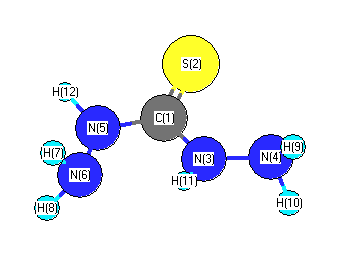
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3393 |
3398 |
0.77 |
|
|
|
| 2 |
A |
3387 |
3393 |
8.38 |
|
|
|
| 3 |
A |
3381 |
3386 |
1.08 |
|
|
|
| 4 |
A |
3331 |
3336 |
25.60 |
|
|
|
| 5 |
A |
3282 |
3288 |
12.04 |
|
|
|
| 6 |
A |
3270 |
3276 |
15.10 |
|
|
|
| 7 |
A |
1631 |
1633 |
28.07 |
|
|
|
| 8 |
A |
1611 |
1614 |
11.66 |
|
|
|
| 9 |
A |
1480 |
1482 |
85.70 |
|
|
|
| 10 |
A |
1431 |
1433 |
110.32 |
|
|
|
| 11 |
A |
1312 |
1314 |
37.24 |
|
|
|
| 12 |
A |
1297 |
1299 |
8.20 |
|
|
|
| 13 |
A |
1255 |
1257 |
15.91 |
|
|
|
| 14 |
A |
1210 |
1212 |
39.80 |
|
|
|
| 15 |
A |
1115 |
1117 |
75.58 |
|
|
|
| 16 |
A |
1030 |
1031 |
39.64 |
|
|
|
| 17 |
A |
888 |
890 |
90.35 |
|
|
|
| 18 |
A |
863 |
864 |
74.35 |
|
|
|
| 19 |
A |
748 |
749 |
19.87 |
|
|
|
| 20 |
A |
617 |
618 |
37.47 |
|
|
|
| 21 |
A |
614 |
615 |
18.69 |
|
|
|
| 22 |
A |
581 |
581 |
35.17 |
|
|
|
| 23 |
A |
513 |
514 |
93.55 |
|
|
|
| 24 |
A |
455 |
455 |
1.50 |
|
|
|
| 25 |
A |
316 |
317 |
31.83 |
|
|
|
| 26 |
A |
264 |
264 |
19.67 |
|
|
|
| 27 |
A |
223 |
224 |
5.21 |
|
|
|
| 28 |
A |
171 |
172 |
3.72 |
|
|
|
| 29 |
A |
118 |
119 |
9.99 |
|
|
|
| 30 |
A |
84 |
84 |
44.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19934.9 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 19966.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.113 |
|
|
|
| 2 |
S |
-0.314 |
|
|
|
| 3 |
N |
-0.098 |
|
|
|
| 4 |
N |
-0.024 |
|
|
|
| 5 |
N |
-0.101 |
|
|
|
| 6 |
N |
-0.081 |
|
|
|
| 7 |
H |
0.093 |
|
|
|
| 8 |
H |
0.095 |
|
|
|
| 9 |
H |
0.082 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
| 11 |
H |
0.068 |
|
|
|
| 12 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.139 |
3.576 |
-0.065 |
6.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.721 |
-7.082 |
2.052 |
| y |
-7.082 |
-37.900 |
-1.226 |
| z |
2.052 |
-1.226 |
-42.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.428 |
-7.082 |
2.052 |
| y |
-7.082 |
2.158 |
-1.226 |
| z |
2.052 |
-1.226 |
-4.586 |
|
| Polar |
|---|
| 3z2-r2 | -9.172 |
|---|
| x2-y2 | 0.180 |
|---|
| xy | -7.082 |
|---|
| xz | 2.052 |
|---|
| yz | -1.226 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.816 |
0.960 |
0.086 |
| y |
0.960 |
10.616 |
-0.084 |
| z |
0.086 |
-0.084 |
5.098 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |