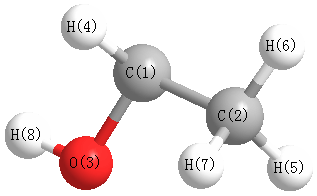
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3682 |
3661 |
40.67 |
|
|
|
| 2 |
A |
3062 |
3044 |
29.15 |
|
|
|
| 3 |
A |
3039 |
3021 |
29.64 |
|
|
|
| 4 |
A |
2986 |
2968 |
18.99 |
|
|
|
| 5 |
A |
2869 |
2852 |
61.83 |
|
|
|
| 6 |
A |
1419 |
1411 |
10.29 |
|
|
|
| 7 |
A |
1403 |
1394 |
9.20 |
|
|
|
| 8 |
A |
1376 |
1368 |
5.03 |
|
|
|
| 9 |
A |
1321 |
1313 |
0.65 |
|
|
|
| 10 |
A |
1233 |
1226 |
138.53 |
|
|
|
| 11 |
A |
1183 |
1176 |
18.30 |
|
|
|
| 12 |
A |
1028 |
1023 |
20.24 |
|
|
|
| 13 |
A |
971 |
965 |
3.59 |
|
|
|
| 14 |
A |
908 |
903 |
11.11 |
|
|
|
| 15 |
A |
550 |
547 |
16.86 |
|
|
|
| 16 |
A |
394 |
392 |
13.15 |
|
|
|
| 17 |
A |
366 |
364 |
113.05 |
|
|
|
| 18 |
A |
170 |
169 |
2.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13979.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 13898.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.075 |
|
|
|
| 2 |
C |
-0.046 |
|
|
|
| 3 |
O |
-0.160 |
|
|
|
| 4 |
H |
0.008 |
|
|
|
| 5 |
H |
0.046 |
|
|
|
| 6 |
H |
0.038 |
|
|
|
| 7 |
H |
0.037 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.718 |
1.013 |
0.299 |
1.277 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.205 |
-1.745 |
-0.187 |
| y |
-1.745 |
-19.306 |
0.520 |
| z |
-0.187 |
0.520 |
-20.305 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.600 |
-1.745 |
-0.187 |
| y |
-1.745 |
-1.552 |
0.520 |
| z |
-0.187 |
0.520 |
-3.049 |
|
| Polar |
|---|
| 3z2-r2 | -6.098 |
|---|
| x2-y2 | 4.101 |
|---|
| xy | -1.745 |
|---|
| xz | -0.187 |
|---|
| yz | 0.520 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.167 |
-0.233 |
0.085 |
| y |
-0.233 |
4.139 |
0.041 |
| z |
0.085 |
0.041 |
3.107 |
<r2> (average value of r
2) Å
2
| <r2> |
51.934 |
| (<r2>)1/2 |
7.206 |