Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.123185 |
| Energy at 298.15K | -235.136642 |
| HF Energy | -234.184705 |
| Nuclear repulsion energy | 248.257452 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3132 |
3005 |
105.54 |
|
|
|
| 2 |
A' |
3127 |
3000 |
22.66 |
|
|
|
| 3 |
A' |
3121 |
2994 |
19.52 |
|
|
|
| 4 |
A' |
3085 |
2960 |
21.36 |
|
|
|
| 5 |
A' |
3071 |
2946 |
14.14 |
|
|
|
| 6 |
A' |
3064 |
2939 |
8.91 |
|
|
|
| 7 |
A' |
3045 |
2922 |
30.18 |
|
|
|
| 8 |
A' |
3042 |
2919 |
27.18 |
|
|
|
| 9 |
A' |
1502 |
1441 |
5.39 |
|
|
|
| 10 |
A' |
1493 |
1432 |
0.95 |
|
|
|
| 11 |
A' |
1491 |
1431 |
5.12 |
|
|
|
| 12 |
A' |
1423 |
1365 |
2.21 |
|
|
|
| 13 |
A' |
1421 |
1364 |
3.58 |
|
|
|
| 14 |
A' |
1386 |
1330 |
1.18 |
|
|
|
| 15 |
A' |
1374 |
1318 |
8.40 |
|
|
|
| 16 |
A' |
1287 |
1235 |
1.30 |
|
|
|
| 17 |
A' |
1255 |
1204 |
0.43 |
|
|
|
| 18 |
A' |
1208 |
1159 |
0.15 |
|
|
|
| 19 |
A' |
1157 |
1110 |
0.39 |
|
|
|
| 20 |
A' |
1060 |
1017 |
0.46 |
|
|
|
| 21 |
A' |
964 |
925 |
0.35 |
|
|
|
| 22 |
A' |
961 |
922 |
0.71 |
|
|
|
| 23 |
A' |
873 |
837 |
0.60 |
|
|
|
| 24 |
A' |
814 |
781 |
0.63 |
|
|
|
| 25 |
A' |
654 |
627 |
1.45 |
|
|
|
| 26 |
A' |
433 |
415 |
0.04 |
|
|
|
| 27 |
A' |
327 |
314 |
0.06 |
|
|
|
| 28 |
A' |
117 |
112 |
0.01 |
|
|
|
| 29 |
A" |
3128 |
3001 |
36.08 |
|
|
|
| 30 |
A" |
3125 |
2999 |
26.63 |
|
|
|
| 31 |
A" |
3121 |
2995 |
9.19 |
|
|
|
| 32 |
A" |
3062 |
2938 |
64.86 |
|
|
|
| 33 |
A" |
1495 |
1434 |
2.66 |
|
|
|
| 34 |
A" |
1493 |
1433 |
0.33 |
|
|
|
| 35 |
A" |
1470 |
1411 |
2.59 |
|
|
|
| 36 |
A" |
1305 |
1252 |
0.70 |
|
|
|
| 37 |
A" |
1278 |
1226 |
0.00 |
|
|
|
| 38 |
A" |
1241 |
1190 |
0.06 |
|
|
|
| 39 |
A" |
1146 |
1099 |
0.15 |
|
|
|
| 40 |
A" |
1134 |
1088 |
0.05 |
|
|
|
| 41 |
A" |
996 |
956 |
1.78 |
|
|
|
| 42 |
A" |
919 |
882 |
0.01 |
|
|
|
| 43 |
A" |
911 |
874 |
0.13 |
|
|
|
| 44 |
A" |
811 |
778 |
0.11 |
|
|
|
| 45 |
A" |
371 |
356 |
0.00 |
|
|
|
| 46 |
A" |
263 |
252 |
0.05 |
|
|
|
| 47 |
A" |
252 |
242 |
0.00 |
|
|
|
| 48 |
A" |
236 |
226 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36820.8 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 35325.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.262 |
0.089 |
1.088 |
| C2 |
0.262 |
0.089 |
-1.088 |
| C3 |
-0.668 |
0.710 |
0.000 |
| C4 |
0.862 |
-0.858 |
0.000 |
| C5 |
-0.881 |
2.222 |
0.000 |
| C6 |
0.262 |
-2.267 |
0.000 |
| H7 |
1.005 |
0.822 |
1.456 |
| H8 |
-0.223 |
-0.390 |
1.959 |
| H9 |
1.005 |
0.822 |
-1.456 |
| H10 |
-0.223 |
-0.390 |
-1.959 |
| H11 |
-1.649 |
0.198 |
0.000 |
| H12 |
1.965 |
-0.926 |
0.000 |
| H13 |
0.089 |
2.755 |
0.000 |
| H14 |
-0.844 |
-2.230 |
0.000 |
| H15 |
-1.447 |
2.545 |
-0.895 |
| H16 |
-1.447 |
2.545 |
0.895 |
| H17 |
0.582 |
-2.832 |
-0.896 |
| H18 |
0.582 |
-2.832 |
0.896 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1762 | 1.5602 | 1.5622 | 2.6534 | 2.5953 | 1.1061 | 1.1060 | 2.7499 | 3.1222 | 2.2022 | 2.2614 | 2.8842 | 2.7905 | 3.5895 | 2.9982 | 3.5460 | 2.9452 |
C2 | 2.1762 | | 1.5602 | 1.5622 | 2.6534 | 2.5953 | 2.7499 | 3.1222 | 1.1061 | 1.1060 | 2.2022 | 2.2614 | 2.8842 | 2.7905 | 2.9982 | 3.5895 | 2.9452 | 3.5460 | C3 | 1.5602 | 1.5602 | | 2.1904 | 1.5270 | 3.1188 | 2.2203 | 2.2901 | 2.2203 | 2.2901 | 1.1068 | 3.0996 | 2.1803 | 2.9451 | 2.1852 | 2.1852 | 3.8617 | 3.8617 | C4 | 1.5622 | 1.5622 | 2.1904 | | 3.5390 | 1.5311 | 2.2280 | 2.2874 | 2.2280 | 2.2874 | 2.7241 | 1.1056 | 3.6943 | 2.1893 | 4.2084 | 4.2084 | 2.1860 | 2.1860 | C5 | 2.6534 | 2.6534 | 1.5270 | 3.5390 | | 4.6323 | 2.7635 | 3.3306 | 2.7635 | 3.3306 | 2.1643 | 4.2439 | 1.1072 | 4.4520 | 1.1069 | 1.1069 | 5.3376 | 5.3376 | C6 | 2.5953 | 2.5953 | 3.1188 | 1.5311 | 4.6323 | | 3.4948 | 2.7559 | 3.4948 | 2.7559 | 3.1196 | 2.1677 | 5.0245 | 1.1074 | 5.1842 | 5.1842 | 1.1066 | 1.1066 | H7 | 1.1061 | 2.7499 | 2.2203 | 2.2280 | 2.7635 | 3.4948 | | 1.7967 | 2.9127 | 3.8260 | 3.0906 | 2.4695 | 2.5871 | 3.8541 | 3.8087 | 3.0484 | 4.3664 | 3.7211 | H8 | 1.1060 | 3.1222 | 2.2901 | 2.2874 | 3.3306 | 2.7559 | 1.7967 | | 3.8260 | 3.9175 | 2.4935 | 2.9852 | 3.7179 | 2.7583 | 4.2727 | 3.3531 | 3.8422 | 2.7826 | H9 | 2.7499 | 1.1061 | 2.2203 | 2.2280 | 2.7635 | 3.4948 | 2.9127 | 3.8260 | | 1.7967 | 3.0906 | 2.4695 | 2.5871 | 3.8541 | 3.0484 | 3.8087 | 3.7211 | 4.3664 | H10 | 3.1222 | 1.1060 | 2.2901 | 2.2874 | 3.3306 | 2.7559 | 3.8260 | 3.9175 | 1.7967 | | 2.4935 | 2.9852 | 3.7179 | 2.7583 | 3.3531 | 4.2727 | 2.7826 | 3.8422 | H11 | 2.2022 | 2.2022 | 1.1068 | 2.7241 | 2.1643 | 3.1196 | 3.0906 | 2.4935 | 3.0906 | 2.4935 | | 3.7852 | 3.0913 | 2.5581 | 2.5194 | 2.5194 | 3.8687 | 3.8687 | H12 | 2.2614 | 2.2614 | 3.0996 | 1.1056 | 4.2439 | 2.1677 | 2.4695 | 2.9852 | 2.4695 | 2.9852 | 3.7852 | | 4.1308 | 3.0976 | 4.9486 | 4.9486 | 2.5200 | 2.5200 | H13 | 2.8842 | 2.8842 | 2.1803 | 3.6943 | 1.1072 | 5.0245 | 2.5871 | 3.7179 | 2.5871 | 3.7179 | 3.0913 | 4.1308 | | 5.0712 | 1.7903 | 1.7903 | 5.6797 | 5.6797 | H14 | 2.7905 | 2.7905 | 2.9451 | 2.1893 | 4.4520 | 1.1074 | 3.8541 | 2.7583 | 3.8541 | 2.7583 | 2.5581 | 3.0976 | 5.0712 | | 4.8950 | 4.8950 | 1.7892 | 1.7892 | H15 | 3.5895 | 2.9982 | 2.1852 | 4.2084 | 1.1069 | 5.1842 | 3.8087 | 4.2727 | 3.0484 | 3.3531 | 2.5194 | 4.9486 | 1.7903 | 4.8950 | | 1.7901 | 5.7473 | 6.0198 | H16 | 2.9982 | 3.5895 | 2.1852 | 4.2084 | 1.1069 | 5.1842 | 3.0484 | 3.3531 | 3.8087 | 4.2727 | 2.5194 | 4.9486 | 1.7903 | 4.8950 | 1.7901 | | 6.0198 | 5.7473 | H17 | 3.5460 | 2.9452 | 3.8617 | 2.1860 | 5.3376 | 1.1066 | 4.3664 | 3.8422 | 3.7211 | 2.7826 | 3.8687 | 2.5200 | 5.6797 | 1.7892 | 5.7473 | 6.0198 | | 1.7918 | H18 | 2.9452 | 3.5460 | 3.8617 | 2.1860 | 5.3376 | 1.1066 | 3.7211 | 2.7826 | 4.3664 | 3.8422 | 3.8687 | 2.5200 | 5.6797 | 1.7892 | 6.0198 | 5.7473 | 1.7918 | |
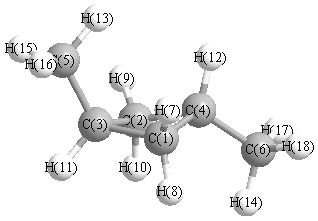 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.439 |
|
C1 |
C3 |
C5 |
118.515 |
| C1 |
C3 |
H11 |
110.162 |
|
C1 |
C4 |
C2 |
88.297 |
| C1 |
C4 |
C6 |
114.064 |
|
C1 |
C4 |
H12 |
114.838 |
| C2 |
C3 |
C5 |
118.515 |
|
C2 |
C3 |
H11 |
110.162 |
| C2 |
C4 |
C6 |
114.064 |
|
C2 |
C4 |
H12 |
114.838 |
| C3 |
C1 |
C4 |
89.095 |
|
C3 |
C1 |
H7 |
111.622 |
| C3 |
C1 |
H8 |
117.383 |
|
C3 |
C2 |
C4 |
89.095 |
| C3 |
C2 |
H9 |
111.622 |
|
C3 |
C2 |
H10 |
117.383 |
| C3 |
C5 |
H13 |
110.713 |
|
C3 |
C5 |
H15 |
111.118 |
| C3 |
C5 |
H16 |
111.118 |
|
C4 |
C1 |
H7 |
112.098 |
| C4 |
C1 |
H8 |
116.996 |
|
C4 |
C2 |
H9 |
112.098 |
| C4 |
C2 |
H10 |
116.996 |
|
C4 |
C6 |
H14 |
111.126 |
| C4 |
C6 |
H17 |
110.912 |
|
C4 |
C6 |
H18 |
110.912 |
| C5 |
C3 |
H11 |
109.486 |
|
C6 |
C4 |
H12 |
109.539 |
| H7 |
C1 |
H8 |
108.625 |
|
H9 |
C2 |
H10 |
108.625 |
| H13 |
C5 |
H15 |
107.918 |
|
H13 |
C5 |
H16 |
107.918 |
| H14 |
C6 |
H17 |
107.822 |
|
H14 |
C6 |
H18 |
107.822 |
| H15 |
C5 |
H16 |
107.918 |
|
H17 |
C6 |
H18 |
108.110 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
