Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.118088 |
| Energy at 298.15K | -235.131558 |
| HF Energy | -234.184858 |
| Nuclear repulsion energy | 248.320951 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3144 |
3008 |
98.65 |
|
|
|
| 2 |
A' |
3139 |
3003 |
21.25 |
|
|
|
| 3 |
A' |
3134 |
2998 |
19.59 |
|
|
|
| 4 |
A' |
3096 |
2962 |
21.62 |
|
|
|
| 5 |
A' |
3082 |
2949 |
14.06 |
|
|
|
| 6 |
A' |
3075 |
2942 |
7.51 |
|
|
|
| 7 |
A' |
3055 |
2923 |
28.28 |
|
|
|
| 8 |
A' |
3052 |
2920 |
26.38 |
|
|
|
| 9 |
A' |
1505 |
1440 |
5.22 |
|
|
|
| 10 |
A' |
1495 |
1431 |
1.31 |
|
|
|
| 11 |
A' |
1494 |
1429 |
5.23 |
|
|
|
| 12 |
A' |
1427 |
1365 |
2.17 |
|
|
|
| 13 |
A' |
1425 |
1363 |
4.07 |
|
|
|
| 14 |
A' |
1390 |
1330 |
1.24 |
|
|
|
| 15 |
A' |
1378 |
1318 |
7.88 |
|
|
|
| 16 |
A' |
1292 |
1236 |
1.29 |
|
|
|
| 17 |
A' |
1258 |
1203 |
0.45 |
|
|
|
| 18 |
A' |
1210 |
1158 |
0.14 |
|
|
|
| 19 |
A' |
1160 |
1110 |
0.35 |
|
|
|
| 20 |
A' |
1063 |
1017 |
0.40 |
|
|
|
| 21 |
A' |
967 |
925 |
0.38 |
|
|
|
| 22 |
A' |
963 |
921 |
0.71 |
|
|
|
| 23 |
A' |
876 |
838 |
0.64 |
|
|
|
| 24 |
A' |
817 |
782 |
0.66 |
|
|
|
| 25 |
A' |
655 |
627 |
1.45 |
|
|
|
| 26 |
A' |
434 |
415 |
0.04 |
|
|
|
| 27 |
A' |
328 |
313 |
0.06 |
|
|
|
| 28 |
A' |
116 |
111 |
0.01 |
|
|
|
| 29 |
A" |
3141 |
3005 |
33.25 |
|
|
|
| 30 |
A" |
3138 |
3003 |
24.91 |
|
|
|
| 31 |
A" |
3134 |
2998 |
9.31 |
|
|
|
| 32 |
A" |
3072 |
2939 |
62.17 |
|
|
|
| 33 |
A" |
1498 |
1433 |
2.79 |
|
|
|
| 34 |
A" |
1496 |
1431 |
0.31 |
|
|
|
| 35 |
A" |
1473 |
1410 |
2.94 |
|
|
|
| 36 |
A" |
1309 |
1252 |
0.64 |
|
|
|
| 37 |
A" |
1282 |
1226 |
0.00 |
|
|
|
| 38 |
A" |
1245 |
1191 |
0.06 |
|
|
|
| 39 |
A" |
1149 |
1099 |
0.17 |
|
|
|
| 40 |
A" |
1137 |
1088 |
0.06 |
|
|
|
| 41 |
A" |
999 |
956 |
1.88 |
|
|
|
| 42 |
A" |
923 |
883 |
0.00 |
|
|
|
| 43 |
A" |
915 |
875 |
0.10 |
|
|
|
| 44 |
A" |
813 |
777 |
0.11 |
|
|
|
| 45 |
A" |
373 |
356 |
0.00 |
|
|
|
| 46 |
A" |
263 |
252 |
0.05 |
|
|
|
| 47 |
A" |
253 |
242 |
0.00 |
|
|
|
| 48 |
A" |
236 |
226 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36937.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35338.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.262 |
0.089 |
1.088 |
| C2 |
0.262 |
0.089 |
-1.088 |
| C3 |
-0.667 |
0.710 |
0.000 |
| C4 |
0.861 |
-0.858 |
0.000 |
| C5 |
-0.880 |
2.222 |
0.000 |
| C6 |
0.262 |
-2.267 |
0.000 |
| H7 |
1.004 |
0.821 |
1.456 |
| H8 |
-0.223 |
-0.390 |
1.958 |
| H9 |
1.004 |
0.821 |
-1.456 |
| H10 |
-0.223 |
-0.390 |
-1.958 |
| H11 |
-1.648 |
0.199 |
0.000 |
| H12 |
1.964 |
-0.925 |
0.000 |
| H13 |
0.090 |
2.754 |
0.000 |
| H14 |
-0.844 |
-2.230 |
0.000 |
| H15 |
-1.446 |
2.545 |
-0.895 |
| H16 |
-1.446 |
2.545 |
0.895 |
| H17 |
0.582 |
-2.832 |
-0.896 |
| H18 |
0.582 |
-2.832 |
0.896 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1754 | 1.5596 | 1.5616 | 2.6529 | 2.5947 | 1.1057 | 1.1055 | 2.7489 | 3.1210 | 2.2013 | 2.2605 | 2.8838 | 2.7901 | 3.5886 | 2.9975 | 3.5450 | 2.9446 |
C2 | 2.1754 | | 1.5596 | 1.5616 | 2.6529 | 2.5947 | 2.7489 | 3.1210 | 1.1057 | 1.1055 | 2.2013 | 2.2605 | 2.8838 | 2.7901 | 2.9975 | 3.5886 | 2.9446 | 3.5450 | C3 | 1.5596 | 1.5596 | | 2.1897 | 1.5269 | 3.1185 | 2.2193 | 2.2893 | 2.2193 | 2.2893 | 1.1064 | 3.0984 | 2.1799 | 2.9453 | 2.1847 | 2.1847 | 3.8610 | 3.8610 | C4 | 1.5616 | 1.5616 | 2.1897 | | 3.5382 | 1.5309 | 2.2271 | 2.2865 | 2.2271 | 2.2865 | 2.7232 | 1.1052 | 3.6936 | 2.1889 | 4.2073 | 4.2073 | 2.1855 | 2.1855 | C5 | 2.6529 | 2.6529 | 1.5269 | 3.5382 | | 4.6318 | 2.7628 | 3.3299 | 2.7628 | 3.3299 | 2.1639 | 4.2426 | 1.1067 | 4.4520 | 1.1065 | 1.1065 | 5.3368 | 5.3368 | C6 | 2.5947 | 2.5947 | 3.1185 | 1.5309 | 4.6318 | | 3.4938 | 2.7552 | 3.4938 | 2.7552 | 3.1192 | 2.1672 | 5.0240 | 1.1070 | 5.1834 | 5.1834 | 1.1062 | 1.1062 | H7 | 1.1057 | 2.7489 | 2.2193 | 2.2271 | 2.7628 | 3.4938 | | 1.7960 | 2.9117 | 3.8246 | 3.0893 | 2.4684 | 2.5868 | 3.8532 | 3.8076 | 3.0475 | 4.3650 | 3.7200 | H8 | 1.1055 | 3.1210 | 2.2893 | 2.2865 | 3.3299 | 2.7552 | 1.7960 | | 3.8246 | 3.9159 | 2.4924 | 2.9841 | 3.7173 | 2.7577 | 4.2715 | 3.3524 | 3.8410 | 2.7820 | H9 | 2.7489 | 1.1057 | 2.2193 | 2.2271 | 2.7628 | 3.4938 | 2.9117 | 3.8246 | | 1.7960 | 3.0893 | 2.4684 | 2.5868 | 3.8532 | 3.0475 | 3.8076 | 3.7200 | 4.3650 | H10 | 3.1210 | 1.1055 | 2.2893 | 2.2865 | 3.3299 | 2.7552 | 3.8246 | 3.9159 | 1.7960 | | 2.4924 | 2.9841 | 3.7173 | 2.7577 | 3.3524 | 4.2715 | 2.7820 | 3.8410 | H11 | 2.2013 | 2.2013 | 1.1064 | 2.7232 | 2.1639 | 3.1192 | 3.0893 | 2.4924 | 3.0893 | 2.4924 | | 3.7838 | 3.0906 | 2.5585 | 2.5188 | 2.5188 | 3.8681 | 3.8681 | H12 | 2.2605 | 2.2605 | 3.0984 | 1.1052 | 4.2426 | 2.1672 | 2.4684 | 2.9841 | 2.4684 | 2.9841 | 3.7838 | | 4.1297 | 3.0968 | 4.9470 | 4.9470 | 2.5194 | 2.5194 | H13 | 2.8838 | 2.8838 | 2.1799 | 3.6936 | 1.1067 | 5.0240 | 2.5868 | 3.7173 | 2.5868 | 3.7173 | 3.0906 | 4.1297 | | 5.0710 | 1.7896 | 1.7896 | 5.6788 | 5.6788 | H14 | 2.7901 | 2.7901 | 2.9453 | 2.1889 | 4.4520 | 1.1070 | 3.8532 | 2.7577 | 3.8532 | 2.7577 | 2.5585 | 3.0968 | 5.0710 | | 4.8949 | 4.8949 | 1.7885 | 1.7885 | H15 | 3.5886 | 2.9975 | 2.1847 | 4.2073 | 1.1065 | 5.1834 | 3.8076 | 4.2715 | 3.0475 | 3.3524 | 2.5188 | 4.9470 | 1.7896 | 4.8949 | | 1.7894 | 5.7463 | 6.0187 | H16 | 2.9975 | 3.5886 | 2.1847 | 4.2073 | 1.1065 | 5.1834 | 3.0475 | 3.3524 | 3.8076 | 4.2715 | 2.5188 | 4.9470 | 1.7896 | 4.8949 | 1.7894 | | 6.0187 | 5.7463 | H17 | 3.5450 | 2.9446 | 3.8610 | 2.1855 | 5.3368 | 1.1062 | 4.3650 | 3.8410 | 3.7200 | 2.7820 | 3.8681 | 2.5194 | 5.6788 | 1.7885 | 5.7463 | 6.0187 | | 1.7911 | H18 | 2.9446 | 3.5450 | 3.8610 | 2.1855 | 5.3368 | 1.1062 | 3.7200 | 2.7820 | 4.3650 | 3.8410 | 3.8681 | 2.5194 | 5.6788 | 1.7885 | 6.0187 | 5.7463 | 1.7911 | |
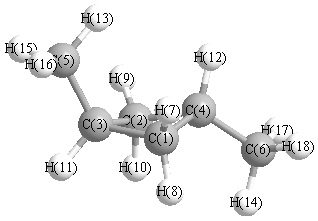 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.438 |
|
C1 |
C3 |
C5 |
118.524 |
| C1 |
C3 |
H11 |
110.152 |
|
C1 |
C4 |
C2 |
88.296 |
| C1 |
C4 |
C6 |
114.070 |
|
C1 |
C4 |
H12 |
114.835 |
| C2 |
C3 |
C5 |
118.524 |
|
C2 |
C3 |
H11 |
110.152 |
| C2 |
C4 |
C6 |
114.070 |
|
C2 |
C4 |
H12 |
114.835 |
| C3 |
C1 |
C4 |
89.101 |
|
C3 |
C1 |
H7 |
111.615 |
| C3 |
C1 |
H8 |
117.388 |
|
C3 |
C2 |
C4 |
89.101 |
| C3 |
C2 |
H9 |
111.615 |
|
C3 |
C2 |
H10 |
117.388 |
| C3 |
C5 |
H13 |
110.720 |
|
C3 |
C5 |
H15 |
111.117 |
| C3 |
C5 |
H16 |
111.117 |
|
C4 |
C1 |
H7 |
112.093 |
| C4 |
C1 |
H8 |
116.995 |
|
C4 |
C2 |
H9 |
112.093 |
| C4 |
C2 |
H10 |
116.995 |
|
C4 |
C6 |
H14 |
111.135 |
| C4 |
C6 |
H17 |
110.911 |
|
C4 |
C6 |
H18 |
110.911 |
| C5 |
C3 |
H11 |
109.486 |
|
C6 |
C4 |
H12 |
109.534 |
| H7 |
C1 |
H8 |
108.626 |
|
H9 |
C2 |
H10 |
108.626 |
| H13 |
C5 |
H15 |
107.916 |
|
H13 |
C5 |
H16 |
107.916 |
| H14 |
C6 |
H17 |
107.819 |
|
H14 |
C6 |
H18 |
107.819 |
| H15 |
C5 |
H16 |
107.917 |
|
H17 |
C6 |
H18 |
108.108 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
