Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.509716 |
| Energy at 298.15K | -235.522904 |
| HF Energy | -235.509716 |
| Nuclear repulsion energy | 247.810027 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3039 |
3021 |
92.30 |
|
|
|
| 2 |
A' |
3035 |
3018 |
18.61 |
|
|
|
| 3 |
A' |
3029 |
3011 |
24.73 |
|
|
|
| 4 |
A' |
2986 |
2968 |
18.99 |
|
|
|
| 5 |
A' |
2973 |
2956 |
11.40 |
|
|
|
| 6 |
A' |
2969 |
2952 |
10.67 |
|
|
|
| 7 |
A' |
2957 |
2940 |
31.66 |
|
|
|
| 8 |
A' |
2954 |
2937 |
29.17 |
|
|
|
| 9 |
A' |
1423 |
1415 |
7.67 |
|
|
|
| 10 |
A' |
1416 |
1408 |
1.01 |
|
|
|
| 11 |
A' |
1414 |
1406 |
4.09 |
|
|
|
| 12 |
A' |
1345 |
1337 |
2.35 |
|
|
|
| 13 |
A' |
1345 |
1337 |
3.57 |
|
|
|
| 14 |
A' |
1318 |
1310 |
0.17 |
|
|
|
| 15 |
A' |
1302 |
1295 |
13.00 |
|
|
|
| 16 |
A' |
1223 |
1216 |
1.19 |
|
|
|
| 17 |
A' |
1204 |
1197 |
0.33 |
|
|
|
| 18 |
A' |
1154 |
1147 |
0.38 |
|
|
|
| 19 |
A' |
1115 |
1108 |
0.35 |
|
|
|
| 20 |
A' |
1024 |
1018 |
1.39 |
|
|
|
| 21 |
A' |
924 |
919 |
1.12 |
|
|
|
| 22 |
A' |
912 |
907 |
0.67 |
|
|
|
| 23 |
A' |
845 |
840 |
1.44 |
|
|
|
| 24 |
A' |
789 |
784 |
0.91 |
|
|
|
| 25 |
A' |
629 |
625 |
1.82 |
|
|
|
| 26 |
A' |
418 |
416 |
0.18 |
|
|
|
| 27 |
A' |
317 |
316 |
0.08 |
|
|
|
| 28 |
A' |
92 |
92 |
0.01 |
|
|
|
| 29 |
A" |
3039 |
3022 |
28.32 |
|
|
|
| 30 |
A" |
3037 |
3019 |
22.12 |
|
|
|
| 31 |
A" |
3029 |
3011 |
8.25 |
|
|
|
| 32 |
A" |
2969 |
2952 |
72.25 |
|
|
|
| 33 |
A" |
1419 |
1411 |
2.83 |
|
|
|
| 34 |
A" |
1416 |
1408 |
1.22 |
|
|
|
| 35 |
A" |
1390 |
1382 |
3.42 |
|
|
|
| 36 |
A" |
1236 |
1229 |
1.28 |
|
|
|
| 37 |
A" |
1215 |
1208 |
0.00 |
|
|
|
| 38 |
A" |
1177 |
1170 |
0.15 |
|
|
|
| 39 |
A" |
1098 |
1091 |
0.37 |
|
|
|
| 40 |
A" |
1086 |
1080 |
0.03 |
|
|
|
| 41 |
A" |
961 |
956 |
3.15 |
|
|
|
| 42 |
A" |
889 |
883 |
0.00 |
|
|
|
| 43 |
A" |
878 |
873 |
0.02 |
|
|
|
| 44 |
A" |
777 |
773 |
0.25 |
|
|
|
| 45 |
A" |
363 |
361 |
0.01 |
|
|
|
| 46 |
A" |
259 |
257 |
0.07 |
|
|
|
| 47 |
A" |
245 |
244 |
0.03 |
|
|
|
| 48 |
A" |
230 |
228 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35430.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 35224.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.251 |
0.081 |
1.087 |
| C2 |
0.251 |
0.081 |
-1.087 |
| C3 |
-0.660 |
0.726 |
0.000 |
| C4 |
0.846 |
-0.868 |
0.000 |
| C5 |
-0.843 |
2.240 |
0.000 |
| C6 |
0.251 |
-2.277 |
0.000 |
| H7 |
1.003 |
0.803 |
1.466 |
| H8 |
-0.246 |
-0.396 |
1.956 |
| H9 |
1.003 |
0.803 |
-1.466 |
| H10 |
-0.246 |
-0.396 |
-1.956 |
| H11 |
-1.656 |
0.236 |
0.000 |
| H12 |
1.952 |
-0.937 |
0.000 |
| H13 |
0.139 |
2.758 |
0.000 |
| H14 |
-0.858 |
-2.249 |
0.000 |
| H15 |
-1.404 |
2.579 |
-0.896 |
| H16 |
-1.404 |
2.579 |
0.896 |
| H17 |
0.573 |
-2.845 |
-0.897 |
| H18 |
0.573 |
-2.845 |
0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1741 | 1.5583 | 1.5607 | 2.6534 | 2.5968 | 1.1088 | 1.1090 | 2.7573 | 3.1199 | 2.2003 | 2.2611 | 2.8910 | 2.8010 | 3.5928 | 3.0023 | 3.5503 | 2.9507 |
C2 | 2.1741 | | 1.5583 | 1.5607 | 2.6534 | 2.5968 | 2.7573 | 3.1199 | 1.1088 | 1.1090 | 2.2003 | 2.2611 | 2.8910 | 2.8010 | 3.0023 | 3.5928 | 2.9507 | 3.5503 | C3 | 1.5583 | 1.5583 | | 2.1929 | 1.5252 | 3.1384 | 2.2183 | 2.2926 | 2.2183 | 2.2926 | 1.1095 | 3.0971 | 2.1831 | 2.9823 | 2.1882 | 2.1882 | 3.8835 | 3.8835 | C4 | 1.5607 | 1.5607 | 2.1929 | | 3.5373 | 1.5297 | 2.2280 | 2.2890 | 2.2280 | 2.2890 | 2.7341 | 1.1088 | 3.6938 | 2.1941 | 4.2122 | 4.2122 | 2.1887 | 2.1887 | C5 | 2.6534 | 2.6534 | 1.5252 | 3.5373 | | 4.6480 | 2.7610 | 3.3365 | 2.7610 | 3.3365 | 2.1628 | 4.2321 | 1.1100 | 4.4899 | 1.1095 | 1.1095 | 5.3549 | 5.3549 | C6 | 2.5968 | 2.5968 | 3.1384 | 1.5297 | 4.6480 | | 3.4928 | 2.7587 | 3.4928 | 2.7587 | 3.1544 | 2.1657 | 5.0361 | 1.1099 | 5.2079 | 5.2079 | 1.1093 | 1.1093 | H7 | 1.1088 | 2.7573 | 2.2183 | 2.2280 | 2.7610 | 3.4928 | | 1.7993 | 2.9315 | 3.8346 | 3.0884 | 2.4651 | 2.5917 | 3.8640 | 3.8109 | 3.0449 | 4.3677 | 3.7176 | H8 | 1.1090 | 3.1199 | 2.2926 | 2.2890 | 3.3365 | 2.7587 | 1.7993 | | 3.8346 | 3.9116 | 2.4923 | 2.9918 | 3.7310 | 2.7633 | 4.2805 | 3.3638 | 3.8479 | 2.7915 | H9 | 2.7573 | 1.1088 | 2.2183 | 2.2280 | 2.7610 | 3.4928 | 2.9315 | 3.8346 | | 1.7993 | 3.0884 | 2.4651 | 2.5917 | 3.8640 | 3.0449 | 3.8109 | 3.7176 | 4.3677 | H10 | 3.1199 | 1.1090 | 2.2926 | 2.2890 | 3.3365 | 2.7587 | 3.8346 | 3.9116 | 1.7993 | | 2.4923 | 2.9918 | 3.7310 | 2.7633 | 3.3638 | 4.2805 | 2.7915 | 3.8479 | H11 | 2.2003 | 2.2003 | 1.1095 | 2.7341 | 2.1628 | 3.1544 | 3.0884 | 2.4923 | 3.0884 | 2.4923 | | 3.7939 | 3.0951 | 2.6102 | 2.5212 | 2.5212 | 3.9070 | 3.9070 | H12 | 2.2611 | 2.2611 | 3.0971 | 1.1088 | 4.2321 | 2.1657 | 2.4651 | 2.9918 | 2.4651 | 2.9918 | 3.7939 | | 4.1159 | 3.1022 | 4.9424 | 4.9424 | 2.5200 | 2.5200 | H13 | 2.8910 | 2.8910 | 2.1831 | 3.6938 | 1.1100 | 5.0361 | 2.5917 | 3.7310 | 2.5917 | 3.7310 | 3.0951 | 4.1159 | | 5.1057 | 1.7925 | 1.7925 | 5.6913 | 5.6913 | H14 | 2.8010 | 2.8010 | 2.9823 | 2.1941 | 4.4899 | 1.1099 | 3.8640 | 2.7633 | 3.8640 | 2.7633 | 2.6102 | 3.1022 | 5.1057 | | 4.9411 | 4.9411 | 1.7908 | 1.7908 | H15 | 3.5928 | 3.0023 | 2.1882 | 4.2122 | 1.1095 | 5.2079 | 3.8109 | 4.2805 | 3.0449 | 3.3638 | 2.5212 | 4.9424 | 1.7925 | 4.9411 | | 1.7914 | 5.7735 | 6.0452 | H16 | 3.0023 | 3.5928 | 2.1882 | 4.2122 | 1.1095 | 5.2079 | 3.0449 | 3.3638 | 3.8109 | 4.2805 | 2.5212 | 4.9424 | 1.7925 | 4.9411 | 1.7914 | | 6.0452 | 5.7735 | H17 | 3.5503 | 2.9507 | 3.8835 | 2.1887 | 5.3549 | 1.1093 | 4.3677 | 3.8479 | 3.7176 | 2.7915 | 3.9070 | 2.5200 | 5.6913 | 1.7908 | 5.7735 | 6.0452 | | 1.7931 | H18 | 2.9507 | 3.5503 | 3.8835 | 2.1887 | 5.3549 | 1.1093 | 3.7176 | 2.7915 | 4.3677 | 3.8479 | 3.9070 | 2.5200 | 5.6913 | 1.7908 | 6.0452 | 5.7735 | 1.7931 | |
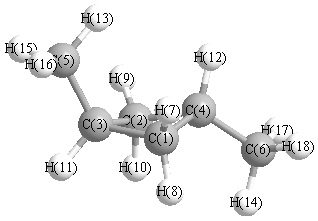 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.468 |
|
C1 |
C3 |
C5 |
118.751 |
| C1 |
C3 |
H11 |
109.990 |
|
C1 |
C4 |
C2 |
88.294 |
| C1 |
C4 |
C6 |
114.336 |
|
C1 |
C4 |
H12 |
114.725 |
| C2 |
C3 |
C5 |
118.751 |
|
C2 |
C3 |
H11 |
109.990 |
| C2 |
C4 |
C6 |
114.336 |
|
C2 |
C4 |
H12 |
114.725 |
| C3 |
C1 |
C4 |
89.347 |
|
C3 |
C1 |
H7 |
111.442 |
| C3 |
C1 |
H8 |
117.543 |
|
C3 |
C2 |
C4 |
89.347 |
| C3 |
C2 |
H9 |
111.442 |
|
C3 |
C2 |
H10 |
117.543 |
| C3 |
C5 |
H13 |
110.901 |
|
C3 |
C5 |
H15 |
111.327 |
| C3 |
C5 |
H16 |
111.327 |
|
C4 |
C1 |
H7 |
112.040 |
| C4 |
C1 |
H8 |
117.045 |
|
C4 |
C2 |
H9 |
112.040 |
| C4 |
C2 |
H10 |
117.045 |
|
C4 |
C6 |
H14 |
111.459 |
| C4 |
C6 |
H17 |
111.073 |
|
C4 |
C6 |
H18 |
111.073 |
| C5 |
C3 |
H11 |
109.334 |
|
C6 |
C4 |
H12 |
109.296 |
| H7 |
C1 |
H8 |
108.456 |
|
H9 |
C2 |
H10 |
108.456 |
| H13 |
C5 |
H15 |
107.727 |
|
H13 |
C5 |
H16 |
107.727 |
| H14 |
C6 |
H17 |
107.606 |
|
H14 |
C6 |
H18 |
107.606 |
| H15 |
C5 |
H16 |
107.658 |
|
H17 |
C6 |
H18 |
107.849 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.075 |
|
|
|
| 2 |
C |
0.075 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.163 |
|
|
|
| 5 |
C |
0.002 |
|
|
|
| 6 |
C |
-0.004 |
|
|
|
| 7 |
H |
0.006 |
|
|
|
| 8 |
H |
0.006 |
|
|
|
| 9 |
H |
0.006 |
|
|
|
| 10 |
H |
0.006 |
|
|
|
| 11 |
H |
-0.004 |
|
|
|
| 12 |
H |
-0.016 |
|
|
|
| 13 |
H |
0.023 |
|
|
|
| 14 |
H |
0.023 |
|
|
|
| 15 |
H |
0.030 |
|
|
|
| 16 |
H |
0.030 |
|
|
|
| 17 |
H |
0.030 |
|
|
|
| 18 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.039 |
0.003 |
0.000 |
0.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.756 |
0.233 |
0.000 |
| y |
0.233 |
-40.580 |
0.000 |
| z |
0.000 |
0.000 |
-40.570 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.819 |
0.233 |
0.000 |
| y |
0.233 |
-0.417 |
0.000 |
| z |
0.000 |
0.000 |
-0.402 |
|
| Polar |
|---|
| 3z2-r2 | -0.805 |
|---|
| x2-y2 | 0.824 |
|---|
| xy | 0.233 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.292 |
-0.650 |
0.000 |
| y |
-0.650 |
11.349 |
0.000 |
| z |
0.000 |
0.000 |
9.671 |
<r2> (average value of r
2) Å
2
| <r2> |
194.278 |
| (<r2>)1/2 |
13.938 |
