Jump to
S1C2
S2C1
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.041837 |
| Energy at 298.15K | -39.041652 |
| HF Energy | -38.926521 |
| Nuclear repulsion energy | 6.059967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.111 |
| H2 |
0.000 |
1.001 |
-0.333 |
| H3 |
0.000 |
-1.001 |
-0.333 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0957 | 1.0957 |
H2 | 1.0957 | | 2.0030 | H3 | 1.0957 | 2.0030 | |
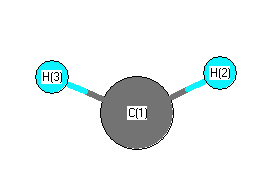 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
132.141 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.030929 |
| Energy at 298.15K | |
| HF Energy | -38.914067 |
| Nuclear repulsion energy | 6.099796 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| H2 |
0.000 |
0.000 |
1.084 |
| H3 |
0.000 |
0.000 |
-1.084 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0839 | 1.0839 |
H2 | 1.0839 | | 2.1677 | H3 | 1.0839 | 2.1677 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.022604 |
| Energy at 298.15K | -39.022440 |
| HF Energy | -38.880465 |
| Nuclear repulsion energy | 5.928208 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.180 |
| H2 |
0.000 |
0.868 |
-0.541 |
| H3 |
0.000 |
-0.868 |
-0.541 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.1292 | 1.1292 |
H2 | 1.1292 | | 1.7368 | H3 | 1.1292 | 1.7368 | |
More geometry information
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.022604 |
| Energy at 298.15K | -39.022440 |
| HF Energy | -38.880465 |
| Nuclear repulsion energy | 5.928208 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
