return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-307.452871
Energy at 298.15K-307.458500
HF Energy-306.367104
Nuclear repulsion energy311.645121
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3267 3135 0.00      
2 Ag 3244 3112 0.00      
3 Ag 3232 3101 0.00      
4 Ag 1710 1640 0.00      
5 Ag 1606 1541 0.00      
6 Ag 1390 1333 0.00      
7 Ag 1346 1291 0.00      
8 Ag 1193 1145 0.00      
9 Ag 1102 1057 0.00      
10 Ag 965 926 0.00      
11 Ag 852 818 0.00      
12 Ag 717 688 0.00      
13 Ag 657 630 0.00      
14 Au 899 863 0.00      
15 Au 861 826 24.13      
16 Au 727 697 76.19      
17 Au 491 471 5.16      
18 Au 293 281 2.18      
19 Au 159 152 5.94      
20 Bg 910 873 0.00      
21 Bg 878 842 0.00      
22 Bg 765 734 0.00      
23 Bg 650 624 0.00      
24 Bg 524 503 0.00      
25 Bu 3267 3135 22.25      
26 Bu 3244 3112 18.80      
27 Bu 3233 3102 10.42      
28 Bu 1693 1624 10.86      
29 Bu 1579 1515 12.88      
30 Bu 1347 1293 14.84      
31 Bu 1265 1213 13.82      
32 Bu 1124 1078 23.93      
33 Bu 1054 1011 6.58      
34 Bu 949 910 0.98      
35 Bu 843 809 5.71      
36 Bu 445 427 3.20      

Unscaled Zero Point Vibrational Energy (zpe) 24238.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23254.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.17039 0.07123 0.05023

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.356 1.104 0.000
C2 2.182 -0.168 0.000
C3 1.356 -1.257 0.000
C4 -1.356 -1.104 0.000
C5 -2.182 0.168 0.000
C6 -1.356 1.257 0.000
C7 0.043 0.741 0.000
C8 -0.043 -0.741 0.000
H9 1.783 2.113 0.000
H10 3.277 -0.183 0.000
H11 1.656 -2.308 0.000
H12 -1.783 -2.113 0.000
H13 -3.277 0.183 0.000
H14 -1.656 2.308 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.51722.36073.49733.65952.71611.36212.31541.09542.31253.42524.49484.72343.2435
C21.51721.36613.65954.37693.81412.32382.29762.31601.09502.20314.41625.47014.5675
C32.36071.36612.71613.81413.69732.38981.49133.39642.20041.09333.25374.85134.6665
C43.49733.65952.71611.51722.36072.31541.36214.49484.72343.24351.09542.31253.4252
C53.65954.37693.81411.51721.36612.29762.32384.41625.47014.56752.31601.09502.2031
C62.71613.81413.69732.36071.36611.49132.38983.25374.85134.66653.39642.20041.0933
C71.36212.32382.38982.31542.29761.49131.48362.21593.36293.44873.38793.36652.3117
C82.31542.29761.49131.36212.32382.38981.48363.38793.36652.31172.21593.36293.4487
H91.09542.31603.39644.49484.41623.25372.21593.38792.73934.42265.52955.41543.4446
H102.31251.09502.20044.72345.47014.85133.36293.36652.73932.67225.41546.56395.5261
H113.42522.20311.09333.24354.56754.66653.44872.31174.42262.67223.44465.52615.6810
H124.49484.41623.25371.09542.31603.39643.38792.21595.52955.41543.44462.73934.4226
H134.72345.47014.85132.31251.09502.20043.36653.36295.41546.56395.52612.73932.6722
H143.24354.56754.66653.42522.20311.09332.31173.44873.44465.52615.68104.42262.6722

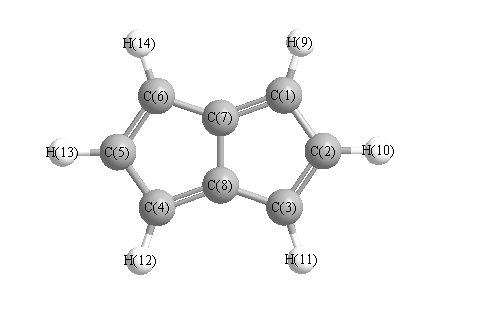
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.811 C1 C2 H10 123.761
C1 C7 C6 144.271 C1 C7 C8 108.831
C2 C1 C7 107.499 C2 C1 H9 124.064
C2 C3 C8 106.961 C2 C3 H11 126.867
C3 C2 H10 126.428 C3 C8 C4 144.271
C3 C8 C7 106.898 C4 C5 C6 109.811
C4 C5 H13 123.761 C4 C8 C7 108.831
C5 C4 C8 107.499 C5 C4 H12 124.064
C5 C6 C7 106.961 C5 C6 H14 126.867
C6 C5 H13 126.428 C6 C7 C8 106.898
C7 C1 H9 128.436 C7 C6 H14 126.172
C8 C3 H11 126.172 C8 C4 H12 128.436
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability