return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-307.442343
Energy at 298.15K-307.447999
HF Energy-306.367730
Nuclear repulsion energy311.922413
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3276 3134 0.00      
2 Ag 3252 3111 0.00      
3 Ag 3241 3100 0.00      
4 Ag 1730 1655 0.00      
5 Ag 1625 1555 0.00      
6 Ag 1396 1336 0.00      
7 Ag 1351 1293 0.00      
8 Ag 1198 1146 0.00      
9 Ag 1106 1058 0.00      
10 Ag 971 929 0.00      
11 Ag 861 824 0.00      
12 Ag 725 693 0.00      
13 Ag 660 631 0.00      
14 Au 907 867 0.00      
15 Au 866 828 26.01      
16 Au 729 697 76.64      
17 Au 493 472 4.66      
18 Au 294 281 2.28      
19 Au 157 150 5.82      
20 Bg 917 877 0.00      
21 Bg 881 843 0.00      
22 Bg 767 733 0.00      
23 Bg 652 624 0.00      
24 Bg 525 503 0.00      
25 Bu 3276 3134 20.69      
26 Bu 3252 3112 17.61      
27 Bu 3241 3101 9.41      
28 Bu 1706 1632 6.78      
29 Bu 1592 1523 11.04      
30 Bu 1353 1294 14.85      
31 Bu 1270 1215 12.78      
32 Bu 1128 1079 19.81      
33 Bu 1059 1013 5.40      
34 Bu 953 912 0.73      
35 Bu 848 811 4.63      
36 Bu 448 428 3.14      

Unscaled Zero Point Vibrational Energy (zpe) 24351.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23297.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.17066 0.07137 0.05032

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.355 1.102 0.000
C2 2.180 -0.170 0.000
C3 1.355 -1.257 0.000
C4 -1.355 -1.102 0.000
C5 -2.180 0.170 0.000
C6 -1.355 1.257 0.000
C7 0.044 0.740 0.000
C8 -0.044 -0.740 0.000
H9 1.782 2.110 0.000
H10 3.274 -0.185 0.000
H11 1.655 -2.308 0.000
H12 -1.782 -2.110 0.000
H13 -3.274 0.185 0.000
H14 -1.655 2.308 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.51622.35883.49293.65512.71391.36032.31271.09512.31123.42314.49014.71873.2419
C21.51621.36423.65514.37233.81142.32182.29502.31481.09472.20114.41135.46524.5652
C32.35881.36422.71393.81143.69572.38871.49073.39412.19811.09303.25104.84834.6649
C43.49293.65512.71391.51622.35882.31271.36034.49014.71873.24191.09512.31123.4231
C53.65514.37233.81141.51621.36422.29502.32184.41135.46524.56522.31481.09472.2011
C62.71393.81143.69572.35881.36421.49072.38873.25104.84834.66493.39412.19811.0930
C71.36032.32182.38872.31272.29501.49071.48252.21383.36053.44733.38493.36372.3113
C82.31272.29501.49071.36032.32182.38871.48253.38493.36372.31132.21383.36053.4473
H91.09512.31483.39414.49014.41133.25102.21383.38492.73794.42005.52455.41033.4425
H102.31121.09472.19814.71875.46524.84833.36053.36372.73792.66975.41036.55875.5234
H113.42312.20111.09303.24194.56524.66493.44732.31134.42002.66973.44255.52345.6793
H124.49014.41133.25101.09512.31483.39413.38492.21385.52455.41033.44252.73794.4200
H134.71875.46524.84832.31121.09472.19813.36373.36055.41036.55875.52342.73792.6697
H143.24194.56524.66493.42312.20111.09302.31133.44733.44255.52345.67934.42002.6697

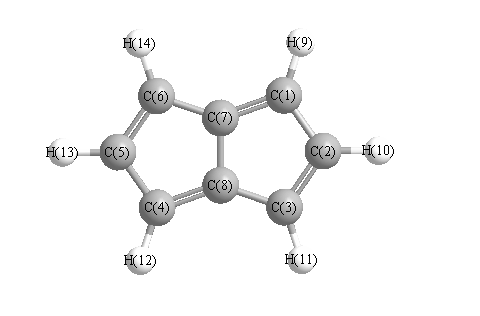
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.844 C1 C2 H10 123.756
C1 C7 C6 144.277 C1 C7 C8 108.811
C2 C1 C7 107.513 C2 C1 H9 124.068
C2 C3 C8 106.920 C2 C3 H11 126.874
C3 C2 H10 126.401 C3 C8 C4 144.277
C3 C8 C7 106.912 C4 C5 C6 109.844
C4 C5 H13 123.756 C4 C8 C7 108.811
C5 C4 C8 107.513 C5 C4 H12 124.068
C5 C6 C7 106.920 C5 C6 H14 126.874
C6 C5 H13 126.401 C6 C7 C8 106.912
C7 C1 H9 128.419 C7 C6 H14 126.207
C8 C3 H11 126.207 C8 C4 H12 128.419
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability