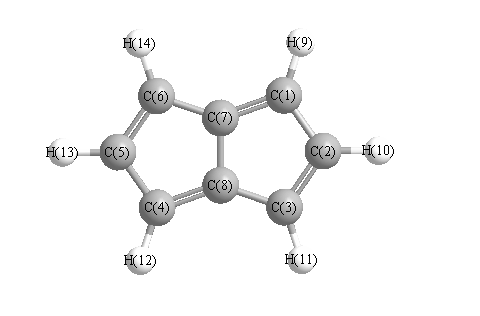
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3176 |
3158 |
0.00 |
|
|
|
| 2 |
Ag |
3154 |
3135 |
0.00 |
|
|
|
| 3 |
Ag |
3135 |
3116 |
0.00 |
|
|
|
| 4 |
Ag |
1600 |
1591 |
0.00 |
|
|
|
| 5 |
Ag |
1496 |
1487 |
0.00 |
|
|
|
| 6 |
Ag |
1350 |
1342 |
0.00 |
|
|
|
| 7 |
Ag |
1270 |
1262 |
0.00 |
|
|
|
| 8 |
Ag |
1143 |
1136 |
0.00 |
|
|
|
| 9 |
Ag |
1054 |
1048 |
0.00 |
|
|
|
| 10 |
Ag |
933 |
927 |
0.00 |
|
|
|
| 11 |
Ag |
835 |
831 |
0.00 |
|
|
|
| 12 |
Ag |
679 |
675 |
0.00 |
|
|
|
| 13 |
Ag |
636 |
632 |
0.00 |
|
|
|
| 14 |
Au |
889 |
884 |
4.86 |
|
|
|
| 15 |
Au |
841 |
837 |
6.21 |
|
|
|
| 16 |
Au |
694 |
690 |
68.74 |
|
|
|
| 17 |
Au |
494 |
491 |
9.55 |
|
|
|
| 18 |
Au |
287 |
286 |
0.85 |
|
|
|
| 19 |
Au |
146 |
145 |
5.01 |
|
|
|
| 20 |
Bg |
890 |
885 |
0.00 |
|
|
|
| 21 |
Bg |
879 |
874 |
0.00 |
|
|
|
| 22 |
Bg |
760 |
756 |
0.00 |
|
|
|
| 23 |
Bg |
645 |
642 |
0.00 |
|
|
|
| 24 |
Bg |
537 |
534 |
0.00 |
|
|
|
| 25 |
Bu |
3176 |
3158 |
21.45 |
|
|
|
| 26 |
Bu |
3155 |
3136 |
10.79 |
|
|
|
| 27 |
Bu |
3135 |
3117 |
16.03 |
|
|
|
| 28 |
Bu |
1608 |
1599 |
69.23 |
|
|
|
| 29 |
Bu |
1499 |
1490 |
14.67 |
|
|
|
| 30 |
Bu |
1286 |
1278 |
22.16 |
|
|
|
| 31 |
Bu |
1213 |
1206 |
17.18 |
|
|
|
| 32 |
Bu |
1091 |
1085 |
41.08 |
|
|
|
| 33 |
Bu |
1023 |
1017 |
1.91 |
|
|
|
| 34 |
Bu |
929 |
924 |
16.73 |
|
|
|
| 35 |
Bu |
817 |
812 |
6.71 |
|
|
|
| 36 |
Bu |
430 |
428 |
3.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23442.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23306.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.063 |
|
|
|
| 2 |
C |
0.030 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
C |
-0.063 |
|
|
|
| 5 |
C |
0.030 |
|
|
|
| 6 |
C |
-0.024 |
|
|
|
| 7 |
C |
0.123 |
|
|
|
| 8 |
C |
0.123 |
|
|
|
| 9 |
H |
-0.020 |
|
|
|
| 10 |
H |
-0.021 |
|
|
|
| 11 |
H |
-0.026 |
|
|
|
| 12 |
H |
-0.020 |
|
|
|
| 13 |
H |
-0.021 |
|
|
|
| 14 |
H |
-0.026 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.072 |
1.259 |
0.000 |
| y |
1.259 |
-40.666 |
0.000 |
| z |
0.000 |
0.000 |
-50.197 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.359 |
1.259 |
0.000 |
| y |
1.259 |
4.969 |
0.000 |
| z |
0.000 |
0.000 |
-9.328 |
|
| Polar |
|---|
| 3z2-r2 | -18.656 |
|---|
| x2-y2 | -0.407 |
|---|
| xy | 1.259 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.679 |
0.825 |
0.000 |
| y |
0.825 |
12.923 |
0.000 |
| z |
0.000 |
0.000 |
4.650 |
<r2> (average value of r
2) Å
2
| <r2> |
220.739 |
| (<r2>)1/2 |
14.857 |