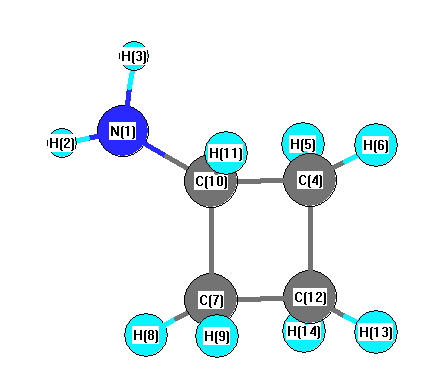
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3308 |
3313 |
9.13 |
|
|
|
| 2 |
A' |
3041 |
3046 |
66.19 |
|
|
|
| 3 |
A' |
3011 |
3016 |
20.09 |
|
|
|
| 4 |
A' |
2982 |
2986 |
36.89 |
|
|
|
| 5 |
A' |
2967 |
2971 |
19.47 |
|
|
|
| 6 |
A' |
2850 |
2855 |
113.05 |
|
|
|
| 7 |
A' |
1593 |
1596 |
10.75 |
|
|
|
| 8 |
A' |
1430 |
1433 |
3.35 |
|
|
|
| 9 |
A' |
1402 |
1404 |
1.06 |
|
|
|
| 10 |
A' |
1320 |
1322 |
23.48 |
|
|
|
| 11 |
A' |
1238 |
1240 |
1.04 |
|
|
|
| 12 |
A' |
1189 |
1191 |
0.39 |
|
|
|
| 13 |
A' |
1110 |
1111 |
7.05 |
|
|
|
| 14 |
A' |
1050 |
1051 |
8.78 |
|
|
|
| 15 |
A' |
938 |
939 |
12.77 |
|
|
|
| 16 |
A' |
854 |
855 |
12.70 |
|
|
|
| 17 |
A' |
835 |
837 |
89.11 |
|
|
|
| 18 |
A' |
789 |
790 |
3.15 |
|
|
|
| 19 |
A' |
654 |
655 |
0.51 |
|
|
|
| 20 |
A' |
390 |
391 |
4.21 |
|
|
|
| 21 |
A' |
110 |
110 |
1.02 |
|
|
|
| 22 |
A" |
3379 |
3384 |
2.62 |
|
|
|
| 23 |
A" |
3018 |
3023 |
5.77 |
|
|
|
| 24 |
A" |
2963 |
2968 |
82.35 |
|
|
|
| 25 |
A" |
1396 |
1399 |
1.16 |
|
|
|
| 26 |
A" |
1295 |
1297 |
0.48 |
|
|
|
| 27 |
A" |
1228 |
1230 |
0.02 |
|
|
|
| 28 |
A" |
1215 |
1217 |
0.29 |
|
|
|
| 29 |
A" |
1181 |
1183 |
0.00 |
|
|
|
| 30 |
A" |
1125 |
1127 |
1.43 |
|
|
|
| 31 |
A" |
997 |
999 |
0.49 |
|
|
|
| 32 |
A" |
912 |
913 |
0.21 |
|
|
|
| 33 |
A" |
883 |
884 |
1.16 |
|
|
|
| 34 |
A" |
738 |
739 |
0.50 |
|
|
|
| 35 |
A" |
375 |
376 |
7.97 |
|
|
|
| 36 |
A" |
270 |
271 |
28.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27017.2 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 27060.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.115 |
|
|
|
| 2 |
H |
0.060 |
|
|
|
| 3 |
H |
0.060 |
|
|
|
| 4 |
C |
0.087 |
|
|
|
| 5 |
H |
-0.021 |
|
|
|
| 6 |
H |
-0.017 |
|
|
|
| 7 |
C |
0.087 |
|
|
|
| 8 |
H |
-0.021 |
|
|
|
| 9 |
H |
-0.017 |
|
|
|
| 10 |
C |
-0.059 |
|
|
|
| 11 |
H |
-0.056 |
|
|
|
| 12 |
C |
0.028 |
|
|
|
| 13 |
H |
-0.015 |
|
|
|
| 14 |
H |
-0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.194 |
1.140 |
0.000 |
1.156 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.073 |
-2.144 |
0.000 |
| y |
-2.144 |
-30.894 |
0.000 |
| z |
0.000 |
0.000 |
-32.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.560 |
-2.144 |
0.000 |
| y |
-2.144 |
1.708 |
0.000 |
| z |
0.000 |
0.000 |
-0.149 |
|
| Polar |
|---|
| 3z2-r2 | -0.297 |
|---|
| x2-y2 | -2.179 |
|---|
| xy | -2.144 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.778 |
-0.664 |
0.000 |
| y |
-0.664 |
7.755 |
0.000 |
| z |
0.000 |
0.000 |
7.807 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |