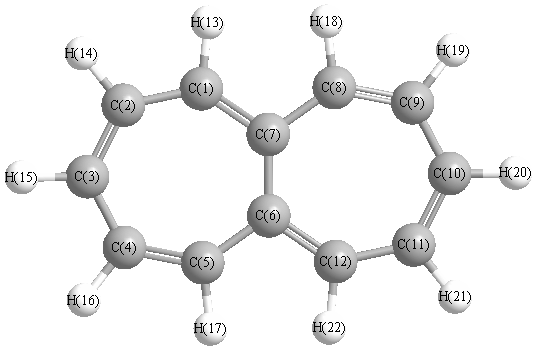
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3116 |
3098 |
2.36 |
|
|
|
| 2 |
A |
3092 |
3074 |
11.44 |
|
|
|
| 3 |
A |
3089 |
3071 |
2.15 |
|
|
|
| 4 |
A |
3071 |
3053 |
0.42 |
|
|
|
| 5 |
A |
3070 |
3052 |
0.00 |
|
|
|
| 6 |
A |
1642 |
1632 |
1.20 |
|
|
|
| 7 |
A |
1592 |
1582 |
3.59 |
|
|
|
| 8 |
A |
1516 |
1508 |
1.33 |
|
|
|
| 9 |
A |
1433 |
1425 |
0.09 |
|
|
|
| 10 |
A |
1400 |
1391 |
0.00 |
|
|
|
| 11 |
A |
1360 |
1352 |
0.39 |
|
|
|
| 12 |
A |
1225 |
1218 |
0.17 |
|
|
|
| 13 |
A |
1210 |
1203 |
0.02 |
|
|
|
| 14 |
A |
1188 |
1181 |
0.39 |
|
|
|
| 15 |
A |
1049 |
1043 |
1.00 |
|
|
|
| 16 |
A |
968 |
963 |
1.04 |
|
|
|
| 17 |
A |
957 |
951 |
0.09 |
|
|
|
| 18 |
A |
942 |
937 |
0.02 |
|
|
|
| 19 |
A |
868 |
863 |
2.07 |
|
|
|
| 20 |
A |
860 |
855 |
9.39 |
|
|
|
| 21 |
A |
790 |
786 |
2.96 |
|
|
|
| 22 |
A |
752 |
748 |
0.62 |
|
|
|
| 23 |
A |
704 |
700 |
61.65 |
|
|
|
| 24 |
A |
635 |
631 |
2.58 |
|
|
|
| 25 |
A |
566 |
563 |
0.07 |
|
|
|
| 26 |
A |
409 |
406 |
4.57 |
|
|
|
| 27 |
A |
377 |
375 |
3.61 |
|
|
|
| 28 |
A |
351 |
349 |
1.61 |
|
|
|
| 29 |
A |
289 |
287 |
2.38 |
|
|
|
| 30 |
A |
119 |
119 |
0.87 |
|
|
|
| 31 |
A |
59 |
59 |
0.03 |
|
|
|
| 32 |
B |
3116 |
3098 |
51.15 |
|
|
|
| 33 |
B |
3093 |
3075 |
51.45 |
|
|
|
| 34 |
B |
3088 |
3070 |
5.29 |
|
|
|
| 35 |
B |
3072 |
3055 |
7.06 |
|
|
|
| 36 |
B |
3069 |
3051 |
16.09 |
|
|
|
| 37 |
B |
1618 |
1608 |
68.14 |
|
|
|
| 38 |
B |
1595 |
1586 |
77.92 |
|
|
|
| 39 |
B |
1533 |
1524 |
56.65 |
|
|
|
| 40 |
B |
1429 |
1420 |
10.32 |
|
|
|
| 41 |
B |
1356 |
1348 |
8.11 |
|
|
|
| 42 |
B |
1288 |
1281 |
2.85 |
|
|
|
| 43 |
B |
1229 |
1222 |
13.78 |
|
|
|
| 44 |
B |
1200 |
1193 |
0.75 |
|
|
|
| 45 |
B |
1113 |
1106 |
3.00 |
|
|
|
| 46 |
B |
1026 |
1020 |
3.98 |
|
|
|
| 47 |
B |
956 |
950 |
1.05 |
|
|
|
| 48 |
B |
951 |
945 |
3.31 |
|
|
|
| 49 |
B |
925 |
920 |
7.65 |
|
|
|
| 50 |
B |
888 |
883 |
0.06 |
|
|
|
| 51 |
B |
845 |
841 |
10.48 |
|
|
|
| 52 |
B |
794 |
790 |
47.15 |
|
|
|
| 53 |
B |
719 |
715 |
7.49 |
|
|
|
| 54 |
B |
641 |
637 |
4.78 |
|
|
|
| 55 |
B |
578 |
575 |
22.09 |
|
|
|
| 56 |
B |
562 |
559 |
2.26 |
|
|
|
| 57 |
B |
411 |
409 |
0.76 |
|
|
|
| 58 |
B |
344 |
342 |
0.84 |
|
|
|
| 59 |
B |
278 |
276 |
0.61 |
|
|
|
| 60 |
B |
150 |
149 |
3.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38281.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 38059.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.005 |
|
|
|
| 2 |
C |
0.028 |
|
|
|
| 3 |
C |
-0.002 |
|
|
|
| 4 |
C |
0.024 |
|
|
|
| 5 |
C |
-0.008 |
|
|
|
| 6 |
C |
0.001 |
|
|
|
| 7 |
C |
0.001 |
|
|
|
| 8 |
C |
-0.008 |
|
|
|
| 9 |
C |
0.024 |
|
|
|
| 10 |
C |
-0.002 |
|
|
|
| 11 |
C |
0.028 |
|
|
|
| 12 |
C |
0.005 |
|
|
|
| 13 |
H |
-0.025 |
|
|
|
| 14 |
H |
-0.008 |
|
|
|
| 15 |
H |
-0.013 |
|
|
|
| 16 |
H |
-0.004 |
|
|
|
| 17 |
H |
0.002 |
|
|
|
| 18 |
H |
0.002 |
|
|
|
| 19 |
H |
-0.004 |
|
|
|
| 20 |
H |
-0.013 |
|
|
|
| 21 |
H |
-0.008 |
|
|
|
| 22 |
H |
-0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.258 |
0.258 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-63.881 |
-0.814 |
0.000 |
| y |
-0.814 |
-62.635 |
0.000 |
| z |
0.000 |
0.000 |
-73.711 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.292 |
-0.814 |
0.000 |
| y |
-0.814 |
6.161 |
0.000 |
| z |
0.000 |
0.000 |
-10.453 |
|
| Polar |
|---|
| 3z2-r2 | -20.905 |
|---|
| x2-y2 | -1.246 |
|---|
| xy | -0.814 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
21.014 |
0.038 |
0.000 |
| y |
0.038 |
34.626 |
0.000 |
| z |
0.000 |
0.000 |
9.483 |
<r2> (average value of r
2) Å
2
| <r2> |
563.259 |
| (<r2>)1/2 |
23.733 |