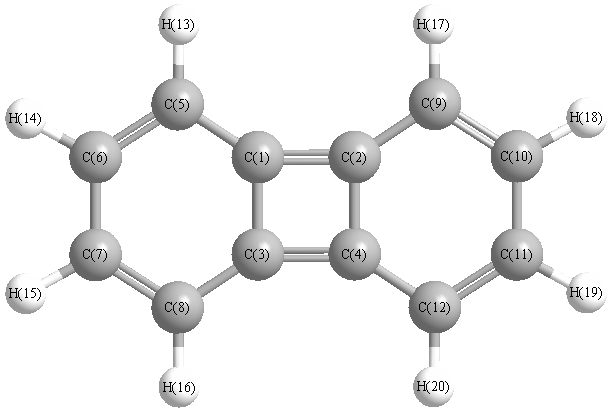
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3137 |
3119 |
0.00 |
|
|
|
| 2 |
Ag |
3121 |
3103 |
0.00 |
|
|
|
| 3 |
Ag |
1686 |
1676 |
0.00 |
|
|
|
| 4 |
Ag |
1497 |
1488 |
0.00 |
|
|
|
| 5 |
Ag |
1384 |
1376 |
0.00 |
|
|
|
| 6 |
Ag |
1156 |
1150 |
0.00 |
|
|
|
| 7 |
Ag |
1114 |
1107 |
0.00 |
|
|
|
| 8 |
Ag |
994 |
988 |
0.00 |
|
|
|
| 9 |
Ag |
761 |
757 |
0.00 |
|
|
|
| 10 |
Ag |
391 |
389 |
0.00 |
|
|
|
| 11 |
Au |
950 |
944 |
0.00 |
|
|
|
| 12 |
Au |
877 |
871 |
0.00 |
|
|
|
| 13 |
Au |
814 |
809 |
0.00 |
|
|
|
| 14 |
Au |
572 |
569 |
0.00 |
|
|
|
| 15 |
Au |
142 |
141 |
0.00 |
|
|
|
| 16 |
B1g |
950 |
945 |
0.00 |
|
|
|
| 17 |
B1g |
852 |
847 |
0.00 |
|
|
|
| 18 |
B1g |
707 |
703 |
0.00 |
|
|
|
| 19 |
B1g |
450 |
448 |
0.00 |
|
|
|
| 20 |
B1u |
3136 |
3118 |
35.63 |
|
|
|
| 21 |
B1u |
3120 |
3102 |
10.88 |
|
|
|
| 22 |
B1u |
1596 |
1587 |
0.96 |
|
|
|
| 23 |
B1u |
1409 |
1401 |
27.22 |
|
|
|
| 24 |
B1u |
1327 |
1319 |
25.95 |
|
|
|
| 25 |
B1u |
1141 |
1135 |
4.73 |
|
|
|
| 26 |
B1u |
1014 |
1008 |
0.06 |
|
|
|
| 27 |
B1u |
982 |
976 |
17.41 |
|
|
|
| 28 |
B1u |
602 |
599 |
3.76 |
|
|
|
| 29 |
B2g |
903 |
898 |
0.00 |
|
|
|
| 30 |
B2g |
734 |
729 |
0.00 |
|
|
|
| 31 |
B2g |
432 |
429 |
0.00 |
|
|
|
| 32 |
B2g |
310 |
309 |
0.00 |
|
|
|
| 33 |
B2u |
3129 |
3111 |
43.81 |
|
|
|
| 34 |
B2u |
3109 |
3091 |
0.04 |
|
|
|
| 35 |
B2u |
1594 |
1585 |
2.20 |
|
|
|
| 36 |
B2u |
1431 |
1423 |
3.48 |
|
|
|
| 37 |
B2u |
1241 |
1234 |
5.06 |
|
|
|
| 38 |
B2u |
1113 |
1106 |
12.45 |
|
|
|
| 39 |
B2u |
1039 |
1033 |
0.78 |
|
|
|
| 40 |
B2u |
712 |
708 |
0.41 |
|
|
|
| 41 |
B2u |
209 |
207 |
0.73 |
|
|
|
| 42 |
B3g |
3129 |
3111 |
0.00 |
|
|
|
| 43 |
B3g |
3108 |
3090 |
0.00 |
|
|
|
| 44 |
B3g |
1605 |
1596 |
0.00 |
|
|
|
| 45 |
B3g |
1431 |
1423 |
0.00 |
|
|
|
| 46 |
B3g |
1250 |
1242 |
0.00 |
|
|
|
| 47 |
B3g |
1080 |
1073 |
0.00 |
|
|
|
| 48 |
B3g |
962 |
956 |
0.00 |
|
|
|
| 49 |
B3g |
589 |
586 |
0.00 |
|
|
|
| 50 |
B3g |
550 |
547 |
0.00 |
|
|
|
| 51 |
B3u |
899 |
894 |
5.73 |
|
|
|
| 52 |
B3u |
722 |
718 |
96.55 |
|
|
|
| 53 |
B3u |
362 |
360 |
2.25 |
|
|
|
| 54 |
B3u |
101 |
100 |
2.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33811.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33615.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.145 |
|
|
|
| 2 |
C |
0.145 |
|
|
|
| 3 |
C |
0.145 |
|
|
|
| 4 |
C |
0.145 |
|
|
|
| 5 |
C |
-0.131 |
|
|
|
| 6 |
C |
0.033 |
|
|
|
| 7 |
C |
0.033 |
|
|
|
| 8 |
C |
-0.131 |
|
|
|
| 9 |
C |
-0.131 |
|
|
|
| 10 |
C |
0.033 |
|
|
|
| 11 |
C |
0.033 |
|
|
|
| 12 |
C |
-0.131 |
|
|
|
| 13 |
H |
-0.030 |
|
|
|
| 14 |
H |
-0.016 |
|
|
|
| 15 |
H |
-0.016 |
|
|
|
| 16 |
H |
-0.030 |
|
|
|
| 17 |
H |
-0.030 |
|
|
|
| 18 |
H |
-0.016 |
|
|
|
| 19 |
H |
-0.016 |
|
|
|
| 20 |
H |
-0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-74.011 |
0.000 |
0.000 |
| y |
0.000 |
-60.756 |
0.000 |
| z |
0.000 |
0.000 |
-60.060 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.603 |
0.000 |
0.000 |
| y |
0.000 |
6.280 |
0.000 |
| z |
0.000 |
0.000 |
7.323 |
|
| Polar |
|---|
| 3z2-r2 | 14.647 |
|---|
| x2-y2 | -13.256 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.576 |
0.000 |
0.000 |
| y |
0.000 |
19.975 |
0.000 |
| z |
0.000 |
0.000 |
33.596 |
<r2> (average value of r
2) Å
2
| <r2> |
568.552 |
| (<r2>)1/2 |
23.844 |