Jump to
S1C2
S2C1
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.075359 |
| Energy at 298.15K | -131.074757 |
| HF Energy | -130.690884 |
| Nuclear repulsion energy | 46.492695 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3329 |
3173 |
|
|
|
|
| 2 |
A' |
1834 |
1748 |
|
|
|
|
| 3 |
A' |
1138 |
1085 |
|
|
|
|
| 4 |
A' |
661 |
630 |
|
|
|
|
| 5 |
A' |
383 |
365 |
|
|
|
|
| 6 |
A" |
418 |
398 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3881.3 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 3699.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.057 |
-1.257 |
0.000 |
| C2 |
0.000 |
0.095 |
0.000 |
| N3 |
-0.159 |
1.293 |
0.000 |
| H4 |
0.770 |
-2.080 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3540 | 2.5600 | 1.0888 |
C2 | 1.3540 | | 1.2085 | 2.3081 | N3 | 2.5600 | 1.2085 | | 3.4994 | H4 | 1.0888 | 2.3081 | 3.4994 | |
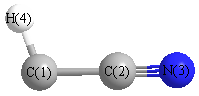 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
174.858 |
|
C2 |
C1 |
H4 |
141.532 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.072487 |
| Energy at 298.15K | |
| HF Energy | -130.688672 |
| Nuclear repulsion energy | 46.633453 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3424 |
3263 |
|
|
|
|
| 2 |
Σ |
1673 |
1595 |
|
|
|
|
| 3 |
Σ |
1222 |
1165 |
|
|
|
|
| 4 |
Π |
420 |
401 |
|
|
|
|
| 4 |
Π |
420 |
401 |
|
|
|
|
| 5 |
Π |
452i |
431i |
|
|
|
|
| 5 |
Π |
452i |
431i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3127.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 2980.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.231 |
| C2 |
0.000 |
0.000 |
0.087 |
| N3 |
0.000 |
0.000 |
1.311 |
| H4 |
0.000 |
0.000 |
-2.311 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3180 | 2.5425 | 1.0797 |
C2 | 1.3180 | | 1.2245 | 2.3977 | N3 | 2.5425 | 1.2245 | | 3.6221 | H4 | 1.0797 | 2.3977 | 3.6221 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at QCISD(T)/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.022 |
|
|
|
| 2 |
C |
0.044 |
|
|
|
| 3 |
N |
-0.145 |
|
|
|
| 4 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
36.994 |
| (<r2>)1/2 |
6.082 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.054746 |
| Energy at 298.15K | -131.054122 |
| HF Energy | -130.626720 |
| Nuclear repulsion energy | 46.092578 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3037 |
2894 |
|
|
|
|
| 2 |
A' |
2050 |
1954 |
|
|
|
|
| 3 |
A' |
1046 |
997 |
|
|
|
|
| 4 |
A' |
977 |
931 |
|
|
|
|
| 5 |
A' |
405 |
386 |
|
|
|
|
| 6 |
A" |
292 |
278 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3902.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 3719.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.111 |
-1.314 |
0.000 |
| C2 |
0.000 |
0.099 |
0.000 |
| N3 |
-0.267 |
1.265 |
0.000 |
| H4 |
1.201 |
-1.566 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4171 | 2.6067 | 1.1179 |
C2 | 1.4171 | | 1.1965 | 2.0526 | N3 | 2.6067 | 1.1965 | | 3.1890 | H4 | 1.1179 | 2.0526 | 3.1890 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.054746 |
| Energy at 298.15K | -131.054122 |
| HF Energy | -130.626720 |
| Nuclear repulsion energy | 46.092578 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
