Jump to
S1C2
S2C1
S2C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.396676 |
| Energy at 298.15K | -131.395875 |
| HF Energy | -131.396676 |
| Nuclear repulsion energy | 46.781259 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3317 |
3323 |
44.20 |
|
|
|
| 2 |
A' |
1772 |
1775 |
20.54 |
|
|
|
| 3 |
A' |
1220 |
1222 |
2.84 |
|
|
|
| 4 |
A' |
435 |
436 |
5.97 |
|
|
|
| 5 |
A' |
321 |
322 |
46.70 |
|
|
|
| 6 |
A" |
433 |
434 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3749.5 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 3755.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.115 |
-1.226 |
0.000 |
| C2 |
0.000 |
0.083 |
0.000 |
| N3 |
-0.190 |
1.290 |
0.000 |
| H4 |
0.641 |
-2.174 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3137 | 2.5342 | 1.0841 |
C2 | 1.3137 | | 1.2220 | 2.3459 | N3 | 2.5342 | 1.2220 | | 3.5620 | H4 | 1.0841 | 2.3459 | 3.5620 | |
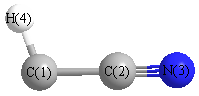 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
176.072 |
|
C2 |
C1 |
H4 |
155.987 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.396426 |
| Energy at 298.15K | |
| HF Energy | -131.396426 |
| Nuclear repulsion energy | 46.822858 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3375 |
3380 |
70.37 |
|
|
|
| 2 |
Σ |
1749 |
1751 |
20.42 |
|
|
|
| 3 |
Σ |
1241 |
1243 |
2.19 |
|
|
|
| 4 |
Π |
433 |
434 |
0.21 |
|
|
|
| 4 |
Π |
433 |
434 |
0.21 |
|
|
|
| 5 |
Π |
205i |
205i |
40.07 |
|
|
|
| 5 |
Π |
205i |
205i |
40.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3410.0 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 3415.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.222 |
| C2 |
0.000 |
0.000 |
0.080 |
| N3 |
0.000 |
0.000 |
1.308 |
| H4 |
0.000 |
0.000 |
-2.301 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3022 | 2.5298 | 1.0790 |
C2 | 1.3022 | | 1.2275 | 2.3812 | N3 | 2.5298 | 1.2275 | | 3.6088 | H4 | 1.0790 | 2.3812 | 3.6088 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.051 |
|
|
|
| 2 |
C |
0.013 |
|
|
|
| 3 |
N |
-0.086 |
|
|
|
| 4 |
H |
0.022 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.975 |
2.975 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.060 |
0.000 |
0.000 |
| y |
0.000 |
-17.060 |
0.000 |
| z |
0.000 |
0.000 |
-15.279 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.891 |
0.000 |
0.000 |
| y |
0.000 |
-0.891 |
0.000 |
| z |
0.000 |
0.000 |
1.782 |
|
| Polar |
|---|
| 3z2-r2 | 3.564 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.829 |
0.000 |
0.000 |
| y |
0.000 |
1.829 |
0.000 |
| z |
0.000 |
0.000 |
6.397 |
<r2> (average value of r
2) Å
2
| <r2> |
36.549 |
| (<r2>)1/2 |
6.046 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.375834 |
| Energy at 298.15K | -131.375250 |
| HF Energy | -131.375834 |
| Nuclear repulsion energy | 46.408024 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2959 |
2964 |
13.94 |
|
|
|
| 2 |
A' |
2005 |
2008 |
16.25 |
|
|
|
| 3 |
A' |
1073 |
1075 |
23.80 |
|
|
|
| 4 |
A' |
913 |
914 |
49.63 |
|
|
|
| 5 |
A' |
439 |
440 |
18.93 |
|
|
|
| 6 |
A" |
307 |
307 |
6.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3847.9 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 3854.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.274 |
-1.265 |
0.000 |
| C2 |
0.000 |
0.092 |
0.000 |
| N3 |
-0.431 |
1.213 |
0.000 |
| H4 |
1.376 |
-1.456 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3834 | 2.5764 | 1.1186 |
C2 | 1.3834 | | 1.2020 | 2.0707 | N3 | 2.5764 | 1.2020 | | 3.2237 | H4 | 1.1186 | 2.0707 | 3.2237 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.007 |
|
|
|
| 2 |
C |
-0.025 |
|
|
|
| 3 |
N |
-0.057 |
|
|
|
| 4 |
H |
0.090 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.763 |
-0.909 |
0.000 |
1.983 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.309 |
-1.560 |
0.000 |
| y |
-1.560 |
-21.118 |
0.000 |
| z |
0.000 |
0.000 |
-15.701 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.101 |
-1.560 |
0.000 |
| y |
-1.560 |
-5.113 |
0.000 |
| z |
0.000 |
0.000 |
3.012 |
|
| Polar |
|---|
| 3z2-r2 | 6.025 |
|---|
| x2-y2 | 4.809 |
|---|
| xy | -1.560 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.961 |
-1.289 |
0.000 |
| y |
-1.289 |
6.004 |
0.000 |
| z |
0.000 |
0.000 |
2.270 |
<r2> (average value of r
2) Å
2
| <r2> |
36.779 |
| (<r2>)1/2 |
6.065 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.375834 |
| Energy at 298.15K | -131.375250 |
| HF Energy | -131.375834 |
| Nuclear repulsion energy | 46.408024 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
