Jump to
S1C2
S2C1
S2C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.666436 |
| Energy at 298.15K | -130.666052 |
| HF Energy | -130.666436 |
| Nuclear repulsion energy | 47.196431 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
2953 |
8.26 |
|
|
|
| 2 |
A' |
2545 |
2190 |
3.80 |
|
|
|
| 3 |
A' |
1098 |
945 |
4.38 |
|
|
|
| 4 |
A' |
1026 |
883 |
13.02 |
|
|
|
| 5 |
A' |
432 |
372 |
4.11 |
|
|
|
| 6 |
A" |
489 |
421 |
5.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4509.9 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 3881.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.294 |
-1.246 |
0.000 |
| C2 |
0.000 |
0.131 |
0.000 |
| N3 |
0.199 |
1.254 |
0.000 |
| H4 |
0.368 |
-2.095 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4080 | 2.5482 | 1.0772 |
C2 | 1.4080 | | 1.1406 | 2.2569 | N3 | 2.5482 | 1.1406 | | 3.3540 | H4 | 1.0772 | 2.2569 | 3.3540 | |
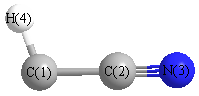 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
178.015 |
|
C2 |
C1 |
H4 |
130.025 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.649995 |
| Energy at 298.15K | |
| HF Energy | -130.649995 |
| Nuclear repulsion energy | 47.569148 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3607 |
3104 |
126.06 |
|
|
|
| 2 |
Σ |
2459 |
2117 |
17.51 |
|
|
|
| 3 |
Σ |
1178 |
1014 |
1.09 |
|
|
|
| 4 |
Π |
490 |
422 |
0.89 |
|
|
|
| 4 |
Π |
490 |
422 |
0.89 |
|
|
|
| 5 |
Π |
968i |
833i |
63.26 |
|
|
|
| 5 |
Π |
968i |
833i |
63.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3144.2 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 2706.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.234 |
| C2 |
0.000 |
0.000 |
0.130 |
| N3 |
0.000 |
0.000 |
1.275 |
| H4 |
0.000 |
0.000 |
-2.296 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3637 | 2.5091 | 1.0619 |
C2 | 1.3637 | | 1.1453 | 2.4256 | N3 | 2.5091 | 1.1453 | | 3.5710 | H4 | 1.0619 | 2.4256 | 3.5710 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.028 |
|
|
|
| 2 |
C |
0.063 |
|
|
|
| 3 |
N |
-0.160 |
|
|
|
| 4 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.864 |
3.864 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.360 |
0.000 |
0.000 |
| y |
0.000 |
-17.360 |
0.000 |
| z |
0.000 |
0.000 |
-15.816 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.772 |
0.000 |
0.000 |
| y |
0.000 |
-0.772 |
0.000 |
| z |
0.000 |
0.000 |
1.544 |
|
| Polar |
|---|
| 3z2-r2 | 3.088 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
36.411 |
| (<r2>)1/2 |
6.034 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.632189 |
| Energy at 298.15K | -130.631719 |
| HF Energy | -130.632189 |
| Nuclear repulsion energy | 47.193563 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3247 |
2794 |
8.73 |
|
|
|
| 2 |
A' |
2359 |
2030 |
36.81 |
|
|
|
| 3 |
A' |
1133 |
975 |
67.06 |
|
|
|
| 4 |
A' |
1033 |
889 |
20.91 |
|
|
|
| 5 |
A' |
479 |
412 |
26.09 |
|
|
|
| 6 |
A" |
349 |
300 |
14.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4299.3 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 3700.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.244 |
-1.277 |
0.000 |
| C2 |
0.000 |
0.111 |
0.000 |
| N3 |
0.106 |
1.256 |
0.000 |
| H4 |
0.723 |
-1.795 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4092 | 2.5564 | 1.0968 |
C2 | 1.4092 | | 1.1493 | 2.0385 | N3 | 2.5564 | 1.1493 | | 3.1122 | H4 | 1.0968 | 2.0385 | 3.1122 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.632189 |
| Energy at 298.15K | -130.631719 |
| HF Energy | -130.632189 |
| Nuclear repulsion energy | 47.193563 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
