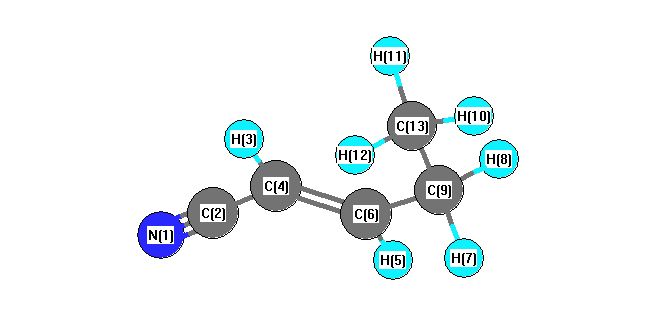
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3200 |
3088 |
1.58 |
|
|
|
| 2 |
A' |
3173 |
3062 |
5.44 |
|
|
|
| 3 |
A' |
3138 |
3028 |
18.33 |
|
|
|
| 4 |
A' |
3049 |
2942 |
16.51 |
|
|
|
| 5 |
A' |
3022 |
2917 |
15.90 |
|
|
|
| 6 |
A' |
2346 |
2264 |
27.58 |
|
|
|
| 7 |
A' |
1714 |
1654 |
28.75 |
|
|
|
| 8 |
A' |
1478 |
1426 |
4.16 |
|
|
|
| 9 |
A' |
1433 |
1382 |
14.87 |
|
|
|
| 10 |
A' |
1404 |
1355 |
5.58 |
|
|
|
| 11 |
A' |
1366 |
1318 |
7.78 |
|
|
|
| 12 |
A' |
1319 |
1273 |
1.29 |
|
|
|
| 13 |
A' |
1276 |
1232 |
1.29 |
|
|
|
| 14 |
A' |
1119 |
1080 |
4.58 |
|
|
|
| 15 |
A' |
1074 |
1037 |
5.96 |
|
|
|
| 16 |
A' |
990 |
955 |
1.39 |
|
|
|
| 17 |
A' |
910 |
878 |
5.66 |
|
|
|
| 18 |
A' |
580 |
560 |
0.48 |
|
|
|
| 19 |
A' |
509 |
492 |
0.02 |
|
|
|
| 20 |
A' |
257 |
248 |
1.22 |
|
|
|
| 21 |
A' |
145 |
140 |
3.35 |
|
|
|
| 22 |
A" |
3131 |
3021 |
20.02 |
|
|
|
| 23 |
A" |
3050 |
2943 |
7.89 |
|
|
|
| 24 |
A" |
1471 |
1419 |
7.84 |
|
|
|
| 25 |
A" |
1276 |
1231 |
0.10 |
|
|
|
| 26 |
A" |
1089 |
1051 |
1.72 |
|
|
|
| 27 |
A" |
994 |
959 |
27.91 |
|
|
|
| 28 |
A" |
848 |
818 |
9.46 |
|
|
|
| 29 |
A" |
732 |
706 |
0.88 |
|
|
|
| 30 |
A" |
493 |
475 |
4.65 |
|
|
|
| 31 |
A" |
283 |
273 |
0.01 |
|
|
|
| 32 |
A" |
181 |
175 |
0.15 |
|
|
|
| 33 |
A" |
127 |
123 |
3.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23587.1 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 22761.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.141 |
|
|
|
| 2 |
C |
-0.029 |
|
|
|
| 3 |
H |
0.054 |
|
|
|
| 4 |
C |
-0.038 |
|
|
|
| 5 |
H |
0.037 |
|
|
|
| 6 |
C |
-0.020 |
|
|
|
| 7 |
H |
0.067 |
|
|
|
| 8 |
H |
0.067 |
|
|
|
| 9 |
C |
-0.069 |
|
|
|
| 10 |
H |
0.051 |
|
|
|
| 11 |
H |
0.059 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
C |
-0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.113 |
-4.301 |
0.000 |
4.792 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.575 |
6.628 |
0.000 |
| y |
6.628 |
-44.303 |
0.000 |
| z |
0.000 |
0.000 |
-36.922 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.037 |
6.628 |
0.000 |
| y |
6.628 |
-7.055 |
0.000 |
| z |
0.000 |
0.000 |
4.017 |
|
| Polar |
|---|
| 3z2-r2 | 8.034 |
|---|
| x2-y2 | 6.728 |
|---|
| xy | 6.628 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
227.265 |
| (<r2>)1/2 |
15.075 |