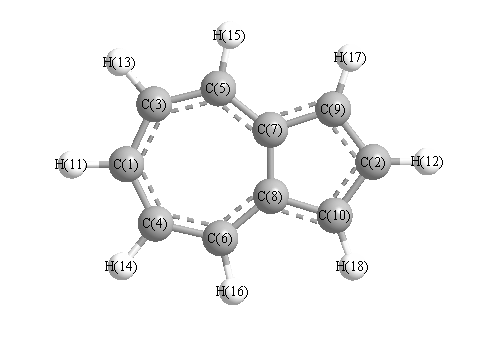
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3169 |
3151 |
8.43 |
|
|
|
| 2 |
A1 |
3141 |
3123 |
6.46 |
|
|
|
| 3 |
A1 |
3114 |
3096 |
18.47 |
|
|
|
| 4 |
A1 |
3085 |
3067 |
6.57 |
|
|
|
| 5 |
A1 |
3075 |
3057 |
0.38 |
|
|
|
| 6 |
A1 |
1613 |
1603 |
48.83 |
|
|
|
| 7 |
A1 |
1538 |
1529 |
7.97 |
|
|
|
| 8 |
A1 |
1445 |
1436 |
29.16 |
|
|
|
| 9 |
A1 |
1380 |
1372 |
49.28 |
|
|
|
| 10 |
A1 |
1270 |
1263 |
0.04 |
|
|
|
| 11 |
A1 |
1200 |
1193 |
0.23 |
|
|
|
| 12 |
A1 |
1044 |
1038 |
7.98 |
|
|
|
| 13 |
A1 |
945 |
939 |
2.14 |
|
|
|
| 14 |
A1 |
881 |
876 |
1.61 |
|
|
|
| 15 |
A1 |
806 |
802 |
3.75 |
|
|
|
| 16 |
A1 |
669 |
665 |
0.72 |
|
|
|
| 17 |
A1 |
399 |
397 |
0.98 |
|
|
|
| 18 |
A2 |
958 |
952 |
0.00 |
|
|
|
| 19 |
A2 |
849 |
844 |
0.00 |
|
|
|
| 20 |
A2 |
772 |
768 |
0.00 |
|
|
|
| 21 |
A2 |
716 |
712 |
0.00 |
|
|
|
| 22 |
A2 |
416 |
414 |
0.00 |
|
|
|
| 23 |
A2 |
160 |
159 |
0.00 |
|
|
|
| 24 |
B1 |
973 |
967 |
0.00 |
|
|
|
| 25 |
B1 |
941 |
936 |
3.81 |
|
|
|
| 26 |
B1 |
905 |
899 |
0.06 |
|
|
|
| 27 |
B1 |
760 |
756 |
77.53 |
|
|
|
| 28 |
B1 |
717 |
713 |
3.18 |
|
|
|
| 29 |
B1 |
593 |
589 |
1.12 |
|
|
|
| 30 |
B1 |
553 |
550 |
7.42 |
|
|
|
| 31 |
B1 |
313 |
311 |
7.80 |
|
|
|
| 32 |
B1 |
165 |
164 |
1.36 |
|
|
|
| 33 |
B2 |
3160 |
3142 |
11.00 |
|
|
|
| 34 |
B2 |
3106 |
3088 |
27.07 |
|
|
|
| 35 |
B2 |
3076 |
3058 |
13.73 |
|
|
|
| 36 |
B2 |
1609 |
1600 |
3.05 |
|
|
|
| 37 |
B2 |
1525 |
1516 |
7.44 |
|
|
|
| 38 |
B2 |
1437 |
1429 |
3.90 |
|
|
|
| 39 |
B2 |
1386 |
1378 |
1.18 |
|
|
|
| 40 |
B2 |
1318 |
1311 |
0.46 |
|
|
|
| 41 |
B2 |
1269 |
1262 |
5.41 |
|
|
|
| 42 |
B2 |
1211 |
1204 |
7.68 |
|
|
|
| 43 |
B2 |
1151 |
1145 |
0.00 |
|
|
|
| 44 |
B2 |
1044 |
1038 |
1.87 |
|
|
|
| 45 |
B2 |
1013 |
1007 |
7.65 |
|
|
|
| 46 |
B2 |
739 |
735 |
0.03 |
|
|
|
| 47 |
B2 |
487 |
484 |
1.43 |
|
|
|
| 48 |
B2 |
327 |
325 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31210.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31029.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.024 |
|
|
|
| 2 |
C |
0.032 |
|
|
|
| 3 |
C |
0.019 |
|
|
|
| 4 |
C |
0.019 |
|
|
|
| 5 |
C |
0.023 |
|
|
|
| 6 |
C |
0.023 |
|
|
|
| 7 |
C |
0.021 |
|
|
|
| 8 |
C |
0.021 |
|
|
|
| 9 |
C |
-0.006 |
|
|
|
| 10 |
C |
-0.006 |
|
|
|
| 11 |
H |
-0.003 |
|
|
|
| 12 |
H |
-0.026 |
|
|
|
| 13 |
H |
-0.009 |
|
|
|
| 14 |
H |
-0.009 |
|
|
|
| 15 |
H |
-0.020 |
|
|
|
| 16 |
H |
-0.020 |
|
|
|
| 17 |
H |
-0.042 |
|
|
|
| 18 |
H |
-0.042 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.922 |
0.922 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-63.153 |
0.000 |
0.000 |
| y |
0.000 |
-51.263 |
0.000 |
| z |
0.000 |
0.000 |
-51.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.615 |
0.000 |
0.000 |
| y |
0.000 |
6.221 |
0.000 |
| z |
0.000 |
0.000 |
5.394 |
|
| Polar |
|---|
| 3z2-r2 | 10.788 |
|---|
| x2-y2 | -11.890 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.681 |
0.000 |
0.000 |
| y |
0.000 |
18.537 |
0.000 |
| z |
0.000 |
0.000 |
27.627 |
<r2> (average value of r
2) Å
2
| <r2> |
370.828 |
| (<r2>)1/2 |
19.257 |