return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3C(OH)=NH (Ethaninidic acid)

using model chemistry: CISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CISD/cc-pVDZ
 hartrees
Energy at 0K-208.542156
Energy at 298.15K-208.548616
HF Energy-207.976615
Nuclear repulsion energy121.943334
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3939 3631 70.12      
2 A' 3656 3370 2.66      
3 A' 3268 3013 12.21      
4 A' 3160 2913 6.84      
5 A' 1864 1718 243.15      
6 A' 1537 1417 50.16      
7 A' 1512 1394 50.65      
8 A' 1433 1321 0.86      
9 A' 1331 1227 100.39      
10 A' 1170 1079 187.12      
11 A' 1053 970 25.02      
12 A' 917 845 0.90      
13 A' 569 524 42.94      
14 A' 434 400 1.69      
15 A" 3243 2990 6.86      
16 A" 1513 1394 7.28      
17 A" 1108 1021 8.87      
18 A" 893 823 24.57      
19 A" 645 595 123.13      
20 A" 538 495 28.56      
21 A" 133 123 0.90      

Unscaled Zero Point Vibrational Energy (zpe) 16957.2 cm-1
Scaled (by 0.9218) Zero Point Vibrational Energy (zpe) 15631.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/cc-pVDZ
ABC
0.36422 0.31172 0.17341

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.134 0.000
C2 0.893 -1.075 0.000
N3 0.315 1.361 0.000
O4 -1.300 -0.223 0.000
H5 1.948 -0.789 0.000
H6 0.679 -1.687 0.884
H7 0.679 -1.687 -0.884
H8 1.326 1.461 0.000
H9 -1.795 0.602 0.000

Atom - Atom Distances (Å)
  C1 C2 N3 O4 H5 H6 H7 H8 H9
C11.50281.26611.34802.15552.13532.13531.87591.8550
C21.50282.50292.35201.09351.09631.09632.57253.1677
N31.26612.50292.26172.69913.19403.19401.01622.2425
O41.34802.35202.26173.29662.61562.61563.11950.9619
H52.15551.09352.69913.29661.78831.78832.33393.9929
H62.13531.09633.19402.61561.78831.76813.33303.4845
H72.13531.09633.19402.61561.78831.76813.33303.4845
H81.87592.57251.01623.11952.33393.33303.33303.2375
H91.85503.16772.24250.96193.99293.48453.48453.2375

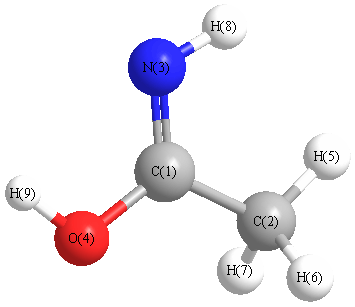
picture of Ethaninidic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 111.267 C1 C2 H6 109.490
C1 C2 H7 109.490 C1 N3 H8 110.076
C1 O4 H9 105.625 C2 C1 N3 129.155
C2 C1 O4 111.070 N3 C1 O4 119.775
H5 C2 H6 109.511 H5 C2 H7 109.511
H6 C2 H7 107.491
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability