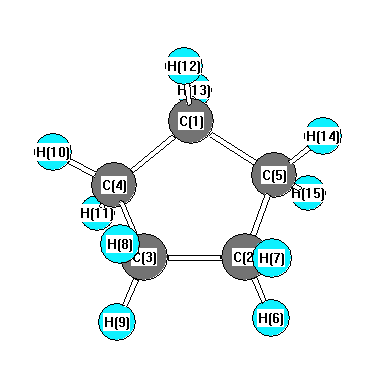
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3044 |
3027 |
82.94 |
|
|
|
| 2 |
A |
3032 |
3014 |
13.37 |
|
|
|
| 3 |
A |
3028 |
3011 |
26.69 |
|
|
|
| 4 |
A |
3025 |
3008 |
17.37 |
|
|
|
| 5 |
A |
3017 |
2999 |
29.72 |
|
|
|
| 6 |
A |
2991 |
2973 |
46.62 |
|
|
|
| 7 |
A |
2974 |
2957 |
51.67 |
|
|
|
| 8 |
A |
2973 |
2956 |
0.21 |
|
|
|
| 9 |
A |
2963 |
2946 |
37.38 |
|
|
|
| 10 |
A |
2958 |
2941 |
18.24 |
|
|
|
| 11 |
A |
1444 |
1435 |
0.05 |
|
|
|
| 12 |
A |
1416 |
1407 |
3.61 |
|
|
|
| 13 |
A |
1415 |
1407 |
2.45 |
|
|
|
| 14 |
A |
1410 |
1401 |
3.72 |
|
|
|
| 15 |
A |
1403 |
1395 |
0.14 |
|
|
|
| 16 |
A |
1294 |
1286 |
0.82 |
|
|
|
| 17 |
A |
1291 |
1283 |
0.91 |
|
|
|
| 18 |
A |
1289 |
1282 |
0.00 |
|
|
|
| 19 |
A |
1269 |
1261 |
0.41 |
|
|
|
| 20 |
A |
1242 |
1234 |
0.02 |
|
|
|
| 21 |
A |
1231 |
1224 |
0.87 |
|
|
|
| 22 |
A |
1186 |
1179 |
0.10 |
|
|
|
| 23 |
A |
1179 |
1173 |
0.43 |
|
|
|
| 24 |
A |
1155 |
1148 |
0.15 |
|
|
|
| 25 |
A |
1138 |
1131 |
0.02 |
|
|
|
| 26 |
A |
1036 |
1030 |
0.04 |
|
|
|
| 27 |
A |
1020 |
1014 |
0.01 |
|
|
|
| 28 |
A |
964 |
958 |
0.29 |
|
|
|
| 29 |
A |
936 |
930 |
0.95 |
|
|
|
| 30 |
A |
905 |
899 |
1.34 |
|
|
|
| 31 |
A |
892 |
887 |
1.45 |
|
|
|
| 32 |
A |
880 |
875 |
0.78 |
|
|
|
| 33 |
A |
848 |
843 |
0.83 |
|
|
|
| 34 |
A |
797 |
793 |
0.14 |
|
|
|
| 35 |
A |
738 |
734 |
0.91 |
|
|
|
| 36 |
A |
599 |
595 |
0.16 |
|
|
|
| 37 |
A |
530 |
527 |
1.88 |
|
|
|
| 38 |
A |
269 |
267 |
0.01 |
|
|
|
| 39 |
A |
17 |
17 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29897.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29724.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.034 |
|
|
|
| 2 |
C |
-0.034 |
|
|
|
| 3 |
C |
-0.029 |
|
|
|
| 4 |
C |
-0.034 |
|
|
|
| 5 |
C |
-0.034 |
|
|
|
| 6 |
H |
0.022 |
|
|
|
| 7 |
H |
0.011 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.019 |
|
|
|
| 10 |
H |
0.022 |
|
|
|
| 11 |
H |
0.011 |
|
|
|
| 12 |
H |
0.011 |
|
|
|
| 13 |
H |
0.021 |
|
|
|
| 14 |
H |
0.021 |
|
|
|
| 15 |
H |
0.011 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.012 |
-0.000 |
0.000 |
0.012 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.868 |
-0.000 |
0.000 |
| y |
-0.000 |
-33.874 |
-0.022 |
| z |
0.000 |
-0.022 |
-32.783 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.540 |
-0.000 |
0.000 |
| y |
-0.000 |
-0.548 |
-0.022 |
| z |
0.000 |
-0.022 |
1.087 |
|
| Polar |
|---|
| 3z2-r2 | 2.175 |
|---|
| x2-y2 | 0.005 |
|---|
| xy | -0.000 |
|---|
| xz | 0.000 |
|---|
| yz | -0.022 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.517 |
0.000 |
0.000 |
| y |
0.000 |
8.534 |
0.007 |
| z |
0.000 |
0.007 |
7.602 |
<r2> (average value of r
2) Å
2
| <r2> |
116.726 |
| (<r2>)1/2 |
10.804 |