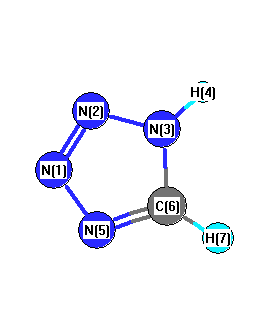
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3532 |
3511 |
63.87 |
|
|
|
| 2 |
A' |
3216 |
3197 |
1.30 |
|
|
|
| 3 |
A' |
1445 |
1436 |
11.08 |
|
|
|
| 4 |
A' |
1407 |
1399 |
6.86 |
|
|
|
| 5 |
A' |
1237 |
1230 |
7.67 |
|
|
|
| 6 |
A' |
1204 |
1197 |
6.34 |
|
|
|
| 7 |
A' |
1105 |
1098 |
14.30 |
|
|
|
| 8 |
A' |
1024 |
1018 |
22.33 |
|
|
|
| 9 |
A' |
1013 |
1007 |
18.00 |
|
|
|
| 10 |
A' |
970 |
964 |
2.37 |
|
|
|
| 11 |
A' |
929 |
924 |
4.00 |
|
|
|
| 12 |
A" |
807 |
803 |
20.42 |
|
|
|
| 13 |
A" |
720 |
715 |
7.77 |
|
|
|
| 14 |
A" |
672 |
668 |
6.75 |
|
|
|
| 15 |
A" |
566 |
563 |
66.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9922.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 9864.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.187 |
|
|
|
| 2 |
N |
-0.028 |
|
|
|
| 3 |
N |
-0.071 |
|
|
|
| 4 |
N |
-0.089 |
|
|
|
| 5 |
N |
-0.172 |
|
|
|
| 6 |
H |
0.054 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.296 |
4.530 |
0.000 |
5.078 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.608 |
1.666 |
0.000 |
| y |
1.666 |
-25.914 |
0.000 |
| z |
0.000 |
0.000 |
-27.977 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.662 |
1.666 |
0.000 |
| y |
1.666 |
2.378 |
0.000 |
| z |
0.000 |
0.000 |
-0.716 |
|
| Polar |
|---|
| 3z2-r2 | -1.432 |
|---|
| x2-y2 | -2.694 |
|---|
| xy | 1.666 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.845 |
0.182 |
0.000 |
| y |
0.182 |
5.727 |
0.000 |
| z |
0.000 |
0.000 |
2.417 |
<r2> (average value of r
2) Å
2
| <r2> |
71.021 |
| (<r2>)1/2 |
8.427 |