return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H14 (cycloheptane)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-274.343220
Energy at 298.15K-274.360030
HF Energy-273.246860
Nuclear repulsion energy327.264724
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3093 2967 3.82      
2 A 3085 2960 22.76      
3 A 3079 2954 78.25      
4 A 3054 2930 20.09      
5 A 3042 2919 10.41      
6 A 3036 2913 24.86      
7 A 3036 2912 0.68      
8 A 1507 1446 0.30      
9 A 1493 1432 4.08      
10 A 1482 1422 0.10      
11 A 1480 1419 0.02      
12 A 1416 1358 0.39      
13 A 1414 1356 1.12      
14 A 1385 1329 3.78      
15 A 1319 1265 0.06      
16 A 1308 1255 1.88      
17 A 1239 1188 0.67      
18 A 1225 1175 0.36      
19 A 1149 1103 0.04      
20 A 1056 1013 0.03      
21 A 1045 1002 0.01      
22 A 971 931 1.18      
23 A 862 827 0.06      
24 A 851 816 0.94      
25 A 752 722 0.01      
26 A 516 495 0.07      
27 A 397 380 0.00      
28 A 309 297 0.00      
29 A 171 164 0.00      
30 B 3101 2975 122.09      
31 B 3091 2965 87.70      
32 B 3085 2960 51.96      
33 B 3075 2950 9.79      
34 B 3043 2920 38.82      
35 B 3040 2917 32.13      
36 B 3031 2908 21.78      
37 B 1499 1438 13.48      
38 B 1485 1425 5.32      
39 B 1478 1418 0.75      
40 B 1409 1351 1.61      
41 B 1401 1344 0.80      
42 B 1393 1337 2.87      
43 B 1357 1302 0.14      
44 B 1310 1257 1.37      
45 B 1237 1187 0.40      
46 B 1210 1161 1.08      
47 B 1164 1117 0.32      
48 B 1043 1000 0.17      
49 B 1031 989 0.35      
50 B 974 934 1.96      
51 B 877 841 0.18      
52 B 827 793 1.49      
53 B 739 709 0.70      
54 B 492 472 0.19      
55 B 333 319 0.02      
56 B 275 264 0.01      
57 B 132 127 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 43948.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 42164.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.10370 0.09413 0.05678

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.781
C2 0.725 1.093 0.966
C3 -0.725 -1.093 0.966
C4 0.000 1.575 -0.308
C5 0.000 -1.575 -0.308
C6 0.302 0.707 -1.546
C7 -0.302 -0.707 -1.546
H8 -0.744 0.478 2.449
H9 0.744 -0.478 2.449
H10 1.730 0.731 0.670
H11 -1.730 -0.731 0.670
H12 0.904 1.953 1.638
H13 -0.904 -1.953 1.638
H14 0.330 2.607 -0.532
H15 -0.330 -2.607 -0.532
H16 -1.094 1.633 -0.134
H17 1.094 -1.633 -0.134
H18 1.404 0.631 -1.649
H19 -1.404 -0.631 -1.649
H20 -0.055 1.233 -2.453
H21 0.055 -1.233 -2.453

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21
C11.54391.54392.61612.61613.41453.41451.10801.10802.18182.18182.15732.15733.50103.50102.74442.74443.75913.75914.41024.4102
C21.54392.62241.54293.04382.57603.25622.17562.16041.10853.07191.10693.51932.16644.12892.19322.96262.74053.78663.50954.1890
C31.54392.62243.04381.54293.25622.57602.16042.17563.07191.10853.51931.10694.12892.16642.96262.19323.78662.74054.18903.5095
C42.61611.54293.04383.14961.54122.61373.05913.51692.15873.04362.17964.12981.10704.20121.10943.39372.15852.93842.17253.5340
C52.61613.04381.54293.14962.61371.54123.51693.05913.04362.15874.12982.17964.20121.10703.39371.10942.93842.15853.53402.1725
C63.41452.57603.25621.54122.61371.53814.13574.19032.63593.33223.47214.32132.15373.52352.19032.84561.10982.17071.10782.1564
C73.41453.25622.57602.61371.54121.53814.19034.13573.33222.63594.32133.47213.52352.15372.84562.19032.17071.10982.15641.1078
H81.10802.17562.16043.05913.51694.13574.19031.76793.05722.36622.35582.56783.81764.31002.85143.80884.62884.29605.00735.2532
H91.10802.16042.17563.51693.05914.19034.13571.76792.36623.05722.56782.35584.31003.81763.80882.85144.29604.62885.25325.0073
H102.18181.10853.07192.15873.04362.63593.33223.05722.36623.75531.76463.88342.63134.10263.07152.57642.34324.12913.63184.0512
H112.18183.07191.10853.04362.15873.33222.63592.36623.05723.75533.88341.76464.10262.63132.57643.07154.12912.34324.05123.6318
H122.15731.10693.51932.17964.12983.47214.32132.35582.56781.76463.88344.30512.33855.19962.69074.00503.57814.77634.26375.2551
H132.15733.51931.10694.12982.17964.32133.47212.56782.35583.88341.76464.30515.19962.33854.00502.69074.77633.57815.25514.2637
H143.50102.16644.12891.10704.20122.15373.52353.81764.31002.63134.10262.33855.19965.25641.77114.32692.51113.83952.39304.3031
H153.50104.12892.16644.20121.10703.52352.15374.31003.81764.10262.63135.19962.33855.25644.32691.77113.83952.51114.30312.3930
H162.74442.19322.96261.10943.39372.19032.84562.85143.80883.07152.57642.69074.00501.77114.32693.93133.08842.74122.57193.8618
H172.74442.96262.19323.39371.10942.84562.19033.80882.85142.57643.07154.00502.69074.32691.77113.93132.74123.08843.86182.5719
H183.75912.74053.78662.15852.93841.10982.17074.62884.29602.34324.12913.57814.77632.51113.83953.08842.74123.07881.77202.4376
H193.75913.78662.74052.93842.15852.17071.10984.29604.62884.12912.34324.77633.57813.83952.51112.74123.08843.07882.43761.7720
H204.41023.50954.18902.17253.53401.10782.15645.00735.25323.63184.05124.26375.25512.39304.30312.57193.86181.77202.43762.4693
H214.41024.18903.50953.53402.17252.15641.10785.25325.00734.05123.63185.25514.26374.30312.39303.86182.57192.43761.77202.4693

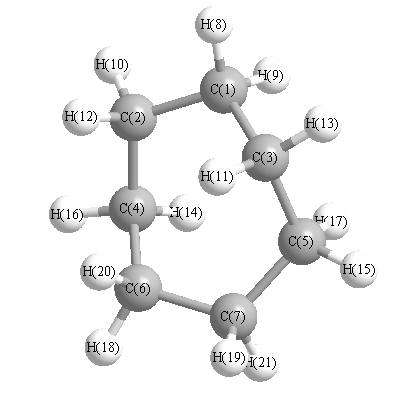
picture of cycloheptane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 115.889 C1 C2 H10 109.595
C1 C2 H12 107.804 C1 C3 C5 115.889
C1 C3 H11 109.595 C1 C3 H13 107.804
C2 C1 C3 116.272 C2 C1 H8 109.144
C2 C1 H9 107.977 C2 C4 C6 113.288
C2 C4 H14 108.558 C2 C4 H16 110.500
C3 C1 H8 107.977 C3 C1 H9 109.144
C3 C5 C7 113.288 C3 C5 H15 108.558
C3 C5 H17 110.500 C4 C2 H10 107.890
C4 C2 H12 109.586 C4 C6 C7 116.160
C4 C6 H18 107.913 C4 C6 H20 109.101
C5 C3 H11 107.890 C5 C3 H13 109.586
C5 C7 C6 116.160 C5 C7 H19 107.913
C5 C7 H21 109.101 C6 C4 H14 107.706
C6 C4 H16 110.389 C6 C7 H19 109.052
C6 C7 H21 108.076 C7 C5 H15 107.706
C7 C5 H17 110.389 C7 C6 H18 109.052
C7 C6 H20 108.076 H8 C1 H9 105.839
H10 C2 H12 105.599 H11 C3 H13 105.599
H14 C4 H16 106.087 H15 C5 H17 106.087
H18 C6 H20 106.087 H19 C7 H21 106.087
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability