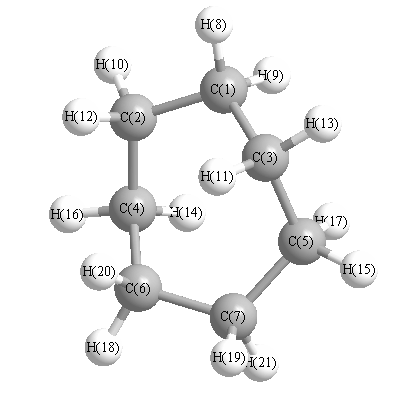
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3000 |
2983 |
3.76 |
|
|
|
| 2 |
A |
2992 |
2975 |
32.03 |
|
|
|
| 3 |
A |
2987 |
2969 |
71.55 |
|
|
|
| 4 |
A |
2959 |
2942 |
24.90 |
|
|
|
| 5 |
A |
2949 |
2932 |
1.38 |
|
|
|
| 6 |
A |
2946 |
2929 |
4.90 |
|
|
|
| 7 |
A |
2939 |
2922 |
15.46 |
|
|
|
| 8 |
A |
1428 |
1420 |
0.32 |
|
|
|
| 9 |
A |
1412 |
1403 |
6.03 |
|
|
|
| 10 |
A |
1403 |
1395 |
0.12 |
|
|
|
| 11 |
A |
1400 |
1392 |
0.39 |
|
|
|
| 12 |
A |
1342 |
1335 |
0.75 |
|
|
|
| 13 |
A |
1339 |
1331 |
1.39 |
|
|
|
| 14 |
A |
1319 |
1312 |
3.37 |
|
|
|
| 15 |
A |
1266 |
1259 |
0.02 |
|
|
|
| 16 |
A |
1256 |
1249 |
1.62 |
|
|
|
| 17 |
A |
1185 |
1178 |
0.77 |
|
|
|
| 18 |
A |
1171 |
1164 |
0.14 |
|
|
|
| 19 |
A |
1109 |
1102 |
0.08 |
|
|
|
| 20 |
A |
1014 |
1008 |
0.13 |
|
|
|
| 21 |
A |
1008 |
1002 |
0.00 |
|
|
|
| 22 |
A |
924 |
919 |
1.53 |
|
|
|
| 23 |
A |
829 |
825 |
0.24 |
|
|
|
| 24 |
A |
819 |
814 |
1.94 |
|
|
|
| 25 |
A |
728 |
724 |
0.04 |
|
|
|
| 26 |
A |
504 |
501 |
0.24 |
|
|
|
| 27 |
A |
382 |
380 |
0.00 |
|
|
|
| 28 |
A |
304 |
303 |
0.00 |
|
|
|
| 29 |
A |
172 |
171 |
0.00 |
|
|
|
| 30 |
B |
3007 |
2990 |
99.00 |
|
|
|
| 31 |
B |
2997 |
2980 |
99.27 |
|
|
|
| 32 |
B |
2992 |
2975 |
46.16 |
|
|
|
| 33 |
B |
2983 |
2965 |
8.73 |
|
|
|
| 34 |
B |
2951 |
2934 |
46.92 |
|
|
|
| 35 |
B |
2944 |
2927 |
19.58 |
|
|
|
| 36 |
B |
2939 |
2922 |
27.04 |
|
|
|
| 37 |
B |
1420 |
1411 |
15.46 |
|
|
|
| 38 |
B |
1406 |
1398 |
7.16 |
|
|
|
| 39 |
B |
1399 |
1390 |
0.82 |
|
|
|
| 40 |
B |
1338 |
1331 |
3.61 |
|
|
|
| 41 |
B |
1329 |
1321 |
1.20 |
|
|
|
| 42 |
B |
1323 |
1315 |
1.65 |
|
|
|
| 43 |
B |
1293 |
1286 |
0.16 |
|
|
|
| 44 |
B |
1259 |
1252 |
1.86 |
|
|
|
| 45 |
B |
1181 |
1174 |
0.37 |
|
|
|
| 46 |
B |
1161 |
1154 |
1.03 |
|
|
|
| 47 |
B |
1121 |
1115 |
0.50 |
|
|
|
| 48 |
B |
1005 |
999 |
0.22 |
|
|
|
| 49 |
B |
992 |
986 |
0.52 |
|
|
|
| 50 |
B |
933 |
927 |
2.30 |
|
|
|
| 51 |
B |
842 |
837 |
0.25 |
|
|
|
| 52 |
B |
798 |
793 |
2.83 |
|
|
|
| 53 |
B |
707 |
703 |
1.21 |
|
|
|
| 54 |
B |
471 |
468 |
0.48 |
|
|
|
| 55 |
B |
325 |
323 |
0.07 |
|
|
|
| 56 |
B |
270 |
268 |
0.05 |
|
|
|
| 57 |
B |
105 |
104 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42287.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 42042.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.020 |
|
|
|
| 2 |
C |
-0.032 |
|
|
|
| 3 |
C |
-0.032 |
|
|
|
| 4 |
C |
-0.018 |
|
|
|
| 5 |
C |
-0.018 |
|
|
|
| 6 |
C |
-0.025 |
|
|
|
| 7 |
C |
-0.025 |
|
|
|
| 8 |
H |
0.016 |
|
|
|
| 9 |
H |
0.016 |
|
|
|
| 10 |
H |
0.015 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
H |
0.009 |
|
|
|
| 13 |
H |
0.009 |
|
|
|
| 14 |
H |
0.011 |
|
|
|
| 15 |
H |
0.011 |
|
|
|
| 16 |
H |
0.010 |
|
|
|
| 17 |
H |
0.010 |
|
|
|
| 18 |
H |
0.015 |
|
|
|
| 19 |
H |
0.015 |
|
|
|
| 20 |
H |
0.009 |
|
|
|
| 21 |
H |
0.009 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.005 |
0.005 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.793 |
-0.097 |
0.000 |
| y |
-0.097 |
-46.721 |
0.000 |
| z |
0.000 |
0.000 |
-47.141 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.138 |
-0.097 |
0.000 |
| y |
-0.097 |
0.246 |
0.000 |
| z |
0.000 |
0.000 |
-0.384 |
|
| Polar |
|---|
| 3z2-r2 | -0.768 |
|---|
| x2-y2 | -0.072 |
|---|
| xy | -0.097 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.524 |
0.424 |
0.000 |
| y |
0.424 |
12.031 |
0.000 |
| z |
0.000 |
0.000 |
12.448 |
<r2> (average value of r
2) Å
2
| <r2> |
231.403 |
| (<r2>)1/2 |
15.212 |