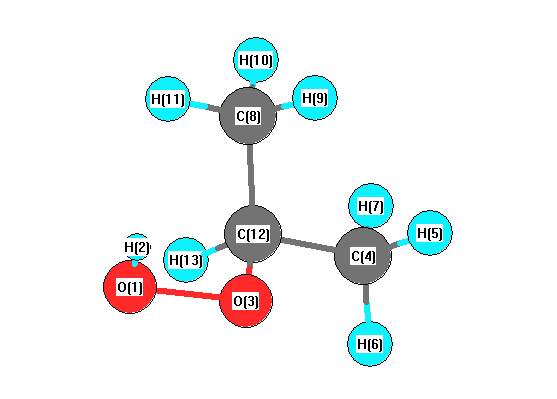
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3589 |
3568 |
25.91 |
|
|
|
| 2 |
A |
3078 |
3060 |
13.54 |
|
|
|
| 3 |
A |
3069 |
3051 |
13.88 |
|
|
|
| 4 |
A |
3067 |
3049 |
24.96 |
|
|
|
| 5 |
A |
3055 |
3038 |
14.71 |
|
|
|
| 6 |
A |
2981 |
2964 |
14.33 |
|
|
|
| 7 |
A |
2971 |
2954 |
17.59 |
|
|
|
| 8 |
A |
2933 |
2916 |
32.84 |
|
|
|
| 9 |
A |
1436 |
1428 |
5.40 |
|
|
|
| 10 |
A |
1414 |
1406 |
2.70 |
|
|
|
| 11 |
A |
1405 |
1397 |
2.72 |
|
|
|
| 12 |
A |
1399 |
1391 |
2.19 |
|
|
|
| 13 |
A |
1339 |
1331 |
4.89 |
|
|
|
| 14 |
A |
1336 |
1329 |
28.83 |
|
|
|
| 15 |
A |
1307 |
1300 |
0.41 |
|
|
|
| 16 |
A |
1301 |
1293 |
20.23 |
|
|
|
| 17 |
A |
1266 |
1259 |
40.72 |
|
|
|
| 18 |
A |
1144 |
1137 |
4.03 |
|
|
|
| 19 |
A |
1134 |
1127 |
23.16 |
|
|
|
| 20 |
A |
1100 |
1094 |
16.35 |
|
|
|
| 21 |
A |
916 |
910 |
2.33 |
|
|
|
| 22 |
A |
894 |
889 |
0.35 |
|
|
|
| 23 |
A |
883 |
878 |
3.85 |
|
|
|
| 24 |
A |
868 |
863 |
7.74 |
|
|
|
| 25 |
A |
803 |
798 |
3.91 |
|
|
|
| 26 |
A |
485 |
482 |
5.24 |
|
|
|
| 27 |
A |
457 |
454 |
3.74 |
|
|
|
| 28 |
A |
338 |
336 |
0.60 |
|
|
|
| 29 |
A |
279 |
277 |
1.98 |
|
|
|
| 30 |
A |
258 |
257 |
32.18 |
|
|
|
| 31 |
A |
213 |
212 |
77.22 |
|
|
|
| 32 |
A |
195 |
193 |
4.84 |
|
|
|
| 33 |
A |
140 |
140 |
5.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23526.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23390.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.147 |
|
|
|
| 2 |
H |
0.171 |
|
|
|
| 3 |
O |
-0.214 |
|
|
|
| 4 |
C |
-0.032 |
|
|
|
| 5 |
H |
0.038 |
|
|
|
| 6 |
H |
0.043 |
|
|
|
| 7 |
H |
0.038 |
|
|
|
| 8 |
C |
-0.030 |
|
|
|
| 9 |
H |
0.028 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.027 |
|
|
|
| 12 |
C |
0.033 |
|
|
|
| 13 |
H |
0.013 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.677 |
1.301 |
0.621 |
1.593 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.273 |
-2.292 |
-3.054 |
| y |
-2.292 |
-32.534 |
0.549 |
| z |
-3.054 |
0.549 |
-30.917 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.452 |
-2.292 |
-3.054 |
| y |
-2.292 |
-2.439 |
0.549 |
| z |
-3.054 |
0.549 |
-0.013 |
|
| Polar |
|---|
| 3z2-r2 | -0.026 |
|---|
| x2-y2 | 3.261 |
|---|
| xy | -2.292 |
|---|
| xz | -3.054 |
|---|
| yz | 0.549 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.990 |
-0.410 |
-0.107 |
| y |
-0.410 |
6.518 |
0.063 |
| z |
-0.107 |
0.063 |
5.932 |
<r2> (average value of r
2) Å
2
| <r2> |
133.775 |
| (<r2>)1/2 |
11.566 |