return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6 (1-Methylcyclopropene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-155.467037
Energy at 298.15K 
HF Energy-154.877777
Nuclear repulsion energy107.094870
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3299 3165 0.16      
2 A' 3156 3027 16.66      
3 A' 3071 2946 62.53      
4 A' 3059 2934 28.85      
5 A' 1862 1786 11.00      
6 A' 1524 1462 6.02      
7 A' 1482 1422 7.19      
8 A' 1410 1353 0.76      
9 A' 1216 1167 2.27      
10 A' 1100 1055 13.47      
11 A' 1073 1030 20.76      
12 A' 990 950 5.60      
13 A' 945 906 11.59      
14 A' 683 655 0.62      
15 A' 319 306 1.35      
16 A" 3142 3014 45.59      
17 A" 3134 3007 21.63      
18 A" 1479 1418 6.86      
19 A" 1114 1069 1.57      
20 A" 1045 1003 0.26      
21 A" 992 951 1.85      
22 A" 711 682 27.15      
23 A" 290 278 10.25      
24 A" 153 147 0.39      

Unscaled Zero Point Vibrational Energy (zpe) 18624.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17867.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.67588 0.20775 0.16951

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.180 0.000
C2 -0.062 -1.340 0.000
C3 1.162 -0.433 0.000
C4 -0.831 1.417 0.000
H5 -0.346 -1.874 0.923
H6 -0.346 -1.874 -0.923
H7 2.251 -0.394 0.000
H8 -0.200 2.322 0.000
H9 -1.485 1.438 -0.891
H10 -1.485 1.438 0.891

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.52191.31381.49052.27862.27862.32272.15132.14082.1408
C21.52191.52382.86301.10361.10362.49903.66513.24673.2467
C31.31381.52382.71962.28082.28081.08943.07323.36223.3622
C41.49052.86302.71963.45283.45283.57471.10351.10531.1053
H52.27861.10362.28083.45281.84693.12804.29913.94503.5033
H62.27861.10362.28083.45281.84693.12804.29913.50333.9450
H72.32272.49901.08943.57473.12803.12803.65804.25554.2555
H82.15133.66513.07321.10354.29914.29913.65801.79651.7965
H92.14083.24673.36221.10533.94503.50334.25551.79651.7812
H102.14083.24673.36221.10533.50333.94504.25551.79651.7812

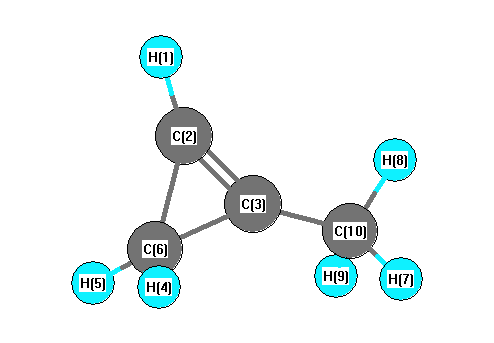
picture of 1-Methylcyclopropene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 51.109 H1 C2 C6 119.582
C2 C3 C6 24.873 C2 C3 C10 72.478
C2 C6 C3 35.504 C2 C6 H4 49.561
C2 C6 H5 33.198 C3 C2 C6 119.623
C3 C6 H4 51.858 C3 C6 H5 66.117
C3 C10 H7 9.456 C3 C10 H8 65.096
C3 C10 H9 74.640 H4 C6 H5 74.488
C6 C3 C10 86.474 H7 C10 H8 58.645
H7 C10 H9 77.920 H8 C10 H9 60.281
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability