return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6 (1-Methylcyclopropene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-155.462880
Energy at 298.15K 
HF Energy-154.878089
Nuclear repulsion energy107.192000
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3308 3165 0.17      
2 A' 3166 3029 15.59      
3 A' 3080 2946 61.58      
4 A' 3067 2934 26.96      
5 A' 1880 1798 9.57      
6 A' 1529 1462 5.87      
7 A' 1484 1420 7.28      
8 A' 1414 1353 0.53      
9 A' 1224 1171 2.12      
10 A' 1106 1058 14.24      
11 A' 1081 1034 17.80      
12 A' 995 951 5.63      
13 A' 949 908 11.93      
14 A' 689 659 0.64      
15 A' 321 307 1.31      
16 A" 3153 3016 44.78      
17 A" 3145 3009 20.61      
18 A" 1481 1417 7.04      
19 A" 1119 1070 1.56      
20 A" 1050 1004 0.32      
21 A" 995 952 1.91      
22 A" 724 692 27.01      
23 A" 292 279 10.45      
24 A" 154 147 0.39      

Unscaled Zero Point Vibrational Energy (zpe) 18700.7 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 17890.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.67708 0.20813 0.16982

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.180 0.000
C2 -0.058 -1.339 0.000
C3 1.162 -0.428 0.000
C4 -0.835 1.413 0.000
H5 -0.339 -1.874 0.923
H6 -0.339 -1.874 -0.923
H7 2.250 -0.386 0.000
H8 -0.207 2.320 0.000
H9 -1.489 1.433 -0.890
H10 -1.489 1.433 0.890

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.52021.31121.48972.27722.27722.32002.15042.14022.1402
C21.52021.52232.86041.10321.10322.49683.66263.24443.2444
C31.31121.52232.71642.27912.27911.08903.07033.35913.3591
C41.48972.86042.71643.45073.45073.57161.10311.10491.1049
H52.27721.10322.27913.45071.84613.12534.29683.94303.5015
H62.27721.10322.27913.45071.84613.12534.29683.50153.9430
H72.32002.49681.08903.57163.12533.12533.65524.25244.2524
H82.15043.66263.07031.10314.29684.29683.65521.79571.7957
H92.14023.24443.35911.10493.94303.50154.25241.79571.7804
H102.14023.24443.35911.10493.50153.94304.25241.79571.7804

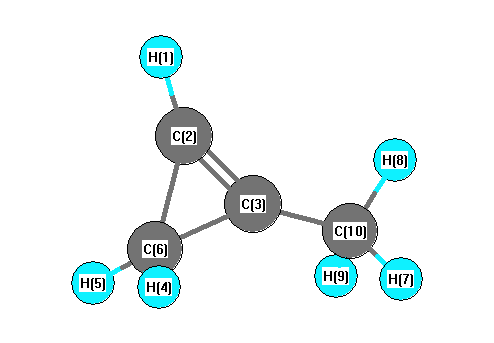
picture of 1-Methylcyclopropene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 51.058 H1 C2 C6 119.616
C2 C3 C6 24.885 C2 C3 C10 72.503
C2 C6 C3 35.496 C2 C6 H4 49.524
C2 C6 H5 33.208 C3 C2 C6 119.619
C3 C6 H4 51.821 C3 C6 H5 66.109
C3 C10 H7 9.454 C3 C10 H8 65.091
C3 C10 H9 74.632 H4 C6 H5 74.485
C6 C3 C10 86.517 H7 C10 H8 58.646
H7 C10 H9 77.916 H8 C10 H9 60.281
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability