return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H3ClO (Furan, 2-chloro)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-688.432439
Energy at 298.15K-688.436249
HF Energy-687.556591
Nuclear repulsion energy263.598646
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3321 3186 0.05      
2 A' 3306 3171 0.58      
3 A' 3284 3151 1.81      
4 A' 1642 1575 19.83      
5 A' 1562 1498 58.56      
6 A' 1422 1364 11.45      
7 A' 1255 1204 36.20      
8 A' 1223 1173 36.65      
9 A' 1159 1112 4.91      
10 A' 1096 1051 16.54      
11 A' 1023 982 24.86      
12 A' 935 897 26.64      
13 A' 893 856 1.14      
14 A' 498 478 3.39      
15 A' 313 300 0.30      
16 A" 860 825        
17 A" 809 776 17.19      
18 A" 750 720 50.85      
19 A" 629 604 6.02      
20 A" 599 574 4.47      
21 A" 228 219 1.96      

Unscaled Zero Point Vibrational Energy (zpe) 13402.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12858.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.30428 0.07069 0.05737

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.267 0.000
C2 1.318 -0.096 0.000
C3 1.307 -1.547 0.000
C4 -0.010 -1.915 0.000
O5 -0.829 -0.812 0.000
Cl6 -0.757 1.809 0.000
H7 2.175 0.576 0.000
H8 2.168 -2.215 0.000
H9 -0.521 -2.876 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 Cl6 H7 H8 H9
C11.36662.23492.18201.36001.71852.19673.29513.1858
C21.36661.45112.25292.26302.81671.08892.28363.3337
C32.23491.45111.36762.25863.93962.29341.09022.2602
C42.18202.25291.36761.37433.79873.31402.19881.0882
O51.36002.26302.25861.37432.62173.30893.30932.0877
Cl61.71852.81673.93963.79872.62173.18084.97504.6915
H72.19671.08892.29343.31403.30893.18082.79114.3803
H83.29512.28361.09022.19883.30934.97502.79112.7691
H93.18583.33372.26021.08822.08774.69154.38032.7691

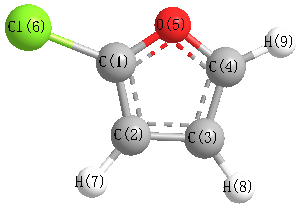
picture of Furan, 2-chloro state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 104.924 C1 C2 H7 126.551
C1 O5 C4 105.877 C2 C1 O5 112.195
C2 C1 Cl6 131.510 C2 C3 C4 106.084
C2 C3 H8 127.370 C3 C2 H7 128.525
C3 C4 O5 110.919 C3 C4 H9 133.632
C4 C3 H8 126.546 O5 C1 Cl6 116.295
O5 C4 H9 115.449
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability