return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHFClCHClF (ethane, 1,2-dichloro-1,2-difluoro-)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CI 1AG
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-1195.727028
Energy at 298.15K-1195.730568
HF Energy-1194.778208
Nuclear repulsion energy374.813752
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3161 3025 0.00      
2 Ag 1434 1372 0.00      
3 Ag 1335 1277 0.00      
4 Ag 1174 1123 0.00      
5 Ag 1119 1070 0.00      
6 Ag 858 821 0.00      
7 Ag 526 504 0.00      
8 Ag 384 367 0.00      
9 Ag 276 264 0.00      
10 Au 3171 3034 16.42      
11 Au 1344 1286 32.52      
12 Au 1250 1196 41.95      
13 Au 1174 1123 255.33      
14 Au 807 772 181.47      
15 Au 410 392 3.79      
16 Au 372 356 29.06      
17 Au 175 168 1.34      
18 Au 75 72 0.75      

Unscaled Zero Point Vibrational Energy (zpe) 9523.0 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 9110.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.13391 0.04883 0.03696

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Ci

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.765
C2 0.000 0.000 0.765
H3 1.029 0.000 -1.156
H4 -1.029 0.000 1.156
F5 -0.666 -1.099 -1.207
F6 0.666 1.099 1.207
Cl7 -0.820 1.466 -1.360
Cl8 0.820 -1.466 1.360

Atom - Atom Distances (Å)
  C1 C2 H3 H4 F5 F6 Cl7 Cl8
C11.53061.10062.17971.35852.35401.78192.7087
C21.53062.17971.10062.35401.35852.70871.7819
H31.10062.17973.09542.02012.63172.36852.9192
H42.17971.10063.09542.63172.02012.91922.3685
F51.35852.35402.02012.63173.52572.57402.9885
F62.35401.35852.63172.02013.52572.98852.5740
Cl71.78192.70872.36852.91922.57402.98854.3222
Cl82.70871.78192.91922.36852.98852.57404.3222

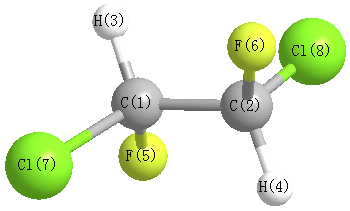
picture of ethane, 1,2-dichloro-1,2-difluoro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H4 110.809 C1 C2 F6 108.986
C1 C2 Cl8 109.480 C2 C1 H3 110.809
C2 C1 F5 108.986 C2 C1 Cl7 109.480
H3 C1 F5 110.021 H3 C1 Cl7 108.170
H4 C2 F6 110.021 H4 C2 Cl8 108.170
F5 C1 Cl7 109.355 F6 C2 Cl8 109.355
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability