Jump to
S1C2
Energy calculated at ROMP2/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -957.789922 |
| Energy at 298.15K | |
| HF Energy | -957.396258 |
| Nuclear repulsion energy | 125.363130 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at ROMP2/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.675 |
| H2 |
0.000 |
0.000 |
1.763 |
| Cl3 |
0.000 |
1.479 |
-0.171 |
| Cl4 |
0.000 |
-1.479 |
-0.171 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0880 | 1.7041 | 1.7041 |
H2 | 1.0880 | | 2.4351 | 2.4351 | Cl3 | 1.7041 | 2.4351 | | 2.9580 | Cl4 | 1.7041 | 2.4351 | 2.9580 | |
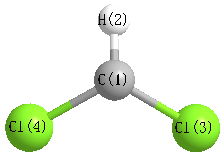 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.785 |
|
Cl3 |
C1 |
Cl4 |
120.431 |
| Cl4 |
C1 |
H2 |
119.785 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROMP2/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.056 |
|
|
|
| 2 |
H |
0.133 |
|
|
|
| 3 |
Cl |
-0.039 |
|
|
|
| 4 |
Cl |
-0.039 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
100.491 |
| (<r2>)1/2 |
10.025 |
Jump to
S1C1
Energy calculated at ROMP2/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -957.791204 |
| Energy at 298.15K | -957.792076 |
| HF Energy | -957.399596 |
| Nuclear repulsion energy | 125.158575 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at ROMP2/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.013 |
0.703 |
0.000 |
| H2 |
-0.528 |
1.652 |
0.000 |
| Cl3 |
0.013 |
-0.173 |
1.475 |
| Cl4 |
0.013 |
-0.173 |
-1.475 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.0926 | 1.7153 | 1.7153 |
H2 | 1.0926 | | 2.4081 | 2.4081 | Cl3 | 1.7153 | 2.4081 | | 2.9498 | Cl4 | 1.7153 | 2.4081 | 2.9498 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
116.335 |
|
Cl3 |
C1 |
Cl4 |
118.602 |
| Cl4 |
C1 |
H2 |
116.335 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROMP2/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.027 |
|
|
|
| 2 |
H |
0.130 |
|
|
|
| 3 |
Cl |
-0.052 |
|
|
|
| 4 |
Cl |
-0.052 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
100.330 |
| (<r2>)1/2 |
10.016 |
